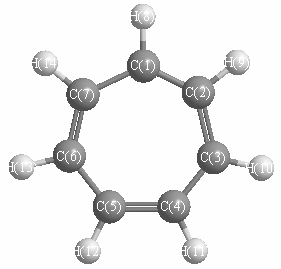
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3210 |
3072 |
4.21 |
|
|
|
| 2 |
A1 |
3198 |
3061 |
37.02 |
|
|
|
| 3 |
A1 |
3166 |
3030 |
3.34 |
|
|
|
| 4 |
A1 |
3158 |
3023 |
6.22 |
|
|
|
| 5 |
A1 |
1646 |
1576 |
3.74 |
|
|
|
| 6 |
A1 |
1557 |
1490 |
11.41 |
|
|
|
| 7 |
A1 |
1479 |
1415 |
5.45 |
|
|
|
| 8 |
A1 |
1282 |
1227 |
0.79 |
|
|
|
| 9 |
A1 |
1194 |
1143 |
3.15 |
|
|
|
| 10 |
A1 |
976 |
934 |
2.57 |
|
|
|
| 11 |
A1 |
913 |
874 |
0.76 |
|
|
|
| 12 |
A1 |
857 |
821 |
0.30 |
|
|
|
| 13 |
A1 |
434 |
416 |
0.25 |
|
|
|
| 14 |
A2 |
1007 |
964 |
0.00 |
|
|
|
| 15 |
A2 |
856 |
819 |
0.00 |
|
|
|
| 16 |
A2 |
766 |
733 |
0.00 |
|
|
|
| 17 |
A2 |
570 |
545 |
0.00 |
|
|
|
| 18 |
A2 |
146 |
140 |
0.00 |
|
|
|
| 19 |
B1 |
1021 |
977 |
0.00 |
|
|
|
| 20 |
B1 |
998 |
955 |
0.35 |
|
|
|
| 21 |
B1 |
777 |
743 |
0.99 |
|
|
|
| 22 |
B1 |
652 |
624 |
69.91 |
|
|
|
| 23 |
B1 |
518 |
496 |
37.30 |
|
|
|
| 24 |
B1 |
288 |
275 |
0.00 |
|
|
|
| 25 |
B2 |
3202 |
3065 |
28.35 |
|
|
|
| 26 |
B2 |
3183 |
3047 |
16.95 |
|
|
|
| 27 |
B2 |
3160 |
3024 |
5.37 |
|
|
|
| 28 |
B2 |
1662 |
1591 |
9.29 |
|
|
|
| 29 |
B2 |
1488 |
1424 |
2.58 |
|
|
|
| 30 |
B2 |
1414 |
1353 |
0.01 |
|
|
|
| 31 |
B2 |
1302 |
1246 |
0.28 |
|
|
|
| 32 |
B2 |
1248 |
1194 |
1.83 |
|
|
|
| 33 |
B2 |
986 |
943 |
6.19 |
|
|
|
| 34 |
B2 |
902 |
863 |
1.79 |
|
|
|
| 35 |
B2 |
408 |
391 |
0.36 |
|
|
|
| 36 |
B2 |
151i |
144i |
38.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24736.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23675.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.069 |
|
|
|
| 2 |
C |
-0.174 |
|
|
|
| 3 |
C |
-0.174 |
|
|
|
| 4 |
C |
-0.089 |
|
|
|
| 5 |
C |
-0.089 |
|
|
|
| 6 |
C |
-0.135 |
|
|
|
| 7 |
C |
-0.135 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
| 13 |
H |
0.123 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.085 |
0.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.904 |
0.000 |
0.000 |
| y |
0.000 |
-36.970 |
0.000 |
| z |
0.000 |
0.000 |
-36.968 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.935 |
0.000 |
0.000 |
| y |
0.000 |
4.466 |
0.000 |
| z |
0.000 |
0.000 |
4.469 |
|
| Polar |
|---|
| 3z2-r2 | 8.938 |
|---|
| x2-y2 | -8.934 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.775 |
0.000 |
0.000 |
| y |
0.000 |
14.300 |
0.000 |
| z |
0.000 |
0.000 |
14.281 |
<r2> (average value of r
2) Å
2
| <r2> |
184.094 |
| (<r2>)1/2 |
13.568 |