return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-307.804814
Energy at 298.15K-307.810519
HF Energy-306.433589
Nuclear repulsion energy314.008587
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3290 3124 6.83      
2 A1 3253 3089 8.39      
3 A1 3236 3072 7.66      
4 A1 1709 1623 1.84      
5 A1 1545 1467 1.30      
6 A1 1528 1451 3.22      
7 A1 1455 1381 8.56      
8 A1 1199 1139 0.18      
9 A1 1138 1081 0.26      
10 A1 1076 1022 0.59      
11 A1 1008 957 3.63      
12 A1 820 779 0.49      
13 A1 549 521 0.08      
14 A2 965 916 0.00      
15 A2 920 874 0.00      
16 A2 890 845 0.00      
17 A2 767 728 0.00      
18 A2 589 559 0.00      
19 A2 298 283 0.00      
20 B1 938 890 7.29      
21 B1 768 729 97.92      
22 B1 725 689 8.25      
23 B1 366 347 3.75      
24 B1 237 225 12.27      
25 B2 3255 3090 1.50      
26 B2 3244 3080 18.06      
27 B2 3215 3052 0.00      
28 B2 1659 1575 0.56      
29 B2 1473 1398 3.68      
30 B2 1311 1245 4.83      
31 B2 1263 1200 14.12      
32 B2 1122 1065 1.98      
33 B2 1065 1011 1.24      
34 B2 873 828 0.49      
35 B2 653 620 1.24      
36 B2 418 397 3.50      

Unscaled Zero Point Vibrational Energy (zpe) 24409.3 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23174.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.16256 0.07406 0.05088

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.436 -0.619
C2 0.000 0.685 -1.832
C3 0.000 -0.685 -1.832
C4 0.000 -1.436 -0.619
C5 0.000 -0.675 2.031
C6 0.000 0.675 2.031
C7 0.000 0.712 0.524
C8 0.000 -0.712 0.524
H9 0.000 2.512 -0.648
H10 0.000 1.216 -2.768
H11 0.000 -1.216 -2.768
H12 0.000 -2.512 -0.648
H13 0.000 -1.438 2.788
H14 0.000 1.438 2.788

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.42672.44262.87113.38822.75721.35272.43271.07662.16053.41283.94744.45723.4069
C21.42671.36912.44264.09583.86352.35622.73882.17751.07622.11823.40875.08444.6811
C32.44261.36911.42673.86354.09582.73882.35623.40872.11821.07622.17754.68115.0844
C42.87112.44261.42672.75723.38822.43271.35273.94743.41282.16051.07663.40694.4572
C53.38824.09583.86352.75721.35062.04861.50794.16355.15864.82993.24801.07452.2450
C62.75723.86354.09583.38821.35061.50792.04863.24804.82995.15864.16352.24501.0745
C71.35272.35622.73882.43272.04861.50791.42402.14753.33053.81503.43003.12242.3776
C82.43272.73882.35621.35271.50792.04861.42403.43003.81503.33052.14752.37763.1224
H91.07662.17753.40873.94744.16353.24802.14753.43002.48544.28835.02355.23513.5993
H102.16051.07622.11823.41285.15864.82993.33053.81502.48542.43114.28836.15745.5606
H113.41282.11821.07622.16054.82995.15863.81503.33054.28832.43112.48545.56066.1574
H123.94743.40872.17751.07663.24804.16353.43002.14755.02354.28832.48543.59935.2351
H134.45725.08444.68113.40691.07452.24503.12242.37765.23516.15745.56063.59932.8768
H143.40694.68115.08444.45722.24501.07452.37763.12243.59935.56066.15745.23512.8768

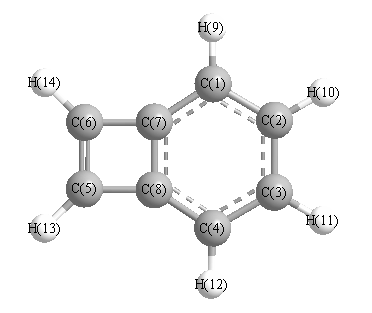
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.759 C1 C2 H10 118.680
C1 C7 C6 149.058 C1 C7 N8 122.337
C2 C1 C7 115.904 C2 C1 H9 120.233
C2 C3 C4 121.759 C2 C3 H11 119.561
C3 C2 H10 119.561 C3 C4 N8 115.904
C3 C4 H12 120.233 C4 C3 H11 118.680
C4 N8 C5 149.058 C4 N8 C7 122.337
C5 C6 C7 91.395 C5 C6 H14 135.249
C5 N8 C7 88.605 C6 C5 N8 91.395
C6 C5 H13 135.249 C6 C7 N8 88.605
C7 C1 H9 123.864 C7 C6 H14 133.356
N8 C4 H12 123.864 N8 C5 H13 133.356
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability