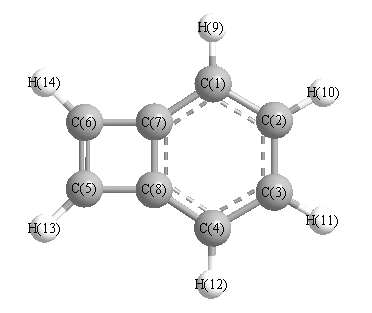
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3261 |
3128 |
8.59 |
|
|
|
| 2 |
A1 |
3219 |
3088 |
11.90 |
|
|
|
| 3 |
A1 |
3203 |
3073 |
8.76 |
|
|
|
| 4 |
A1 |
1726 |
1656 |
3.28 |
|
|
|
| 5 |
A1 |
1594 |
1529 |
1.49 |
|
|
|
| 6 |
A1 |
1521 |
1458 |
3.28 |
|
|
|
| 7 |
A1 |
1454 |
1394 |
10.51 |
|
|
|
| 8 |
A1 |
1192 |
1143 |
0.05 |
|
|
|
| 9 |
A1 |
1117 |
1072 |
0.19 |
|
|
|
| 10 |
A1 |
1074 |
1030 |
0.54 |
|
|
|
| 11 |
A1 |
992 |
951 |
5.42 |
|
|
|
| 12 |
A1 |
828 |
794 |
0.55 |
|
|
|
| 13 |
A1 |
551 |
529 |
0.24 |
|
|
|
| 14 |
A2 |
987 |
947 |
0.00 |
|
|
|
| 15 |
A2 |
938 |
900 |
0.00 |
|
|
|
| 16 |
A2 |
893 |
856 |
0.00 |
|
|
|
| 17 |
A2 |
791 |
759 |
0.00 |
|
|
|
| 18 |
A2 |
592 |
568 |
0.00 |
|
|
|
| 19 |
A2 |
297 |
285 |
0.00 |
|
|
|
| 20 |
B1 |
932 |
894 |
8.36 |
|
|
|
| 21 |
B1 |
759 |
728 |
108.33 |
|
|
|
| 22 |
B1 |
729 |
700 |
0.01 |
|
|
|
| 23 |
B1 |
363 |
348 |
2.88 |
|
|
|
| 24 |
B1 |
235 |
225 |
12.43 |
|
|
|
| 25 |
B2 |
3231 |
3099 |
1.42 |
|
|
|
| 26 |
B2 |
3211 |
3080 |
25.76 |
|
|
|
| 27 |
B2 |
3191 |
3061 |
0.00 |
|
|
|
| 28 |
B2 |
1676 |
1608 |
0.37 |
|
|
|
| 29 |
B2 |
1471 |
1411 |
4.29 |
|
|
|
| 30 |
B2 |
1313 |
1259 |
5.21 |
|
|
|
| 31 |
B2 |
1264 |
1212 |
12.90 |
|
|
|
| 32 |
B2 |
1106 |
1061 |
2.64 |
|
|
|
| 33 |
B2 |
1063 |
1019 |
0.51 |
|
|
|
| 34 |
B2 |
882 |
846 |
0.63 |
|
|
|
| 35 |
B2 |
658 |
631 |
0.79 |
|
|
|
| 36 |
B2 |
424 |
407 |
4.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24367.5 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 23373.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.149 |
|
|
|
| 2 |
C |
-0.104 |
|
|
|
| 3 |
C |
-0.104 |
|
|
|
| 4 |
C |
-0.149 |
|
|
|
| 5 |
C |
-0.133 |
|
|
|
| 6 |
C |
-0.133 |
|
|
|
| 7 |
C |
0.084 |
|
|
|
| 8 |
C |
0.084 |
|
|
|
| 9 |
H |
0.094 |
|
|
|
| 10 |
H |
0.101 |
|
|
|
| 11 |
H |
0.101 |
|
|
|
| 12 |
H |
0.094 |
|
|
|
| 13 |
H |
0.107 |
|
|
|
| 14 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.732 |
0.732 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.367 |
0.000 |
0.000 |
| y |
0.000 |
-40.644 |
0.000 |
| z |
0.000 |
0.000 |
-40.212 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.940 |
0.000 |
0.000 |
| y |
0.000 |
4.646 |
0.000 |
| z |
0.000 |
0.000 |
5.293 |
|
| Polar |
|---|
| 3z2-r2 | 10.587 |
|---|
| x2-y2 | -9.724 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.270 |
0.000 |
0.000 |
| y |
0.000 |
13.800 |
0.000 |
| z |
0.000 |
0.000 |
16.866 |
<r2> (average value of r
2) Å
2
| <r2> |
218.609 |
| (<r2>)1/2 |
14.785 |