Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.392039 |
| Energy at 298.15K | -235.404370 |
| HF Energy | -234.263315 |
| Nuclear repulsion energy | 228.370053 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3190 |
3029 |
28.32 |
|
|
|
| 2 |
A |
3177 |
3016 |
18.03 |
|
|
|
| 3 |
A |
3164 |
3004 |
27.07 |
|
|
|
| 4 |
A |
3163 |
3003 |
6.75 |
|
|
|
| 5 |
A |
3158 |
2998 |
37.86 |
|
|
|
| 6 |
A |
3140 |
2981 |
14.18 |
|
|
|
| 7 |
A |
3124 |
2966 |
17.10 |
|
|
|
| 8 |
A |
3100 |
2943 |
6.83 |
|
|
|
| 9 |
A |
3089 |
2933 |
40.98 |
|
|
|
| 10 |
A |
3086 |
2930 |
11.04 |
|
|
|
| 11 |
A |
3080 |
2924 |
27.24 |
|
|
|
| 12 |
A |
3064 |
2909 |
18.74 |
|
|
|
| 13 |
A |
1745 |
1657 |
0.49 |
|
|
|
| 14 |
A |
1532 |
1454 |
6.39 |
|
|
|
| 15 |
A |
1527 |
1450 |
6.77 |
|
|
|
| 16 |
A |
1519 |
1442 |
10.07 |
|
|
|
| 17 |
A |
1515 |
1439 |
1.03 |
|
|
|
| 18 |
A |
1506 |
1430 |
6.16 |
|
|
|
| 19 |
A |
1496 |
1420 |
2.07 |
|
|
|
| 20 |
A |
1423 |
1351 |
1.60 |
|
|
|
| 21 |
A |
1422 |
1350 |
3.96 |
|
|
|
| 22 |
A |
1395 |
1324 |
0.87 |
|
|
|
| 23 |
A |
1355 |
1286 |
1.32 |
|
|
|
| 24 |
A |
1344 |
1276 |
0.44 |
|
|
|
| 25 |
A |
1330 |
1262 |
0.10 |
|
|
|
| 26 |
A |
1300 |
1235 |
2.66 |
|
|
|
| 27 |
A |
1264 |
1200 |
0.51 |
|
|
|
| 28 |
A |
1196 |
1135 |
0.05 |
|
|
|
| 29 |
A |
1135 |
1077 |
1.31 |
|
|
|
| 30 |
A |
1122 |
1066 |
2.45 |
|
|
|
| 31 |
A |
1083 |
1028 |
0.49 |
|
|
|
| 32 |
A |
1082 |
1027 |
1.70 |
|
|
|
| 33 |
A |
1055 |
1001 |
5.79 |
|
|
|
| 34 |
A |
1025 |
973 |
34.08 |
|
|
|
| 35 |
A |
939 |
891 |
5.28 |
|
|
|
| 36 |
A |
920 |
873 |
1.70 |
|
|
|
| 37 |
A |
884 |
839 |
0.30 |
|
|
|
| 38 |
A |
800 |
760 |
0.17 |
|
|
|
| 39 |
A |
757 |
719 |
2.07 |
|
|
|
| 40 |
A |
536 |
509 |
0.12 |
|
|
|
| 41 |
A |
395 |
375 |
1.22 |
|
|
|
| 42 |
A |
311 |
296 |
0.78 |
|
|
|
| 43 |
A |
290 |
275 |
1.21 |
|
|
|
| 44 |
A |
262 |
249 |
0.04 |
|
|
|
| 45 |
A |
218 |
207 |
0.72 |
|
|
|
| 46 |
A |
149 |
142 |
0.29 |
|
|
|
| 47 |
A |
93 |
88 |
0.06 |
|
|
|
| 48 |
A |
79 |
75 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36768.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 34907.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.145 |
0.231 |
-0.157 |
| C2 |
1.774 |
-0.343 |
-0.254 |
| C3 |
0.732 |
0.051 |
0.474 |
| C4 |
-0.645 |
-0.502 |
0.361 |
| C5 |
-1.648 |
0.529 |
-0.138 |
| C6 |
-3.057 |
-0.027 |
-0.211 |
| H7 |
3.182 |
1.024 |
0.583 |
| H8 |
3.870 |
-0.529 |
0.121 |
| H9 |
3.466 |
0.642 |
-1.112 |
| H10 |
1.627 |
-1.141 |
-0.968 |
| H11 |
0.877 |
0.850 |
1.188 |
| H12 |
-0.980 |
-0.866 |
1.333 |
| H13 |
-0.640 |
-1.358 |
-0.312 |
| H14 |
-1.331 |
0.879 |
-1.118 |
| H15 |
-1.626 |
1.394 |
0.522 |
| H16 |
-3.762 |
0.715 |
-0.571 |
| H17 |
-3.098 |
-0.881 |
-0.882 |
| H18 |
-3.391 |
-0.360 |
0.768 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4893 | 2.5007 | 3.8948 | 4.8019 | 6.2075 | 1.0853 | 1.0874 | 1.0874 | 2.2010 | 2.7087 | 4.5209 | 4.1080 | 4.6236 | 4.9575 | 6.9364 | 6.3827 | 6.6270 |
C2 | 1.4893 | | 1.3310 | 2.5011 | 3.5336 | 4.8419 | 2.1332 | 2.1374 | 2.1376 | 1.0807 | 2.0757 | 3.2211 | 2.6195 | 3.4475 | 3.8968 | 5.6458 | 4.9423 | 5.2650 | C3 | 2.5007 | 1.3310 | | 1.4881 | 2.5032 | 3.8512 | 2.6383 | 3.2111 | 3.2155 | 2.0736 | 1.0818 | 2.1232 | 2.1178 | 2.7341 | 2.7144 | 4.6616 | 4.1686 | 4.1533 | C4 | 3.8948 | 2.5011 | 1.4881 | | 1.5227 | 2.5242 | 4.1257 | 4.5218 | 4.5141 | 2.7080 | 2.1977 | 1.0905 | 1.0888 | 2.1364 | 2.1416 | 3.4741 | 2.7762 | 2.7793 | C5 | 4.8019 | 3.5336 | 2.5032 | 1.5227 | | 1.5169 | 4.9081 | 5.6248 | 5.2069 | 3.7683 | 2.8697 | 2.1343 | 2.1467 | 1.0874 | 1.0881 | 2.1664 | 2.1556 | 2.1558 | C6 | 6.2075 | 4.8419 | 3.8512 | 2.5242 | 1.5169 | | 6.3764 | 6.9536 | 6.6187 | 4.8733 | 4.2667 | 2.7208 | 2.7611 | 2.1493 | 2.1461 | 1.0853 | 1.0865 | 1.0865 | H7 | 1.0853 | 2.1332 | 2.6383 | 4.1257 | 4.9081 | 6.3764 | | 1.7605 | 1.7606 | 3.0838 | 2.3891 | 4.6315 | 4.5914 | 4.8252 | 4.8226 | 7.0461 | 6.7242 | 6.7189 | H8 | 1.0874 | 2.1374 | 3.2111 | 4.5218 | 5.6248 | 6.9536 | 1.7605 | | 1.7479 | 2.5680 | 3.4644 | 5.0107 | 4.6064 | 5.5294 | 5.8372 | 7.7644 | 7.0493 | 7.2917 | H9 | 1.0874 | 2.1376 | 3.2155 | 4.5141 | 5.2069 | 6.6187 | 1.7606 | 1.7479 | | 2.5655 | 3.4692 | 5.2929 | 4.6366 | 4.8031 | 5.4004 | 7.2487 | 6.7426 | 7.1797 | H10 | 2.2010 | 1.0807 | 2.0736 | 2.7080 | 3.7683 | 4.8733 | 3.0838 | 2.5680 | 2.5655 | | 3.0290 | 3.4877 | 2.3696 | 3.5846 | 4.3849 | 5.7133 | 4.7329 | 5.3661 | H11 | 2.7087 | 2.0757 | 1.0818 | 2.1977 | 2.8697 | 4.2667 | 2.3891 | 3.4644 | 3.4692 | 3.0290 | | 2.5326 | 3.0707 | 3.1927 | 2.6467 | 4.9634 | 4.8047 | 4.4557 | H12 | 4.5209 | 3.2211 | 2.1232 | 1.0905 | 2.1343 | 2.7208 | 4.6315 | 5.0107 | 5.2929 | 3.4877 | 2.5326 | | 1.7506 | 3.0285 | 2.4867 | 3.7238 | 3.0649 | 2.5273 | H13 | 4.1080 | 2.6195 | 2.1178 | 1.0888 | 2.1467 | 2.7611 | 4.5914 | 4.6064 | 4.6366 | 2.3696 | 3.0707 | 1.7506 | | 2.4757 | 3.0405 | 3.7569 | 2.5682 | 3.1191 | H14 | 4.6236 | 3.4475 | 2.7341 | 2.1364 | 1.0874 | 2.1493 | 4.8252 | 5.5294 | 4.8031 | 3.5846 | 3.1927 | 3.0285 | 2.4757 | | 1.7440 | 2.4969 | 2.5047 | 3.0543 | H15 | 4.9575 | 3.8968 | 2.7144 | 2.1416 | 1.0881 | 2.1461 | 4.8226 | 5.8372 | 5.4004 | 4.3849 | 2.6467 | 2.4867 | 3.0405 | 1.7440 | | 2.4938 | 3.0521 | 2.5001 | H16 | 6.9364 | 5.6458 | 4.6616 | 3.4741 | 2.1664 | 1.0853 | 7.0461 | 7.7644 | 7.2487 | 5.7133 | 4.9634 | 3.7238 | 3.7569 | 2.4969 | 2.4938 | | 1.7565 | 1.7568 | H17 | 6.3827 | 4.9423 | 4.1686 | 2.7762 | 2.1556 | 1.0865 | 6.7242 | 7.0493 | 6.7426 | 4.7329 | 4.8047 | 3.0649 | 2.5682 | 2.5047 | 3.0521 | 1.7565 | | 1.7545 | H18 | 6.6270 | 5.2650 | 4.1533 | 2.7793 | 2.1558 | 1.0865 | 6.7189 | 7.2917 | 7.1797 | 5.3661 | 4.4557 | 2.5273 | 3.1191 | 3.0543 | 2.5001 | 1.7568 | 1.7545 | |
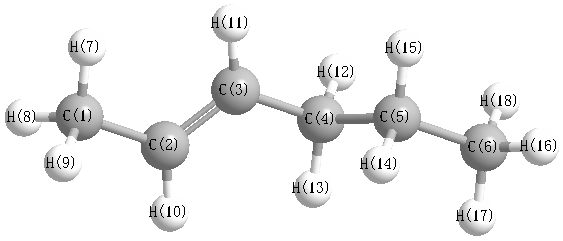 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.832 |
|
C1 |
C2 |
H10 |
116.947 |
| C2 |
C1 |
H7 |
110.924 |
|
C2 |
C1 |
H8 |
111.139 |
| C2 |
C1 |
H9 |
111.153 |
|
C2 |
C3 |
C4 |
124.960 |
| C2 |
C3 |
H11 |
118.341 |
|
C3 |
C2 |
H10 |
118.221 |
| C3 |
C4 |
C5 |
112.483 |
|
C3 |
C4 |
H12 |
109.897 |
| C3 |
C4 |
H13 |
109.572 |
|
C4 |
C3 |
H11 |
116.680 |
| C4 |
C5 |
C6 |
112.288 |
|
C4 |
C5 |
H14 |
108.725 |
| C4 |
C5 |
H15 |
109.088 |
|
C5 |
C4 |
H12 |
108.383 |
| C5 |
C4 |
H13 |
109.450 |
|
C5 |
C6 |
H16 |
111.646 |
| C5 |
C6 |
H17 |
110.705 |
|
C5 |
C6 |
H18 |
110.719 |
| C6 |
C5 |
H14 |
110.145 |
|
C6 |
C5 |
H15 |
109.847 |
| H7 |
C1 |
H8 |
108.244 |
|
H7 |
C1 |
H9 |
108.254 |
| H8 |
C1 |
H9 |
106.973 |
|
H12 |
C4 |
H13 |
106.893 |
| H14 |
C5 |
H15 |
106.574 |
|
H16 |
C6 |
H17 |
107.953 |
| H16 |
C6 |
H18 |
107.979 |
|
H17 |
C6 |
H18 |
107.690 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
