Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.783455 |
| Energy at 298.15K | -235.795659 |
| HF Energy | -235.783455 |
| Nuclear repulsion energy | 227.804595 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3009 |
52.85 |
|
|
|
| 2 |
A |
3129 |
2995 |
21.71 |
|
|
|
| 3 |
A |
3127 |
2993 |
34.14 |
|
|
|
| 4 |
A |
3125 |
2991 |
4.75 |
|
|
|
| 5 |
A |
3120 |
2986 |
50.04 |
|
|
|
| 6 |
A |
3094 |
2961 |
19.61 |
|
|
|
| 7 |
A |
3089 |
2957 |
16.74 |
|
|
|
| 8 |
A |
3068 |
2936 |
8.40 |
|
|
|
| 9 |
A |
3050 |
2920 |
47.02 |
|
|
|
| 10 |
A |
3048 |
2918 |
15.80 |
|
|
|
| 11 |
A |
3042 |
2911 |
36.22 |
|
|
|
| 12 |
A |
3023 |
2893 |
23.35 |
|
|
|
| 13 |
A |
1764 |
1688 |
0.47 |
|
|
|
| 14 |
A |
1508 |
1443 |
6.82 |
|
|
|
| 15 |
A |
1497 |
1433 |
6.33 |
|
|
|
| 16 |
A |
1492 |
1428 |
1.40 |
|
|
|
| 17 |
A |
1491 |
1427 |
10.17 |
|
|
|
| 18 |
A |
1477 |
1414 |
5.34 |
|
|
|
| 19 |
A |
1475 |
1412 |
3.01 |
|
|
|
| 20 |
A |
1410 |
1350 |
0.87 |
|
|
|
| 21 |
A |
1409 |
1349 |
3.97 |
|
|
|
| 22 |
A |
1385 |
1326 |
1.00 |
|
|
|
| 23 |
A |
1342 |
1284 |
1.30 |
|
|
|
| 24 |
A |
1336 |
1279 |
0.39 |
|
|
|
| 25 |
A |
1318 |
1261 |
0.04 |
|
|
|
| 26 |
A |
1287 |
1232 |
4.75 |
|
|
|
| 27 |
A |
1250 |
1197 |
0.39 |
|
|
|
| 28 |
A |
1183 |
1132 |
0.05 |
|
|
|
| 29 |
A |
1119 |
1071 |
0.59 |
|
|
|
| 30 |
A |
1105 |
1057 |
3.87 |
|
|
|
| 31 |
A |
1068 |
1023 |
0.53 |
|
|
|
| 32 |
A |
1066 |
1020 |
2.49 |
|
|
|
| 33 |
A |
1043 |
998 |
7.24 |
|
|
|
| 34 |
A |
1009 |
966 |
33.65 |
|
|
|
| 35 |
A |
928 |
889 |
6.58 |
|
|
|
| 36 |
A |
907 |
868 |
1.92 |
|
|
|
| 37 |
A |
874 |
837 |
0.21 |
|
|
|
| 38 |
A |
787 |
753 |
0.40 |
|
|
|
| 39 |
A |
741 |
709 |
2.79 |
|
|
|
| 40 |
A |
525 |
502 |
0.12 |
|
|
|
| 41 |
A |
387 |
371 |
1.26 |
|
|
|
| 42 |
A |
303 |
290 |
0.86 |
|
|
|
| 43 |
A |
281 |
269 |
1.65 |
|
|
|
| 44 |
A |
240 |
229 |
0.06 |
|
|
|
| 45 |
A |
211 |
202 |
0.64 |
|
|
|
| 46 |
A |
141 |
135 |
0.27 |
|
|
|
| 47 |
A |
97 |
93 |
0.08 |
|
|
|
| 48 |
A |
78 |
75 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36296.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34739.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.164 |
0.212 |
-0.156 |
| C2 |
1.792 |
-0.367 |
-0.213 |
| C3 |
0.736 |
0.110 |
0.429 |
| C4 |
-0.639 |
-0.463 |
0.369 |
| C5 |
-1.673 |
0.518 |
-0.175 |
| C6 |
-3.077 |
-0.059 |
-0.185 |
| H7 |
3.195 |
1.095 |
0.478 |
| H8 |
3.882 |
-0.511 |
0.233 |
| H9 |
3.515 |
0.495 |
-1.150 |
| H10 |
1.666 |
-1.252 |
-0.830 |
| H11 |
0.860 |
0.997 |
1.045 |
| H12 |
-0.951 |
-0.770 |
1.372 |
| H13 |
-0.633 |
-1.366 |
-0.243 |
| H14 |
-1.384 |
0.814 |
-1.184 |
| H15 |
-1.653 |
1.429 |
0.426 |
| H16 |
-3.799 |
0.655 |
-0.576 |
| H17 |
-3.128 |
-0.955 |
-0.803 |
| H18 |
-3.394 |
-0.336 |
0.820 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4893 | 2.4988 | 3.8973 | 4.8461 | 6.2460 | 1.0879 | 1.0908 | 1.0908 | 2.1991 | 2.7142 | 4.4973 | 4.1120 | 4.7007 | 5.0019 | 6.9894 | 6.4314 | 6.6520 |
C2 | 1.4893 | | 1.3246 | 2.5018 | 3.5766 | 4.8788 | 2.1402 | 2.1412 | 2.1414 | 1.0858 | 2.0764 | 3.1934 | 2.6231 | 3.5250 | 3.9374 | 5.6958 | 4.9905 | 5.2880 | C3 | 2.4988 | 1.3246 | | 1.4908 | 2.5170 | 3.8657 | 2.6488 | 3.2120 | 3.2185 | 2.0743 | 1.0867 | 2.1235 | 2.1223 | 2.7553 | 2.7291 | 4.6774 | 4.1934 | 4.1723 | C4 | 3.8973 | 2.5018 | 1.4908 | | 1.5251 | 2.5323 | 4.1393 | 4.5228 | 4.5248 | 2.7154 | 2.1980 | 1.0941 | 1.0913 | 2.1439 | 2.1467 | 3.4828 | 2.7950 | 2.7943 | C5 | 4.8461 | 3.5766 | 2.5170 | 1.5251 | | 1.5175 | 4.9449 | 5.6637 | 5.2783 | 3.8352 | 2.8514 | 2.1383 | 2.1527 | 1.0905 | 1.0915 | 2.1682 | 2.1633 | 2.1631 | C6 | 6.2460 | 4.8788 | 3.8657 | 2.5323 | 1.5175 | | 6.4108 | 6.9854 | 6.6843 | 4.9329 | 4.2567 | 2.7293 | 2.7720 | 2.1505 | 2.1475 | 1.0883 | 1.0895 | 1.0896 | H7 | 1.0879 | 2.1402 | 2.6488 | 4.1393 | 4.9449 | 6.4108 | | 1.7638 | 1.7640 | 3.0906 | 2.4049 | 4.6326 | 4.6070 | 4.8789 | 4.8595 | 7.0866 | 6.7690 | 6.7505 | H8 | 1.0908 | 2.1412 | 3.2120 | 4.5228 | 5.6637 | 6.9854 | 1.7638 | | 1.7488 | 2.5662 | 3.4735 | 4.9713 | 4.6192 | 5.6114 | 5.8679 | 7.8109 | 7.0997 | 7.3010 | H9 | 1.0908 | 2.1414 | 3.2185 | 4.5248 | 5.2783 | 6.6843 | 1.7640 | 1.7488 | | 2.5629 | 3.4808 | 5.2816 | 4.6351 | 4.9089 | 5.4826 | 7.3381 | 6.8078 | 7.2313 | H10 | 2.1991 | 1.0858 | 2.0743 | 2.7154 | 3.8352 | 4.9329 | 3.0906 | 2.5662 | 2.5629 | | 3.0363 | 3.4534 | 2.3755 | 3.7007 | 4.4473 | 5.7941 | 4.8037 | 5.4003 | H11 | 2.7142 | 2.0764 | 1.0867 | 2.1980 | 2.8514 | 4.2567 | 2.4049 | 3.4735 | 3.4808 | 3.0363 | | 2.5508 | 3.0770 | 3.1675 | 2.6234 | 4.9445 | 4.8087 | 4.4627 | H12 | 4.4973 | 3.1934 | 2.1235 | 1.0941 | 2.1383 | 2.7293 | 4.6326 | 4.9713 | 5.2816 | 3.4534 | 2.5508 | | 1.7502 | 3.0378 | 2.4945 | 3.7334 | 3.0829 | 2.5419 | H13 | 4.1120 | 2.6231 | 2.1223 | 1.0913 | 2.1527 | 2.7720 | 4.6070 | 4.6192 | 4.6351 | 2.3755 | 3.0770 | 1.7502 | | 2.4901 | 3.0492 | 3.7710 | 2.5902 | 3.1327 | H14 | 4.7007 | 3.5250 | 2.7553 | 2.1439 | 1.0905 | 2.1505 | 4.8789 | 5.6114 | 4.9089 | 3.7007 | 3.1675 | 3.0378 | 2.4901 | | 1.7443 | 2.4959 | 2.5130 | 3.0621 | H15 | 5.0019 | 3.9374 | 2.7291 | 2.1467 | 1.0915 | 2.1475 | 4.8595 | 5.8679 | 5.4826 | 4.4473 | 2.6234 | 2.4945 | 3.0492 | 1.7443 | | 2.4917 | 3.0604 | 2.5094 | H16 | 6.9894 | 5.6958 | 4.6774 | 3.4828 | 2.1682 | 1.0883 | 7.0866 | 7.8109 | 7.3381 | 5.7941 | 4.9445 | 3.7334 | 3.7710 | 2.4959 | 2.4917 | | 1.7585 | 1.7590 | H17 | 6.4314 | 4.9905 | 4.1934 | 2.7950 | 2.1633 | 1.0895 | 6.7690 | 7.0997 | 6.8078 | 4.8037 | 4.8087 | 3.0829 | 2.5902 | 2.5130 | 3.0604 | 1.7585 | | 1.7572 | H18 | 6.6520 | 5.2880 | 4.1723 | 2.7943 | 2.1631 | 1.0896 | 6.7505 | 7.3010 | 7.2313 | 5.4003 | 4.4627 | 2.5419 | 3.1327 | 3.0621 | 2.5094 | 1.7590 | 1.7572 | |
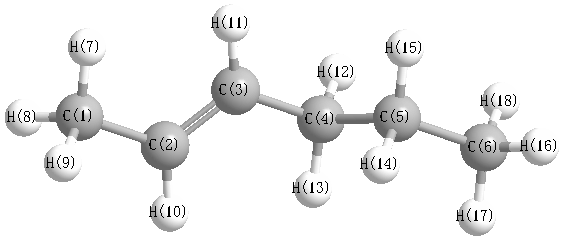 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.151 |
|
C1 |
C2 |
H10 |
116.428 |
| C2 |
C1 |
H7 |
111.342 |
|
C2 |
C1 |
H8 |
111.232 |
| C2 |
C1 |
H9 |
111.260 |
|
C2 |
C3 |
C4 |
125.294 |
| C2 |
C3 |
H11 |
118.551 |
|
C3 |
C2 |
H10 |
118.420 |
| C3 |
C4 |
C5 |
113.135 |
|
C3 |
C4 |
H12 |
109.519 |
| C3 |
C4 |
H13 |
109.587 |
|
C4 |
C3 |
H11 |
116.151 |
| C4 |
C5 |
C6 |
112.664 |
|
C4 |
C5 |
H14 |
108.961 |
| C4 |
C5 |
H15 |
109.126 |
|
C5 |
C4 |
H12 |
108.329 |
| C5 |
C4 |
H13 |
109.612 |
|
C5 |
C6 |
H16 |
111.565 |
| C5 |
C6 |
H17 |
111.092 |
|
C5 |
C6 |
H18 |
111.076 |
| C6 |
C5 |
H14 |
110.014 |
|
C6 |
C5 |
H15 |
109.721 |
| H7 |
C1 |
H8 |
108.109 |
|
H7 |
C1 |
H9 |
108.131 |
| H8 |
C1 |
H9 |
106.569 |
|
H12 |
C4 |
H13 |
106.431 |
| H14 |
C5 |
H15 |
106.139 |
|
H16 |
C6 |
H17 |
107.699 |
| H16 |
C6 |
H18 |
107.737 |
|
H17 |
C6 |
H18 |
107.486 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.332 |
|
|
|
| 2 |
C |
-0.133 |
|
|
|
| 3 |
C |
-0.154 |
|
|
|
| 4 |
C |
-0.195 |
|
|
|
| 5 |
C |
-0.196 |
|
|
|
| 6 |
C |
-0.364 |
|
|
|
| 7 |
H |
0.110 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.107 |
|
|
|
| 14 |
H |
0.115 |
|
|
|
| 15 |
H |
0.111 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.112 |
|
|
|
| 18 |
H |
0.110 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.021 |
0.043 |
-0.048 |
0.068 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.110 |
-0.136 |
-0.645 |
| y |
-0.136 |
-39.833 |
1.215 |
| z |
-0.645 |
1.215 |
-39.966 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.789 |
-0.136 |
-0.645 |
| y |
-0.136 |
-0.295 |
1.215 |
| z |
-0.645 |
1.215 |
-0.494 |
|
| Polar |
|---|
| 3z2-r2 | -0.988 |
|---|
| x2-y2 | 0.723 |
|---|
| xy | -0.136 |
|---|
| xz | -0.645 |
|---|
| yz | 1.215 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.785 |
-0.166 |
-0.795 |
| y |
-0.166 |
9.423 |
0.498 |
| z |
-0.795 |
0.498 |
9.109 |
<r2> (average value of r
2) Å
2
| <r2> |
292.419 |
| (<r2>)1/2 |
17.100 |
