return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCHCHCH3 (2-Butenal)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-230.845463
Energy at 298.15K-230.851372
HF Energy-229.887869
Nuclear repulsion energy153.903245
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3242 3078 2.03      
2 A' 3192 3030 17.68      
3 A' 3175 3014 0.98      
4 A' 3087 2931 9.71      
5 A' 2951 2802 91.83      
6 A' 1766 1676 212.12      
7 A' 1720 1633 7.10      
8 A' 1510 1434 16.52      
9 A' 1440 1368 0.92      
10 A' 1421 1349 3.67      
11 A' 1335 1268 3.51      
12 A' 1284 1219 1.34      
13 A' 1185 1125 49.55      
14 A' 1122 1065 14.58      
15 A' 952 904 22.08      
16 A' 546 518 7.00      
17 A' 462 438 2.55      
18 A' 207 196 6.44      
19 A" 3148 2989 6.68      
20 A" 1505 1429 8.21      
21 A" 1088 1033 3.15      
22 A" 1052 998 5.29      
23 A" 1035 983 32.06      
24 A" 819 778 0.01      
25 A" 303 287 7.55      
26 A" 211 200 0.35      
27 A" 133 126 1.70      

Unscaled Zero Point Vibrational Energy (zpe) 19944.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18935.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.11599 0.07348 0.06982

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.374 -0.467 0.000
C2 0.900 -0.646 0.000
C3 0.000 0.337 0.000
C4 -1.434 0.041 0.000
O5 -2.307 0.879 0.000
H6 -1.671 -1.036 0.000
H7 0.283 1.376 0.000
H8 0.539 -1.666 0.000
H9 2.643 0.584 0.000
H10 2.813 -0.940 0.874
H11 2.813 -0.940 -0.874

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.48532.50623.84214.87034.08502.78682.19241.08411.08681.0868
C21.48531.33262.43323.55072.60012.11431.08132.13342.12432.1243
C32.50621.33261.46442.36972.16271.07722.07362.65423.21073.2107
C43.84212.43321.46441.20931.10332.17512.60904.11294.44644.4464
O54.87033.55072.36971.20932.01762.63743.81724.95835.50345.5034
H64.08502.60012.16271.10332.01763.10452.29764.60794.56994.5699
H72.78682.11431.07722.17512.63743.10453.05252.48893.53953.5395
H82.19241.08132.07362.60903.81722.29763.05253.08012.54292.5429
H91.08412.13342.65424.11294.95834.60792.48893.08011.76461.7646
H101.08682.12433.21074.44645.50344.56993.53952.54291.76461.7483
H111.08682.12433.21074.44645.50344.56993.53952.54291.76461.7483

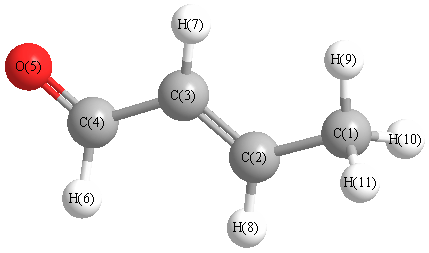
picture of 2-Butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.501 C1 C2 H8 116.461
C2 C1 H9 111.308 C2 C1 H10 110.404
C2 C1 H11 110.404 C2 C3 C4 120.827
C2 C3 H7 122.299 C3 C2 H8 118.038
C3 C4 O5 124.550 C3 C4 H6 114.022
C4 C3 H7 116.874 O5 C4 H6 121.427
H9 C1 H10 108.755 H9 C1 H11 108.755
H10 C1 H11 107.096
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability