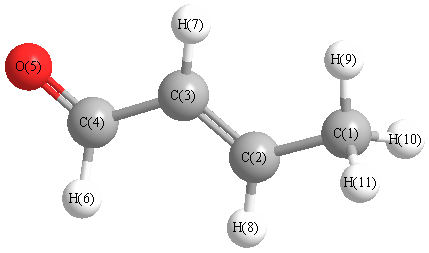
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3191 |
3054 |
5.33 |
|
|
|
| 2 |
A' |
3154 |
3019 |
22.84 |
|
|
|
| 3 |
A' |
3135 |
3001 |
1.29 |
|
|
|
| 4 |
A' |
3049 |
2918 |
10.10 |
|
|
|
| 5 |
A' |
2888 |
2764 |
103.54 |
|
|
|
| 6 |
A' |
1813 |
1735 |
314.79 |
|
|
|
| 7 |
A' |
1729 |
1655 |
58.30 |
|
|
|
| 8 |
A' |
1482 |
1418 |
20.94 |
|
|
|
| 9 |
A' |
1414 |
1353 |
2.18 |
|
|
|
| 10 |
A' |
1406 |
1346 |
4.76 |
|
|
|
| 11 |
A' |
1328 |
1271 |
4.12 |
|
|
|
| 12 |
A' |
1273 |
1218 |
1.09 |
|
|
|
| 13 |
A' |
1172 |
1122 |
60.65 |
|
|
|
| 14 |
A' |
1110 |
1062 |
17.31 |
|
|
|
| 15 |
A' |
942 |
901 |
27.98 |
|
|
|
| 16 |
A' |
543 |
520 |
7.97 |
|
|
|
| 17 |
A' |
457 |
437 |
2.75 |
|
|
|
| 18 |
A' |
201 |
192 |
6.76 |
|
|
|
| 19 |
A" |
3102 |
2969 |
8.73 |
|
|
|
| 20 |
A" |
1476 |
1412 |
8.78 |
|
|
|
| 21 |
A" |
1073 |
1027 |
2.08 |
|
|
|
| 22 |
A" |
1037 |
993 |
1.30 |
|
|
|
| 23 |
A" |
1014 |
970 |
36.74 |
|
|
|
| 24 |
A" |
802 |
767 |
0.01 |
|
|
|
| 25 |
A" |
303 |
290 |
7.87 |
|
|
|
| 26 |
A" |
203 |
194 |
0.34 |
|
|
|
| 27 |
A" |
130 |
125 |
1.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19712.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18867.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.322 |
|
|
|
| 2 |
C |
-0.108 |
|
|
|
| 3 |
C |
-0.133 |
|
|
|
| 4 |
C |
0.121 |
|
|
|
| 5 |
O |
-0.273 |
|
|
|
| 6 |
H |
0.054 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.122 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.541 |
-1.909 |
0.000 |
4.023 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.691 |
3.335 |
0.000 |
| y |
3.335 |
-29.621 |
0.000 |
| z |
0.000 |
0.000 |
-30.863 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.449 |
3.335 |
0.000 |
| y |
3.335 |
3.156 |
0.000 |
| z |
0.000 |
0.000 |
1.293 |
|
| Polar |
|---|
| 3z2-r2 | 2.586 |
|---|
| x2-y2 | -5.070 |
|---|
| xy | 3.335 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.022 |
-1.816 |
0.000 |
| y |
-1.816 |
7.647 |
0.000 |
| z |
0.000 |
0.000 |
4.720 |
<r2> (average value of r
2) Å
2
| <r2> |
159.026 |
| (<r2>)1/2 |
12.611 |