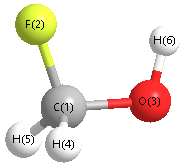
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3868 |
3702 |
47.10 |
67.55 |
0.25 |
0.40 |
| 2 |
A |
3137 |
3003 |
36.94 |
67.38 |
0.47 |
0.64 |
| 3 |
A |
3048 |
2917 |
64.36 |
110.11 |
0.11 |
0.21 |
| 4 |
A |
1535 |
1469 |
1.80 |
5.17 |
0.72 |
0.83 |
| 5 |
A |
1453 |
1391 |
29.93 |
3.06 |
0.74 |
0.85 |
| 6 |
A |
1404 |
1344 |
18.83 |
3.52 |
0.74 |
0.85 |
| 7 |
A |
1275 |
1221 |
7.08 |
4.71 |
0.69 |
0.81 |
| 8 |
A |
1167 |
1117 |
144.69 |
3.15 |
0.23 |
0.37 |
| 9 |
A |
1078 |
1032 |
142.38 |
1.59 |
0.35 |
0.51 |
| 10 |
A |
1037 |
993 |
101.54 |
2.44 |
0.65 |
0.79 |
| 11 |
A |
548 |
525 |
28.66 |
1.60 |
0.59 |
0.74 |
| 12 |
A |
387 |
370 |
113.82 |
3.16 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 9968.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 9540.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.201 |
|
|
|
| 2 |
F |
-0.231 |
|
|
|
| 3 |
O |
-0.302 |
|
|
|
| 4 |
H |
0.040 |
|
|
|
| 5 |
H |
0.074 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.767 |
0.738 |
1.435 |
1.787 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.682 |
1.699 |
-1.920 |
| y |
1.699 |
-15.635 |
-1.367 |
| z |
-1.920 |
-1.367 |
-15.806 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.962 |
1.699 |
-1.920 |
| y |
1.699 |
2.109 |
-1.367 |
| z |
-1.920 |
-1.367 |
1.853 |
|
| Polar |
|---|
| 3z2-r2 | 3.705 |
|---|
| x2-y2 | -4.047 |
|---|
| xy | 1.699 |
|---|
| xz | -1.920 |
|---|
| yz | -1.367 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.870 |
0.173 |
-0.046 |
| y |
0.173 |
2.726 |
-0.113 |
| z |
-0.046 |
-0.113 |
2.604 |
<r2> (average value of r
2) Å
2
| <r2> |
42.064 |
| (<r2>)1/2 |
6.486 |