Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -148.799832 |
| Energy at 298.15K | -148.802134 |
| Nuclear repulsion energy | 59.620371 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3588 |
3442 |
42.86 |
|
|
|
| 2 |
A' |
2388 |
2291 |
111.52 |
|
|
|
| 3 |
A' |
1636 |
1569 |
42.23 |
|
|
|
| 4 |
A' |
1112 |
1067 |
8.79 |
|
|
|
| 5 |
A' |
592 |
568 |
163.70 |
|
|
|
| 6 |
A' |
500 |
479 |
70.48 |
|
|
|
| 7 |
A" |
3682 |
3532 |
64.26 |
|
|
|
| 8 |
A" |
1203 |
1153 |
0.10 |
|
|
|
| 9 |
A" |
421 |
404 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7560.9 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 7252.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.219 |
0.000 |
| N2 |
-0.027 |
1.372 |
0.000 |
| N3 |
0.094 |
-1.114 |
0.000 |
| H4 |
-0.234 |
-1.561 |
0.840 |
| H5 |
-0.234 |
-1.561 |
-0.840 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1531 | 1.3363 | 1.9828 | 1.9828 |
N2 | 1.1531 | | 2.4887 | 3.0584 | 3.0584 | N3 | 1.3363 | 2.4887 | | 1.0072 | 1.0072 | H4 | 1.9828 | 3.0584 | 1.0072 | | 1.6807 | H5 | 1.9828 | 3.0584 | 1.0072 | 1.6807 | |
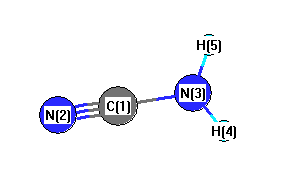 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
114.854 |
|
C1 |
N3 |
H5 |
114.854 |
| N2 |
C1 |
N3 |
177.316 |
|
H4 |
N3 |
H5 |
113.101 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.025 |
|
|
|
| 2 |
N |
-0.128 |
|
|
|
| 3 |
N |
-0.186 |
|
|
|
| 4 |
H |
0.169 |
|
|
|
| 5 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.918 |
-4.511 |
0.000 |
4.603 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.266 |
1.857 |
0.000 |
| y |
1.857 |
-18.344 |
0.000 |
| z |
0.000 |
0.000 |
-14.983 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.602 |
1.857 |
0.000 |
| y |
1.857 |
-1.719 |
0.000 |
| z |
0.000 |
0.000 |
3.321 |
|
| Polar |
|---|
| 3z2-r2 | 6.643 |
|---|
| x2-y2 | 0.078 |
|---|
| xy | 1.857 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.341 |
-0.012 |
0.000 |
| y |
-0.012 |
5.129 |
0.000 |
| z |
0.000 |
0.000 |
2.623 |
<r2> (average value of r
2) Å
2
| <r2> |
39.355 |
| (<r2>)1/2 |
6.273 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -148.798615 |
| Energy at 298.15K | |
| HF Energy | -148.798615 |
| Nuclear repulsion energy | 59.770615 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3656 |
3507 |
67.43 |
|
|
|
| 2 |
A1 |
2387 |
2290 |
140.33 |
|
|
|
| 3 |
A1 |
1619 |
1553 |
50.81 |
|
|
|
| 4 |
A1 |
1145 |
1098 |
12.76 |
|
|
|
| 5 |
B1 |
541 |
519 |
0.44 |
|
|
|
| 6 |
B1 |
418i |
401i |
244.55 |
|
|
|
| 7 |
B2 |
3774 |
3620 |
96.38 |
|
|
|
| 8 |
B2 |
1141 |
1095 |
2.18 |
|
|
|
| 9 |
B2 |
417 |
400 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7130.1 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 6839.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.219 |
| N2 |
0.000 |
0.000 |
1.374 |
| N3 |
0.000 |
0.000 |
-1.102 |
| H4 |
0.000 |
0.864 |
-1.609 |
| H5 |
0.000 |
-0.864 |
-1.609 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1546 | 1.3218 | 2.0216 | 2.0216 |
N2 | 1.1546 | | 2.4764 | 3.1050 | 3.1050 | N3 | 1.3218 | 2.4764 | | 1.0009 | 1.0009 | H4 | 2.0216 | 3.1050 | 1.0009 | | 1.7271 | H5 | 2.0216 | 3.1050 | 1.0009 | 1.7271 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.373 |
|
C1 |
N3 |
H5 |
120.373 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
119.253 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.018 |
|
|
|
| 2 |
N |
-0.142 |
|
|
|
| 3 |
N |
-0.199 |
|
|
|
| 4 |
H |
0.180 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.854 |
4.854 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.502 |
0.000 |
0.000 |
| y |
0.000 |
-14.743 |
0.000 |
| z |
0.000 |
0.000 |
-17.664 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.299 |
0.000 |
0.000 |
| y |
0.000 |
3.340 |
0.000 |
| z |
0.000 |
0.000 |
-1.041 |
|
| Polar |
|---|
| 3z2-r2 | -2.082 |
|---|
| x2-y2 | -3.759 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.282 |
0.000 |
0.000 |
| y |
0.000 |
2.575 |
0.000 |
| z |
0.000 |
0.000 |
5.144 |
<r2> (average value of r
2) Å
2
| <r2> |
39.275 |
| (<r2>)1/2 |
6.267 |
