All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)
using model chemistry: MP4=FULL/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.115751 |
| Energy at 298.15K | |
| HF Energy | -217.203922 |
| Nuclear repulsion energy | 133.178455 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.282 |
0.245 |
0.000 |
| F2 |
-0.875 |
1.034 |
0.000 |
| H3 |
1.123 |
0.936 |
0.000 |
| C4 |
0.282 |
-0.582 |
1.262 |
| C5 |
0.282 |
-0.582 |
-1.262 |
| H6 |
1.187 |
-1.183 |
1.318 |
| H7 |
1.187 |
-1.183 |
-1.318 |
| H8 |
0.234 |
0.060 |
2.137 |
| H9 |
0.234 |
0.060 |
-2.137 |
| H10 |
-0.580 |
-1.246 |
1.267 |
| H11 |
-0.580 |
-1.246 |
-1.267 |
Atom - Atom Distances (Å)
| |
C1 |
F2 |
H3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.3999 | 1.0888 | 1.5086 | 1.5086 | 2.1443 | 2.1443 | 2.1456 | 2.1456 | 2.1379 | 2.1379 |
F2 | 1.3999 | | 2.0004 | 2.3539 | 2.3539 | 3.3024 | 3.3024 | 2.5970 | 2.5970 | 2.6249 | 2.6249 | H3 | 1.0888 | 2.0004 | | 2.1457 | 2.1457 | 2.4967 | 2.4967 | 2.4750 | 2.4750 | 3.0442 | 3.0442 | C4 | 1.5086 | 2.3539 | 2.1457 | | 2.5235 | 1.0884 | 2.7996 | 1.0866 | 3.4593 | 1.0881 | 2.7525 | C5 | 1.5086 | 2.3539 | 2.1457 | 2.5235 | | 2.7996 | 1.0884 | 3.4593 | 1.0866 | 2.7525 | 1.0881 | H6 | 2.1443 | 3.3024 | 2.4967 | 1.0884 | 2.7996 | | 2.6365 | 1.7680 | 3.7940 | 1.7690 | 3.1317 | H7 | 2.1443 | 3.3024 | 2.4967 | 2.7996 | 1.0884 | 2.6365 | | 3.7940 | 1.7680 | 3.1317 | 1.7690 | H8 | 2.1456 | 2.5970 | 2.4750 | 1.0866 | 3.4593 | 1.7680 | 3.7940 | | 4.2742 | 1.7681 | 3.7354 | H9 | 2.1456 | 2.5970 | 2.4750 | 3.4593 | 1.0866 | 3.7940 | 1.7680 | 4.2742 | | 3.7354 | 1.7681 | H10 | 2.1379 | 2.6249 | 3.0442 | 1.0881 | 2.7525 | 1.7690 | 3.1317 | 1.7681 | 3.7354 | | 2.5331 | H11 | 2.1379 | 2.6249 | 3.0442 | 2.7525 | 1.0881 | 3.1317 | 1.7690 | 3.7354 | 1.7681 | 2.5331 | |
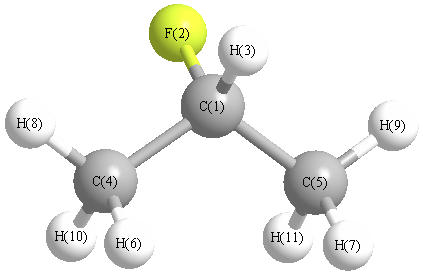 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
H8 |
110.480 |
|
C1 |
C4 |
H10 |
109.773 |
| C1 |
C5 |
H7 |
110.268 |
|
C1 |
C5 |
H9 |
110.480 |
| C1 |
C5 |
H11 |
109.773 |
|
F2 |
C1 |
H3 |
106.314 |
| F2 |
C1 |
C4 |
107.996 |
|
F2 |
C1 |
C5 |
107.996 |
| H3 |
C1 |
C4 |
110.353 |
|
H3 |
C1 |
C5 |
110.353 |
| C4 |
C1 |
C5 |
113.517 |
|
H7 |
C5 |
H9 |
108.756 |
| H7 |
C5 |
H11 |
108.734 |
|
H8 |
C4 |
H10 |
108.788 |
| H9 |
C5 |
H11 |
108.788 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
