Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -377.089075 |
| Energy at 298.15K | |
| HF Energy | -375.936788 |
| Nuclear repulsion energy | 191.941192 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3195 |
3033 |
20.01 |
28.91 |
0.58 |
0.73 |
| 2 |
A |
3166 |
3006 |
19.58 |
85.78 |
0.33 |
0.50 |
| 3 |
A |
3136 |
2977 |
14.28 |
111.37 |
0.07 |
0.13 |
| 4 |
A |
1532 |
1454 |
4.90 |
4.76 |
0.74 |
0.85 |
| 5 |
A |
1495 |
1419 |
16.48 |
2.22 |
0.35 |
0.52 |
| 6 |
A |
1438 |
1365 |
17.37 |
2.89 |
0.73 |
0.85 |
| 7 |
A |
1378 |
1308 |
14.00 |
1.73 |
0.75 |
0.86 |
| 8 |
A |
1301 |
1235 |
11.75 |
4.96 |
0.72 |
0.83 |
| 9 |
A |
1201 |
1140 |
85.84 |
3.96 |
0.72 |
0.84 |
| 10 |
A |
1168 |
1109 |
19.80 |
1.72 |
0.42 |
0.59 |
| 11 |
A |
1152 |
1093 |
226.32 |
0.43 |
0.38 |
0.55 |
| 12 |
A |
1126 |
1069 |
24.56 |
5.18 |
0.46 |
0.63 |
| 13 |
A |
935 |
888 |
32.57 |
3.78 |
0.29 |
0.45 |
| 14 |
A |
590 |
560 |
3.99 |
1.76 |
0.34 |
0.51 |
| 15 |
A |
487 |
463 |
18.10 |
0.63 |
0.72 |
0.84 |
| 16 |
A |
437 |
415 |
5.40 |
1.28 |
0.50 |
0.67 |
| 17 |
A |
246 |
233 |
8.05 |
0.06 |
0.60 |
0.75 |
| 18 |
A |
132 |
125 |
7.90 |
0.03 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12056.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 11446.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.762 |
-0.595 |
-0.276 |
| C2 |
0.465 |
0.021 |
0.328 |
| F3 |
-1.867 |
0.103 |
0.145 |
| F4 |
1.530 |
-0.748 |
-0.011 |
| F5 |
0.660 |
1.257 |
-0.180 |
| H6 |
-0.688 |
-0.542 |
-1.356 |
| H7 |
-0.845 |
-1.626 |
0.046 |
| H8 |
0.413 |
0.099 |
1.407 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.4999 | 1.3736 | 2.3124 | 2.3375 | 1.0838 | 1.0835 | 2.1675 |
C2 | 1.4999 | | 2.3406 | 1.3565 | 1.3514 | 2.1174 | 2.1229 | 1.0831 | F3 | 1.3736 | 2.3406 | | 3.5056 | 2.7971 | 2.0152 | 2.0117 | 2.6064 | F4 | 2.3124 | 1.3565 | 3.5056 | | 2.1923 | 2.6024 | 2.5327 | 1.9939 | F5 | 2.3375 | 1.3514 | 2.7971 | 2.1923 | | 2.5377 | 3.2606 | 1.9802 | H6 | 1.0838 | 2.1174 | 2.0152 | 2.6024 | 2.5377 | | 1.7795 | 3.0433 | H7 | 1.0835 | 2.1229 | 2.0117 | 2.5327 | 3.2606 | 1.7795 | | 2.5324 | H8 | 2.1675 | 1.0831 | 2.6064 | 1.9939 | 1.9802 | 3.0433 | 2.5324 | |
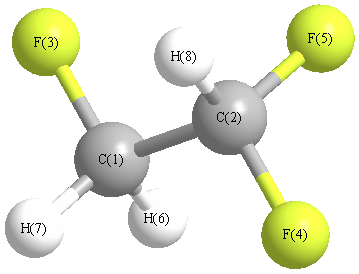 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.999 |
|
C1 |
C2 |
F5 |
110.019 |
| C1 |
C2 |
H8 |
113.110 |
|
C2 |
C1 |
F3 |
109.003 |
| C2 |
C1 |
H6 |
109.010 |
|
C2 |
C1 |
H7 |
109.461 |
| F3 |
C1 |
H6 |
109.614 |
|
F3 |
C1 |
H7 |
109.350 |
| F4 |
C2 |
F5 |
108.117 |
|
F4 |
C2 |
H8 |
109.120 |
| F5 |
C2 |
H8 |
108.360 |
|
H6 |
C1 |
H7 |
110.381 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -377.086968 |
| Energy at 298.15K | |
| HF Energy | -375.934453 |
| Nuclear repulsion energy | 194.683878 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3143 |
2984 |
51.92 |
115.42 |
0.07 |
0.12 |
| 2 |
A' |
3125 |
2967 |
4.34 |
66.04 |
0.25 |
0.40 |
| 3 |
A' |
1527 |
1450 |
7.49 |
4.21 |
0.75 |
0.85 |
| 4 |
A' |
1473 |
1398 |
26.28 |
2.85 |
0.27 |
0.42 |
| 5 |
A' |
1450 |
1376 |
15.91 |
1.62 |
0.70 |
0.82 |
| 6 |
A' |
1224 |
1162 |
111.19 |
3.74 |
0.37 |
0.54 |
| 7 |
A' |
1138 |
1081 |
21.78 |
2.81 |
0.75 |
0.86 |
| 8 |
A' |
904 |
858 |
32.88 |
5.68 |
0.13 |
0.23 |
| 9 |
A' |
778 |
739 |
49.16 |
2.30 |
0.53 |
0.69 |
| 10 |
A' |
528 |
502 |
11.95 |
1.25 |
0.74 |
0.85 |
| 11 |
A' |
240 |
228 |
1.75 |
0.16 |
0.46 |
0.63 |
| 12 |
A" |
3179 |
3018 |
13.26 |
46.01 |
0.75 |
0.86 |
| 13 |
A" |
1432 |
1360 |
24.43 |
1.20 |
0.75 |
0.86 |
| 14 |
A" |
1319 |
1252 |
18.96 |
8.72 |
0.75 |
0.86 |
| 15 |
A" |
1175 |
1116 |
109.86 |
1.21 |
0.75 |
0.86 |
| 16 |
A" |
982 |
932 |
58.36 |
2.25 |
0.75 |
0.86 |
| 17 |
A" |
379 |
360 |
0.21 |
0.21 |
0.75 |
0.86 |
| 18 |
A" |
128 |
122 |
1.89 |
0.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12061.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 11451.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.736 |
-0.825 |
0.000 |
| C2 |
0.355 |
0.625 |
0.000 |
| F3 |
-0.386 |
-1.609 |
0.000 |
| F4 |
-0.386 |
0.916 |
1.094 |
| F5 |
-0.386 |
0.916 |
-1.094 |
| H6 |
1.321 |
-1.036 |
-0.889 |
| H7 |
1.321 |
-1.036 |
0.889 |
| H8 |
1.233 |
1.263 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.4990 | 1.3692 | 2.3418 | 2.3418 | 1.0847 | 1.0847 | 2.1461 |
C2 | 1.4990 | | 2.3538 | 1.3527 | 1.3527 | 2.1171 | 2.1171 | 1.0855 | F3 | 1.3692 | 2.3538 | | 2.7511 | 2.7511 | 2.0080 | 2.0080 | 3.2971 | F4 | 2.3418 | 1.3527 | 2.7511 | | 2.1872 | 3.2639 | 2.6010 | 1.9849 | F5 | 2.3418 | 1.3527 | 2.7511 | 2.1872 | | 2.6010 | 3.2639 | 1.9849 | H6 | 1.0847 | 2.1171 | 2.0080 | 3.2639 | 2.6010 | | 1.7775 | 2.4665 | H7 | 1.0847 | 2.1171 | 2.0080 | 2.6010 | 3.2639 | 1.7775 | | 2.4665 | H8 | 2.1461 | 1.0855 | 3.2971 | 1.9849 | 1.9849 | 2.4665 | 2.4665 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.301 |
|
C1 |
C2 |
F5 |
110.301 |
| C1 |
C2 |
H8 |
111.268 |
|
C2 |
C1 |
F3 |
110.217 |
| C2 |
C1 |
H6 |
108.999 |
|
C2 |
C1 |
H7 |
108.999 |
| F3 |
C1 |
H6 |
109.285 |
|
F3 |
C1 |
H7 |
109.285 |
| F4 |
C2 |
F5 |
107.887 |
|
F4 |
C2 |
H8 |
108.495 |
| F5 |
C2 |
H8 |
108.495 |
|
H6 |
C1 |
H7 |
110.046 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
