Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -377.573589 |
| Energy at 298.15K | |
| HF Energy | -377.573589 |
| Nuclear repulsion energy | 191.719716 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3009 |
21.80 |
37.29 |
0.69 |
0.82 |
| 2 |
A |
3100 |
2967 |
26.95 |
85.92 |
0.32 |
0.49 |
| 3 |
A |
3080 |
2948 |
15.12 |
128.78 |
0.07 |
0.13 |
| 4 |
A |
1496 |
1432 |
4.39 |
5.34 |
0.73 |
0.84 |
| 5 |
A |
1461 |
1398 |
16.16 |
2.28 |
0.40 |
0.57 |
| 6 |
A |
1405 |
1345 |
19.14 |
3.28 |
0.73 |
0.85 |
| 7 |
A |
1345 |
1288 |
14.01 |
1.87 |
0.75 |
0.86 |
| 8 |
A |
1273 |
1218 |
14.91 |
5.23 |
0.72 |
0.84 |
| 9 |
A |
1179 |
1128 |
92.91 |
3.98 |
0.68 |
0.81 |
| 10 |
A |
1143 |
1094 |
122.45 |
1.66 |
0.66 |
0.79 |
| 11 |
A |
1139 |
1090 |
148.28 |
0.75 |
0.43 |
0.60 |
| 12 |
A |
1116 |
1068 |
16.54 |
4.80 |
0.46 |
0.63 |
| 13 |
A |
918 |
879 |
30.47 |
3.93 |
0.32 |
0.49 |
| 14 |
A |
584 |
559 |
3.87 |
1.74 |
0.37 |
0.54 |
| 15 |
A |
473 |
453 |
17.37 |
0.63 |
0.73 |
0.84 |
| 16 |
A |
428 |
410 |
5.44 |
1.37 |
0.50 |
0.67 |
| 17 |
A |
238 |
228 |
7.90 |
0.06 |
0.60 |
0.75 |
| 18 |
A |
114 |
109 |
7.79 |
0.03 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 11817.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 11310.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.773 |
-0.581 |
-0.286 |
| C2 |
0.465 |
0.020 |
0.325 |
| F3 |
-1.872 |
0.110 |
0.153 |
| F4 |
1.521 |
-0.761 |
-0.005 |
| F5 |
0.685 |
1.252 |
-0.181 |
| H6 |
-0.709 |
-0.512 |
-1.370 |
| H7 |
-0.858 |
-1.623 |
0.019 |
| H8 |
0.409 |
0.102 |
1.411 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5057 | 1.3705 | 2.3185 | 2.3446 | 1.0888 | 1.0885 | 2.1773 |
C2 | 1.5057 | | 2.3449 | 1.3548 | 1.3498 | 2.1299 | 2.1314 | 1.0898 | F3 | 1.3705 | 2.3449 | | 3.5071 | 2.8205 | 2.0147 | 2.0119 | 2.6045 | F4 | 2.3185 | 1.3548 | 3.5071 | | 2.1864 | 2.6272 | 2.5311 | 1.9968 | F5 | 2.3446 | 1.3498 | 2.8205 | 2.1864 | | 2.5434 | 3.2688 | 1.9827 | H6 | 1.0888 | 2.1299 | 2.0147 | 2.6272 | 2.5434 | | 1.7852 | 3.0595 | H7 | 1.0885 | 2.1314 | 2.0119 | 2.5311 | 3.2688 | 1.7852 | | 2.5528 | H8 | 2.1773 | 1.0898 | 2.6045 | 1.9968 | 1.9827 | 3.0595 | 2.5528 | |
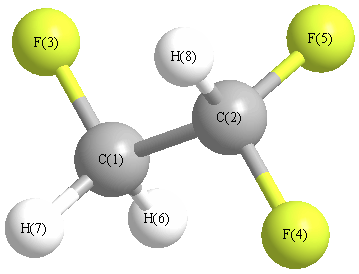 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.180 |
|
C1 |
C2 |
F5 |
110.271 |
| C1 |
C2 |
H8 |
113.076 |
|
C2 |
C1 |
F3 |
109.146 |
| C2 |
C1 |
H6 |
109.311 |
|
C2 |
C1 |
H7 |
109.442 |
| F3 |
C1 |
H6 |
109.486 |
|
F3 |
C1 |
H7 |
109.278 |
| F4 |
C2 |
F5 |
107.874 |
|
F4 |
C2 |
H8 |
109.058 |
| F5 |
C2 |
H8 |
108.252 |
|
H6 |
C1 |
H7 |
110.160 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.064 |
|
|
|
| 2 |
C |
0.303 |
|
|
|
| 3 |
F |
-0.219 |
|
|
|
| 4 |
F |
-0.194 |
|
|
|
| 5 |
F |
-0.186 |
|
|
|
| 6 |
H |
0.088 |
|
|
|
| 7 |
H |
0.082 |
|
|
|
| 8 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.275 |
-1.402 |
0.348 |
1.470 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.911 |
1.536 |
1.393 |
| y |
1.536 |
-27.593 |
0.503 |
| z |
1.393 |
0.503 |
-24.648 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.791 |
1.536 |
1.393 |
| y |
1.536 |
0.687 |
0.503 |
| z |
1.393 |
0.503 |
5.104 |
|
| Polar |
|---|
| 3z2-r2 | 10.208 |
|---|
| x2-y2 | -4.318 |
|---|
| xy | 1.536 |
|---|
| xz | 1.393 |
|---|
| yz | 0.503 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.002 |
-0.023 |
0.016 |
| y |
-0.023 |
3.919 |
0.059 |
| z |
0.016 |
0.059 |
3.764 |
<r2> (average value of r
2) Å
2
| <r2> |
110.264 |
| (<r2>)1/2 |
10.501 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -377.571547 |
| Energy at 298.15K | |
| HF Energy | -377.571547 |
| Nuclear repulsion energy | 194.506900 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3079 |
2947 |
59.01 |
147.92 |
0.02 |
0.04 |
| 2 |
A' |
3065 |
2934 |
3.08 |
50.92 |
0.63 |
0.78 |
| 3 |
A' |
1491 |
1427 |
6.65 |
4.88 |
0.73 |
0.85 |
| 4 |
A' |
1440 |
1378 |
24.28 |
2.64 |
0.29 |
0.45 |
| 5 |
A' |
1418 |
1357 |
19.43 |
2.02 |
0.66 |
0.79 |
| 6 |
A' |
1206 |
1155 |
128.55 |
3.41 |
0.35 |
0.52 |
| 7 |
A' |
1130 |
1082 |
26.90 |
2.96 |
0.74 |
0.85 |
| 8 |
A' |
885 |
847 |
28.31 |
5.88 |
0.17 |
0.29 |
| 9 |
A' |
769 |
736 |
49.37 |
2.37 |
0.54 |
0.70 |
| 10 |
A' |
522 |
499 |
12.18 |
1.25 |
0.75 |
0.86 |
| 11 |
A' |
228 |
218 |
1.72 |
0.15 |
0.45 |
0.62 |
| 12 |
A" |
3128 |
2994 |
17.50 |
52.03 |
0.75 |
0.86 |
| 13 |
A" |
1396 |
1336 |
26.71 |
1.18 |
0.75 |
0.86 |
| 14 |
A" |
1290 |
1235 |
19.98 |
9.56 |
0.75 |
0.86 |
| 15 |
A" |
1157 |
1107 |
116.95 |
1.18 |
0.75 |
0.86 |
| 16 |
A" |
963 |
922 |
56.08 |
2.34 |
0.75 |
0.86 |
| 17 |
A" |
366 |
350 |
0.23 |
0.22 |
0.75 |
0.86 |
| 18 |
A" |
111 |
107 |
1.91 |
0.05 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11821.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 11314.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.733 |
-0.831 |
0.000 |
| C2 |
0.355 |
0.625 |
0.000 |
| F3 |
-0.386 |
-1.614 |
0.000 |
| F4 |
-0.386 |
0.921 |
1.090 |
| F5 |
-0.386 |
0.921 |
-1.090 |
| H6 |
1.322 |
-1.046 |
-0.891 |
| H7 |
1.322 |
-1.046 |
0.891 |
| H8 |
1.241 |
1.264 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5045 | 1.3658 | 2.3475 | 2.3475 | 1.0896 | 1.0896 | 2.1554 |
C2 | 1.5045 | | 2.3589 | 1.3510 | 1.3510 | 2.1266 | 2.1266 | 1.0922 | F3 | 1.3658 | 2.3589 | | 2.7601 | 2.7601 | 2.0083 | 2.0083 | 3.3060 | F4 | 2.3475 | 1.3510 | 2.7601 | | 2.1810 | 3.2731 | 2.6126 | 1.9882 | F5 | 2.3475 | 1.3510 | 2.7601 | 2.1810 | | 2.6126 | 3.2731 | 1.9882 | H6 | 1.0896 | 2.1266 | 2.0083 | 3.2731 | 2.6126 | | 1.7824 | 2.4770 | H7 | 1.0896 | 2.1266 | 2.0083 | 2.6126 | 3.2731 | 1.7824 | | 2.4770 | H8 | 2.1554 | 1.0922 | 3.3060 | 1.9882 | 1.9882 | 2.4770 | 2.4770 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.478 |
|
C1 |
C2 |
F5 |
110.478 |
| C1 |
C2 |
H8 |
111.212 |
|
C2 |
C1 |
F3 |
110.441 |
| C2 |
C1 |
H6 |
109.081 |
|
C2 |
C1 |
H7 |
109.081 |
| F3 |
C1 |
H6 |
109.237 |
|
F3 |
C1 |
H7 |
109.237 |
| F4 |
C2 |
F5 |
107.641 |
|
F4 |
C2 |
H8 |
108.462 |
| F5 |
C2 |
H8 |
108.462 |
|
H6 |
C1 |
H7 |
109.749 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.077 |
|
|
|
| 2 |
C |
0.308 |
|
|
|
| 3 |
F |
-0.210 |
|
|
|
| 4 |
F |
-0.187 |
|
|
|
| 5 |
F |
-0.187 |
|
|
|
| 6 |
H |
0.076 |
|
|
|
| 7 |
H |
0.076 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.040 |
0.057 |
0.000 |
3.040 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.054 |
-0.315 |
0.000 |
| y |
-0.315 |
-28.500 |
0.000 |
| z |
0.000 |
0.000 |
-28.247 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.320 |
-0.315 |
0.000 |
| y |
-0.315 |
-1.850 |
0.000 |
| z |
0.000 |
0.000 |
-1.470 |
|
| Polar |
|---|
| 3z2-r2 | -2.940 |
|---|
| x2-y2 | 3.446 |
|---|
| xy | -0.315 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.902 |
-0.018 |
0.000 |
| y |
-0.018 |
3.949 |
0.000 |
| z |
0.000 |
0.000 |
3.857 |
<r2> (average value of r
2) Å
2
| <r2> |
102.061 |
| (<r2>)1/2 |
10.103 |
