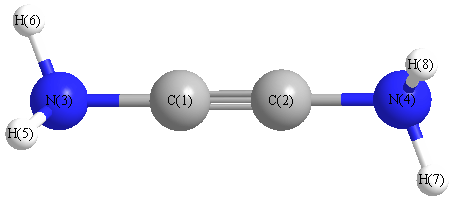
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3656 |
3507 |
23.74 |
|
|
|
| 2 |
A |
3570 |
3425 |
1.77 |
|
|
|
| 3 |
A |
2387 |
2289 |
0.20 |
|
|
|
| 4 |
A |
1655 |
1588 |
7.63 |
|
|
|
| 5 |
A |
1201 |
1152 |
0.00 |
|
|
|
| 6 |
A |
840 |
806 |
1.90 |
|
|
|
| 7 |
A |
527 |
506 |
124.87 |
|
|
|
| 8 |
A |
434 |
416 |
12.78 |
|
|
|
| 9 |
A |
429 |
412 |
17.95 |
|
|
|
| 10 |
A |
203 |
195 |
24.54 |
|
|
|
| 11 |
B |
3656 |
3507 |
22.99 |
|
|
|
| 12 |
B |
3573 |
3428 |
7.05 |
|
|
|
| 13 |
B |
1656 |
1589 |
29.06 |
|
|
|
| 14 |
B |
1398 |
1341 |
124.41 |
|
|
|
| 15 |
B |
1201 |
1152 |
0.02 |
|
|
|
| 16 |
B |
583 |
559 |
292.27 |
|
|
|
| 17 |
B |
437 |
419 |
24.92 |
|
|
|
| 18 |
B |
203 |
195 |
25.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13804.9 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 13241.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.097 |
|
|
|
| 2 |
C |
-0.097 |
|
|
|
| 3 |
N |
-0.205 |
|
|
|
| 4 |
N |
-0.205 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.281 |
1.281 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.566 |
3.716 |
0.000 |
| y |
3.716 |
-13.420 |
0.000 |
| z |
0.000 |
0.000 |
-23.698 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.008 |
3.716 |
0.000 |
| y |
3.716 |
10.213 |
0.000 |
| z |
0.000 |
0.000 |
-5.205 |
|
| Polar |
|---|
| 3z2-r2 | -10.410 |
|---|
| x2-y2 | -10.147 |
|---|
| xy | 3.716 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.795 |
0.211 |
0.000 |
| y |
0.211 |
9.638 |
0.000 |
| z |
0.000 |
0.000 |
3.787 |
<r2> (average value of r
2) Å
2
| <r2> |
96.198 |
| (<r2>)1/2 |
9.808 |