Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.119514 |
| Energy at 298.15K | |
| HF Energy | -153.507228 |
| Nuclear repulsion energy | 74.685353 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.115 |
-0.369 |
0.000 |
| C2 |
0.000 |
0.537 |
0.000 |
| C3 |
1.261 |
-0.256 |
0.000 |
| H4 |
-1.912 |
0.169 |
0.000 |
| H5 |
-0.034 |
1.180 |
0.887 |
| H6 |
-0.034 |
1.180 |
-0.887 |
| H7 |
1.667 |
-0.633 |
-0.928 |
| H8 |
1.667 |
-0.633 |
0.928 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4369 | 2.3786 | 0.9615 | 2.0867 | 2.0867 | 2.9446 | 2.9446 |
C2 | 1.4369 | | 1.4897 | 1.9472 | 1.0955 | 1.0955 | 2.2385 | 2.2385 | C3 | 2.3786 | 1.4897 | | 3.2013 | 2.1268 | 2.1268 | 1.0812 | 1.0812 | H4 | 0.9615 | 1.9472 | 3.2013 | | 2.3102 | 2.3102 | 3.7834 | 3.7834 | H5 | 2.0867 | 1.0955 | 2.1268 | 2.3102 | | 1.7738 | 3.0779 | 2.4860 | H6 | 2.0867 | 1.0955 | 2.1268 | 2.3102 | 1.7738 | | 2.4860 | 3.0779 | H7 | 2.9446 | 2.2385 | 1.0812 | 3.7834 | 3.0779 | 2.4860 | | 1.8565 | H8 | 2.9446 | 2.2385 | 1.0812 | 3.7834 | 2.4860 | 3.0779 | 1.8565 | |
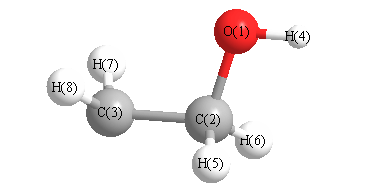 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.716 |
|
O1 |
C2 |
H5 |
110.241 |
| O1 |
C2 |
H6 |
110.241 |
|
C2 |
O1 |
H4 |
106.884 |
| C2 |
C3 |
H7 |
120.247 |
|
C2 |
C3 |
H8 |
120.247 |
| C3 |
C2 |
H5 |
109.764 |
|
C3 |
C2 |
H6 |
109.764 |
| H5 |
C2 |
H6 |
108.107 |
|
H7 |
C3 |
H8 |
118.315 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.099590 |
| Energy at 298.15K | |
| HF Energy | -153.507508 |
| Nuclear repulsion energy | 74.827151 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C1
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
