Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.113232 |
| Energy at 298.15K | |
| HF Energy | -153.507600 |
| Nuclear repulsion energy | 75.152327 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3865 |
3669 |
26.18 |
|
|
|
| 2 |
A' |
3240 |
3076 |
6.07 |
|
|
|
| 3 |
A' |
3070 |
2915 |
52.32 |
|
|
|
| 4 |
A' |
1556 |
1477 |
0.31 |
|
|
|
| 5 |
A' |
1517 |
1441 |
4.27 |
|
|
|
| 6 |
A' |
1457 |
1383 |
6.01 |
|
|
|
| 7 |
A' |
1249 |
1186 |
43.38 |
|
|
|
| 8 |
A' |
1086 |
1031 |
87.43 |
|
|
|
| 9 |
A' |
1021 |
969 |
36.82 |
|
|
|
| 10 |
A' |
633 |
601 |
17.44 |
|
|
|
| 11 |
A' |
386 |
366 |
30.33 |
|
|
|
| 12 |
A" |
3337 |
3168 |
5.10 |
|
|
|
| 13 |
A" |
3101 |
2944 |
38.58 |
|
|
|
| 14 |
A" |
1320 |
1253 |
0.28 |
|
|
|
| 15 |
A" |
1192 |
1132 |
0.79 |
|
|
|
| 16 |
A" |
826 |
784 |
0.24 |
|
|
|
| 17 |
A" |
260 |
246 |
115.40 |
|
|
|
| 18 |
A" |
93i |
89i |
0.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14510.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 13776.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.105 |
-0.372 |
0.000 |
| C2 |
0.000 |
0.537 |
0.000 |
| C3 |
1.252 |
-0.249 |
0.000 |
| H4 |
-1.906 |
0.157 |
0.000 |
| H5 |
-0.035 |
1.175 |
0.881 |
| H6 |
-0.035 |
1.175 |
-0.881 |
| H7 |
1.650 |
-0.629 |
-0.921 |
| H8 |
1.650 |
-0.629 |
0.921 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4304 | 2.3598 | 0.9603 | 2.0767 | 2.0767 | 2.9157 | 2.9157 |
C2 | 1.4304 | | 1.4784 | 1.9439 | 1.0884 | 1.0884 | 2.2201 | 2.2201 | C3 | 2.3598 | 1.4784 | | 3.1844 | 2.1119 | 2.1119 | 1.0727 | 1.0727 | H4 | 0.9603 | 1.9439 | 3.1844 | | 2.3056 | 2.3056 | 3.7566 | 3.7566 | H5 | 2.0767 | 1.0884 | 2.1119 | 2.3056 | | 1.7626 | 3.0559 | 2.4682 | H6 | 2.0767 | 1.0884 | 2.1119 | 2.3056 | 1.7626 | | 2.4682 | 3.0559 | H7 | 2.9157 | 2.2201 | 1.0727 | 3.7566 | 3.0559 | 2.4682 | | 1.8421 | H8 | 2.9157 | 2.2201 | 1.0727 | 3.7566 | 2.4682 | 3.0559 | 1.8421 | |
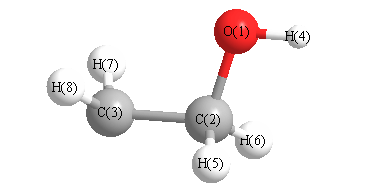 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.432 |
|
O1 |
C2 |
H5 |
110.337 |
| O1 |
C2 |
H6 |
110.337 |
|
C2 |
O1 |
H4 |
107.164 |
| C2 |
C3 |
H7 |
120.145 |
|
C2 |
C3 |
H8 |
120.145 |
| C3 |
C2 |
H5 |
109.798 |
|
C3 |
C2 |
H6 |
109.798 |
| H5 |
C2 |
H6 |
108.136 |
|
H7 |
C3 |
H8 |
118.336 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.114731 |
| Energy at 298.15K | -154.119442 |
| HF Energy | -153.508713 |
| Nuclear repulsion energy | 75.474695 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3888 |
3692 |
36.73 |
|
|
|
| 2 |
A |
3349 |
3179 |
2.05 |
|
|
|
| 3 |
A |
3246 |
3082 |
4.13 |
|
|
|
| 4 |
A |
3048 |
2894 |
45.23 |
|
|
|
| 5 |
A |
2999 |
2848 |
50.25 |
|
|
|
| 6 |
A |
1533 |
1455 |
4.32 |
|
|
|
| 7 |
A |
1503 |
1427 |
6.00 |
|
|
|
| 8 |
A |
1453 |
1379 |
1.69 |
|
|
|
| 9 |
A |
1289 |
1224 |
44.23 |
|
|
|
| 10 |
A |
1251 |
1188 |
14.46 |
|
|
|
| 11 |
A |
1148 |
1090 |
47.78 |
|
|
|
| 12 |
A |
1094 |
1038 |
27.98 |
|
|
|
| 13 |
A |
990 |
940 |
9.84 |
|
|
|
| 14 |
A |
891 |
846 |
16.68 |
|
|
|
| 15 |
A |
475 |
451 |
38.16 |
|
|
|
| 16 |
A |
422 |
400 |
21.29 |
|
|
|
| 17 |
A |
271 |
257 |
89.86 |
|
|
|
| 18 |
A |
149 |
142 |
23.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14498.9 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 13765.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.101 |
-0.378 |
-0.049 |
| C2 |
-0.009 |
0.522 |
0.032 |
| C3 |
1.241 |
-0.256 |
-0.024 |
| H4 |
-1.905 |
0.128 |
0.082 |
| H5 |
-0.051 |
1.095 |
0.966 |
| H6 |
-0.031 |
1.249 |
-0.782 |
| H7 |
2.178 |
0.241 |
-0.179 |
| H8 |
1.225 |
-1.290 |
0.259 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4174 | 2.3450 | 0.9588 | 2.0735 | 2.0805 | 3.3398 | 2.5175 |
C2 | 1.4174 | | 1.4733 | 1.9377 | 1.0955 | 1.0918 | 2.2156 | 2.2041 | C3 | 2.3450 | 1.4733 | | 3.1712 | 2.1148 | 2.1109 | 1.0724 | 1.0722 | H4 | 0.9588 | 1.9377 | 3.1712 | | 2.2703 | 2.3492 | 4.0938 | 3.4410 | H5 | 2.0735 | 1.0955 | 2.1148 | 2.2703 | | 1.7547 | 2.6479 | 2.7952 | H6 | 2.0805 | 1.0918 | 2.1109 | 2.3492 | 1.7547 | | 2.5020 | 3.0178 | H7 | 3.3398 | 2.2156 | 1.0724 | 4.0938 | 2.6479 | 2.5020 | | 1.8558 | H8 | 2.5175 | 2.2041 | 1.0722 | 3.4410 | 2.7952 | 3.0178 | 1.8558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.418 |
|
O1 |
C2 |
H5 |
110.552 |
| O1 |
C2 |
H6 |
111.358 |
|
C2 |
O1 |
H4 |
107.698 |
| C2 |
C3 |
H7 |
120.175 |
|
C2 |
C3 |
H8 |
119.130 |
| C3 |
C2 |
H5 |
109.956 |
|
C3 |
C2 |
H6 |
109.862 |
| H5 |
C2 |
H6 |
106.690 |
|
H7 |
C3 |
H8 |
119.846 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
