Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.344814 |
| Energy at 298.15K | |
| HF Energy | -154.344814 |
| Nuclear repulsion energy | 75.067798 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3858 |
3693 |
17.68 |
|
|
|
| 2 |
A' |
3167 |
3031 |
9.58 |
|
|
|
| 3 |
A' |
3008 |
2879 |
60.94 |
|
|
|
| 4 |
A' |
1522 |
1457 |
0.23 |
|
|
|
| 5 |
A' |
1478 |
1414 |
5.10 |
|
|
|
| 6 |
A' |
1430 |
1369 |
6.24 |
|
|
|
| 7 |
A' |
1230 |
1177 |
45.23 |
|
|
|
| 8 |
A' |
1061 |
1015 |
87.22 |
|
|
|
| 9 |
A' |
1007 |
964 |
41.49 |
|
|
|
| 10 |
A' |
614 |
588 |
16.30 |
|
|
|
| 11 |
A' |
375 |
359 |
28.54 |
|
|
|
| 12 |
A" |
3275 |
3135 |
10.49 |
|
|
|
| 13 |
A" |
3042 |
2912 |
48.18 |
|
|
|
| 14 |
A" |
1287 |
1232 |
0.37 |
|
|
|
| 15 |
A" |
1169 |
1119 |
0.75 |
|
|
|
| 16 |
A" |
807 |
773 |
0.14 |
|
|
|
| 17 |
A" |
226 |
216 |
112.88 |
|
|
|
| 18 |
A" |
162i |
155i |
2.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14197.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 13588.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.109 |
-0.368 |
0.000 |
| C2 |
0.000 |
0.531 |
0.000 |
| C3 |
1.253 |
-0.251 |
0.000 |
| H4 |
-1.911 |
0.156 |
0.000 |
| H5 |
-0.036 |
1.174 |
0.884 |
| H6 |
-0.036 |
1.174 |
-0.884 |
| H7 |
1.670 |
-0.618 |
-0.925 |
| H8 |
1.670 |
-0.618 |
0.925 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4280 | 2.3655 | 0.9583 | 2.0763 | 2.0763 | 2.9394 | 2.9394 |
C2 | 1.4280 | | 1.4769 | 1.9477 | 1.0938 | 1.0938 | 2.2275 | 2.2275 | C3 | 2.3655 | 1.4769 | | 3.1906 | 2.1152 | 2.1152 | 1.0787 | 1.0787 | H4 | 0.9583 | 1.9477 | 3.1906 | | 2.3092 | 2.3092 | 3.7785 | 3.7785 | H5 | 2.0763 | 1.0938 | 2.1152 | 2.3092 | | 1.7682 | 3.0647 | 2.4741 | H6 | 2.0763 | 1.0938 | 2.1152 | 2.3092 | 1.7682 | | 2.4741 | 3.0647 | H7 | 2.9394 | 2.2275 | 1.0787 | 3.7785 | 3.0647 | 2.4741 | | 1.8500 | H8 | 2.9394 | 2.2275 | 1.0787 | 3.7785 | 2.4741 | 3.0647 | 1.8500 | |
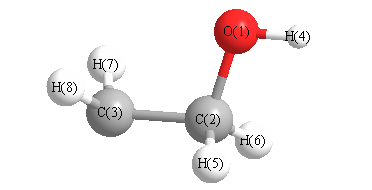 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.027 |
|
O1 |
C2 |
H5 |
110.140 |
| O1 |
C2 |
H6 |
110.140 |
|
C2 |
O1 |
H4 |
107.787 |
| C2 |
C3 |
H7 |
120.492 |
|
C2 |
C3 |
H8 |
120.492 |
| C3 |
C2 |
H5 |
109.836 |
|
C3 |
C2 |
H6 |
109.836 |
| H5 |
C2 |
H6 |
107.851 |
|
H7 |
C3 |
H8 |
118.089 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.342 |
|
|
|
| 2 |
C |
0.008 |
|
|
|
| 3 |
C |
-0.285 |
|
|
|
| 4 |
H |
0.207 |
|
|
|
| 5 |
H |
0.072 |
|
|
|
| 6 |
H |
0.072 |
|
|
|
| 7 |
H |
0.134 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.180 |
1.467 |
0.000 |
1.478 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.846 |
-2.705 |
0.000 |
| y |
-2.705 |
-20.083 |
0.000 |
| z |
0.000 |
0.000 |
-18.785 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.588 |
-2.705 |
0.000 |
| y |
-2.705 |
-2.267 |
0.000 |
| z |
0.000 |
0.000 |
-0.321 |
|
| Polar |
|---|
| 3z2-r2 | -0.642 |
|---|
| x2-y2 | 3.237 |
|---|
| xy | -2.705 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.004 |
-0.398 |
0.000 |
| y |
-0.398 |
3.916 |
0.000 |
| z |
0.000 |
0.000 |
3.894 |
<r2> (average value of r
2) Å
2
| <r2> |
50.069 |
| (<r2>)1/2 |
7.076 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -154.347062 |
| Energy at 298.15K | -154.351612 |
| HF Energy | -154.347062 |
| Nuclear repulsion energy | 75.485239 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3894 |
3727 |
32.18 |
|
|
|
| 2 |
A |
3297 |
3155 |
4.68 |
|
|
|
| 3 |
A |
3182 |
3045 |
5.34 |
|
|
|
| 4 |
A |
2956 |
2829 |
56.65 |
|
|
|
| 5 |
A |
2910 |
2785 |
59.31 |
|
|
|
| 6 |
A |
1493 |
1429 |
9.30 |
|
|
|
| 7 |
A |
1463 |
1401 |
3.74 |
|
|
|
| 8 |
A |
1419 |
1358 |
0.63 |
|
|
|
| 9 |
A |
1269 |
1215 |
64.70 |
|
|
|
| 10 |
A |
1222 |
1170 |
5.39 |
|
|
|
| 11 |
A |
1144 |
1095 |
60.54 |
|
|
|
| 12 |
A |
1066 |
1020 |
17.04 |
|
|
|
| 13 |
A |
957 |
916 |
9.94 |
|
|
|
| 14 |
A |
880 |
842 |
17.13 |
|
|
|
| 15 |
A |
443 |
424 |
24.73 |
|
|
|
| 16 |
A |
406 |
388 |
39.06 |
|
|
|
| 17 |
A |
274 |
262 |
83.59 |
|
|
|
| 18 |
A |
90 |
86 |
25.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14180.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 13572.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.101 |
-0.376 |
-0.057 |
| C2 |
-0.008 |
0.518 |
0.034 |
| C3 |
1.241 |
-0.253 |
-0.016 |
| H4 |
-1.901 |
0.122 |
0.108 |
| H5 |
-0.064 |
1.091 |
0.973 |
| H6 |
-0.030 |
1.255 |
-0.779 |
| H7 |
2.177 |
0.242 |
-0.218 |
| H8 |
1.234 |
-1.292 |
0.268 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4152 | 2.3456 | 0.9568 | 2.0713 | 2.0806 | 3.3395 | 2.5298 |
C2 | 1.4152 | | 1.4685 | 1.9351 | 1.1016 | 1.0969 | 2.2166 | 2.2081 | C3 | 2.3456 | 1.4685 | | 3.1665 | 2.1177 | 2.1140 | 1.0776 | 1.0774 | H4 | 0.9568 | 1.9351 | 3.1665 | | 2.2497 | 2.3602 | 4.0924 | 3.4434 | H5 | 2.0713 | 1.1016 | 2.1177 | 2.2497 | | 1.7598 | 2.6756 | 2.8035 | H6 | 2.0806 | 1.0969 | 2.1140 | 2.3602 | 1.7598 | | 2.4919 | 3.0300 | H7 | 3.3395 | 2.2166 | 1.0776 | 4.0924 | 2.6756 | 2.4919 | | 1.8644 | H8 | 2.5298 | 2.2081 | 1.0774 | 3.4434 | 2.8035 | 3.0300 | 1.8644 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.846 |
|
O1 |
C2 |
H5 |
110.145 |
| O1 |
C2 |
H6 |
111.205 |
|
C2 |
O1 |
H4 |
107.768 |
| C2 |
C3 |
H7 |
120.279 |
|
C2 |
C3 |
H8 |
119.502 |
| C3 |
C2 |
H5 |
110.149 |
|
C3 |
C2 |
H6 |
110.139 |
| H5 |
C2 |
H6 |
106.343 |
|
H7 |
C3 |
H8 |
119.804 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.337 |
|
|
|
| 2 |
C |
0.028 |
|
|
|
| 3 |
C |
-0.316 |
|
|
|
| 4 |
H |
0.207 |
|
|
|
| 5 |
H |
0.071 |
|
|
|
| 6 |
H |
0.073 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.347 |
1.636 |
0.317 |
1.703 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.159 |
-1.728 |
-0.445 |
| y |
-1.728 |
-19.138 |
-0.291 |
| z |
-0.445 |
-0.291 |
-20.347 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.584 |
-1.728 |
-0.445 |
| y |
-1.728 |
-0.885 |
-0.291 |
| z |
-0.445 |
-0.291 |
-2.699 |
|
| Polar |
|---|
| 3z2-r2 | -5.397 |
|---|
| x2-y2 | 2.979 |
|---|
| xy | -1.728 |
|---|
| xz | -0.445 |
|---|
| yz | -0.291 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.019 |
-0.258 |
-0.094 |
| y |
-0.258 |
4.340 |
-0.033 |
| z |
-0.094 |
-0.033 |
3.464 |
<r2> (average value of r
2) Å
2
| <r2> |
49.825 |
| (<r2>)1/2 |
7.059 |
