Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.039172 |
| Energy at 298.15K | -218.047083 |
| HF Energy | -217.197601 |
| Nuclear repulsion energy | 128.951648 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3177 |
3016 |
19.49 |
|
|
|
| 2 |
A' |
3113 |
2956 |
34.48 |
|
|
|
| 3 |
A' |
3098 |
2941 |
26.60 |
|
|
|
| 4 |
A' |
3094 |
2937 |
10.39 |
|
|
|
| 5 |
A' |
1552 |
1474 |
3.37 |
|
|
|
| 6 |
A' |
1535 |
1458 |
5.19 |
|
|
|
| 7 |
A' |
1520 |
1443 |
0.37 |
|
|
|
| 8 |
A' |
1456 |
1382 |
14.52 |
|
|
|
| 9 |
A' |
1428 |
1355 |
2.32 |
|
|
|
| 10 |
A' |
1351 |
1282 |
0.32 |
|
|
|
| 11 |
A' |
1164 |
1105 |
1.41 |
|
|
|
| 12 |
A' |
1107 |
1051 |
88.49 |
|
|
|
| 13 |
A' |
1072 |
1018 |
18.24 |
|
|
|
| 14 |
A' |
922 |
875 |
8.00 |
|
|
|
| 15 |
A' |
460 |
437 |
6.09 |
|
|
|
| 16 |
A' |
268 |
254 |
3.20 |
|
|
|
| 17 |
A" |
3169 |
3009 |
56.38 |
|
|
|
| 18 |
A" |
3150 |
2991 |
3.40 |
|
|
|
| 19 |
A" |
3131 |
2973 |
6.49 |
|
|
|
| 20 |
A" |
1530 |
1453 |
7.35 |
|
|
|
| 21 |
A" |
1345 |
1277 |
0.01 |
|
|
|
| 22 |
A" |
1297 |
1231 |
0.62 |
|
|
|
| 23 |
A" |
1212 |
1151 |
1.39 |
|
|
|
| 24 |
A" |
911 |
865 |
1.43 |
|
|
|
| 25 |
A" |
781 |
741 |
1.68 |
|
|
|
| 26 |
A" |
242 |
230 |
0.01 |
|
|
|
| 27 |
A" |
138 |
131 |
3.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21610.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20516.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.127 |
-0.769 |
0.000 |
| C2 |
0.000 |
0.728 |
0.000 |
| C3 |
-1.457 |
1.150 |
0.000 |
| F4 |
1.460 |
-1.141 |
0.000 |
| H5 |
-0.339 |
-1.197 |
0.883 |
| H6 |
-0.339 |
-1.197 |
-0.883 |
| H7 |
0.509 |
1.121 |
-0.875 |
| H8 |
0.509 |
1.121 |
0.875 |
| H9 |
-1.552 |
2.229 |
0.000 |
| H10 |
-1.973 |
0.771 |
-0.878 |
| H11 |
-1.973 |
0.771 |
0.878 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5024 | 2.4880 | 1.3847 | 1.0866 | 1.0866 | 2.1175 | 2.1175 | 3.4361 | 2.7479 | 2.7479 |
C2 | 1.5024 | | 1.5172 | 2.3725 | 2.1453 | 2.1453 | 1.0857 | 1.0857 | 2.1591 | 2.1601 | 2.1601 | C3 | 2.4880 | 1.5172 | | 3.7098 | 2.7455 | 2.7455 | 2.1526 | 2.1526 | 1.0839 | 1.0861 | 1.0861 | F4 | 1.3847 | 2.3725 | 3.7098 | | 2.0053 | 2.0053 | 2.6058 | 2.6058 | 4.5207 | 4.0271 | 4.0271 | H5 | 1.0866 | 2.1453 | 2.7455 | 2.0053 | | 1.7665 | 3.0307 | 2.4688 | 3.7406 | 3.1055 | 2.5581 | H6 | 1.0866 | 2.1453 | 2.7455 | 2.0053 | 1.7665 | | 2.4688 | 3.0307 | 3.7406 | 2.5581 | 3.1055 | H7 | 2.1175 | 1.0857 | 2.1526 | 2.6058 | 3.0307 | 2.4688 | | 1.7492 | 2.4983 | 2.5073 | 3.0588 | H8 | 2.1175 | 1.0857 | 2.1526 | 2.6058 | 2.4688 | 3.0307 | 1.7492 | | 2.4983 | 3.0588 | 2.5073 | H9 | 3.4361 | 2.1591 | 1.0839 | 4.5207 | 3.7406 | 3.7406 | 2.4983 | 2.4983 | | 1.7535 | 1.7535 | H10 | 2.7479 | 2.1601 | 1.0861 | 4.0271 | 3.1055 | 2.5581 | 2.5073 | 3.0588 | 1.7535 | | 1.7551 | H11 | 2.7479 | 2.1601 | 1.0861 | 4.0271 | 2.5581 | 3.1055 | 3.0588 | 2.5073 | 1.7535 | 1.7551 | |
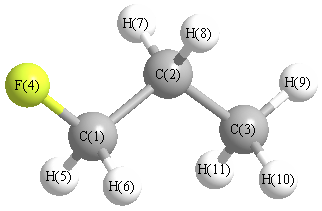 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
110.960 |
|
C1 |
C2 |
H7 |
108.734 |
| C1 |
C2 |
H8 |
108.734 |
|
C2 |
C1 |
F4 |
110.452 |
| C2 |
C1 |
H5 |
110.894 |
|
C2 |
C1 |
H6 |
110.894 |
| C2 |
C3 |
H9 |
111.125 |
|
C2 |
C3 |
H10 |
111.070 |
| C2 |
C3 |
H11 |
111.070 |
|
C3 |
C2 |
H7 |
110.490 |
| C3 |
C2 |
H8 |
110.490 |
|
F4 |
C1 |
H5 |
107.868 |
| F4 |
C1 |
H6 |
107.868 |
|
H5 |
C1 |
H6 |
108.757 |
| H7 |
C2 |
H8 |
107.333 |
|
H9 |
C3 |
H10 |
107.815 |
| H9 |
C3 |
H11 |
107.815 |
|
H10 |
C3 |
H11 |
107.790 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.039691 |
| Energy at 298.15K | |
| HF Energy | -217.197691 |
| Nuclear repulsion energy | 131.371554 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3023 |
15.06 |
|
|
|
| 2 |
A |
3170 |
3009 |
33.40 |
|
|
|
| 3 |
A |
3153 |
2994 |
36.15 |
|
|
|
| 4 |
A |
3135 |
2976 |
13.80 |
|
|
|
| 5 |
A |
3103 |
2946 |
32.28 |
|
|
|
| 6 |
A |
3099 |
2942 |
19.88 |
|
|
|
| 7 |
A |
3098 |
2941 |
10.84 |
|
|
|
| 8 |
A |
1548 |
1470 |
1.71 |
|
|
|
| 9 |
A |
1537 |
1459 |
7.90 |
|
|
|
| 10 |
A |
1526 |
1448 |
6.71 |
|
|
|
| 11 |
A |
1499 |
1424 |
2.97 |
|
|
|
| 12 |
A |
1452 |
1378 |
10.36 |
|
|
|
| 13 |
A |
1428 |
1356 |
4.29 |
|
|
|
| 14 |
A |
1397 |
1326 |
0.05 |
|
|
|
| 15 |
A |
1328 |
1261 |
1.48 |
|
|
|
| 16 |
A |
1303 |
1237 |
1.31 |
|
|
|
| 17 |
A |
1198 |
1138 |
1.61 |
|
|
|
| 18 |
A |
1147 |
1089 |
13.83 |
|
|
|
| 19 |
A |
1125 |
1068 |
44.63 |
|
|
|
| 20 |
A |
1005 |
954 |
40.69 |
|
|
|
| 21 |
A |
939 |
891 |
2.64 |
|
|
|
| 22 |
A |
905 |
859 |
4.15 |
|
|
|
| 23 |
A |
783 |
743 |
0.49 |
|
|
|
| 24 |
A |
488 |
463 |
3.83 |
|
|
|
| 25 |
A |
326 |
310 |
0.95 |
|
|
|
| 26 |
A |
233 |
221 |
1.80 |
|
|
|
| 27 |
A |
153 |
146 |
2.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21630.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20535.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.786 |
0.511 |
0.300 |
| C2 |
-0.591 |
0.653 |
-0.286 |
| C3 |
-1.510 |
-0.483 |
0.118 |
| F4 |
1.382 |
-0.651 |
-0.172 |
| H5 |
1.426 |
1.342 |
0.024 |
| H6 |
0.742 |
0.438 |
1.383 |
| H7 |
-0.502 |
0.702 |
-1.368 |
| H8 |
-0.996 |
1.607 |
0.045 |
| H9 |
-2.492 |
-0.372 |
-0.326 |
| H10 |
-1.098 |
-1.434 |
-0.199 |
| H11 |
-1.631 |
-0.513 |
1.197 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5028 | 2.5080 | 1.3879 | 1.0850 | 1.0867 | 2.1155 | 2.1077 | 3.4524 | 2.7535 | 2.7741 |
C2 | 1.5028 | | 1.5162 | 2.3674 | 2.1532 | 2.1469 | 1.0863 | 1.0878 | 2.1609 | 2.1500 | 2.1551 | C3 | 2.5080 | 1.5162 | | 2.9112 | 3.4579 | 2.7418 | 2.1515 | 2.1529 | 1.0843 | 1.0840 | 1.0864 | F4 | 1.3879 | 2.3674 | 2.9112 | | 2.0030 | 2.0029 | 2.6095 | 3.2861 | 3.8877 | 2.6014 | 3.3125 | H5 | 1.0850 | 2.1532 | 3.4579 | 2.0030 | | 1.7703 | 2.4618 | 2.4363 | 4.2911 | 3.7589 | 3.7635 | H6 | 1.0867 | 2.1469 | 2.7418 | 2.0029 | 1.7703 | | 3.0306 | 2.4858 | 3.7470 | 3.0649 | 2.5632 | H7 | 2.1155 | 1.0863 | 2.1515 | 2.6095 | 2.4618 | 3.0306 | | 1.7487 | 2.4903 | 2.5073 | 3.0550 | H8 | 2.1077 | 1.0878 | 2.1529 | 3.2861 | 2.4363 | 2.4858 | 1.7487 | | 2.5083 | 3.0523 | 2.4951 | H9 | 3.4524 | 2.1609 | 1.0843 | 3.8877 | 4.2911 | 3.7470 | 2.4903 | 2.5083 | | 1.7573 | 1.7562 | H10 | 2.7535 | 2.1500 | 1.0840 | 2.6014 | 3.7589 | 3.0649 | 2.5073 | 3.0523 | 1.7573 | | 1.7555 | H11 | 2.7741 | 2.1551 | 1.0864 | 3.3125 | 3.7635 | 2.5632 | 3.0550 | 2.4951 | 1.7562 | 1.7555 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.344 |
|
C1 |
C2 |
H7 |
108.513 |
| C1 |
C2 |
H8 |
107.818 |
|
C2 |
C1 |
F4 |
109.896 |
| C2 |
C1 |
H5 |
111.602 |
|
C2 |
C1 |
H6 |
110.985 |
| C2 |
C3 |
H9 |
111.305 |
|
C2 |
C3 |
H10 |
110.449 |
| C2 |
C3 |
H11 |
110.712 |
|
C3 |
C2 |
H7 |
110.434 |
| C3 |
C2 |
H8 |
110.457 |
|
F4 |
C1 |
H5 |
107.557 |
| F4 |
C1 |
H6 |
107.447 |
|
H5 |
C1 |
H6 |
109.207 |
| H7 |
C2 |
H8 |
107.092 |
|
H9 |
C3 |
H10 |
108.280 |
| H9 |
C3 |
H11 |
108.010 |
|
H10 |
C3 |
H11 |
107.970 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
