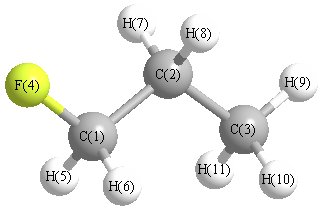
All results from a given calculation for CH2FCH2CH3 (1-Fluoropropane)
using model chemistry: MP4=FULL/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS trans |
1A' |
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.108161 |
| Energy at 298.15K | |
| HF Energy | -217.197471 |
| Nuclear repulsion energy | 128.544774 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.126 |
-0.773 |
0.000 |
| C2 |
0.000 |
0.729 |
0.000 |
| C3 |
-1.462 |
1.156 |
0.000 |
| F4 |
1.465 |
-1.145 |
0.000 |
| H5 |
-0.339 |
-1.203 |
0.886 |
| H6 |
-0.339 |
-1.203 |
-0.886 |
| H7 |
0.511 |
1.123 |
-0.877 |
| H8 |
0.511 |
1.123 |
0.877 |
| H9 |
-1.552 |
2.239 |
0.000 |
| H10 |
-1.980 |
0.777 |
-0.880 |
| H11 |
-1.980 |
0.777 |
0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5074 | 2.4981 | 1.3894 | 1.0892 | 1.0892 | 2.1241 | 2.1241 | 3.4476 | 2.7590 | 2.7590 |
C2 | 1.5074 | | 1.5226 | 2.3782 | 2.1528 | 2.1528 | 1.0885 | 1.0885 | 2.1650 | 2.1671 | 2.1671 | C3 | 2.4981 | 1.5226 | | 3.7222 | 2.7587 | 2.7587 | 2.1586 | 2.1586 | 1.0867 | 1.0892 | 1.0892 | F4 | 1.3894 | 2.3782 | 3.7222 | | 2.0105 | 2.0105 | 2.6117 | 2.6117 | 4.5328 | 4.0414 | 4.0414 | H5 | 1.0892 | 2.1528 | 2.7587 | 2.0105 | | 1.7720 | 3.0400 | 2.4766 | 3.7555 | 3.1200 | 2.5720 | H6 | 1.0892 | 2.1528 | 2.7587 | 2.0105 | 1.7720 | | 2.4766 | 3.0400 | 3.7555 | 2.5720 | 3.1200 | H7 | 2.1241 | 1.0885 | 2.1586 | 2.6117 | 3.0400 | 2.4766 | | 1.7540 | 2.5034 | 2.5141 | 3.0673 | H8 | 2.1241 | 1.0885 | 2.1586 | 2.6117 | 2.4766 | 3.0400 | 1.7540 | | 2.5034 | 3.0673 | 2.5141 | H9 | 3.4476 | 2.1650 | 1.0867 | 4.5328 | 3.7555 | 3.7555 | 2.5034 | 2.5034 | | 1.7588 | 1.7588 | H10 | 2.7590 | 2.1671 | 1.0892 | 4.0414 | 3.1200 | 2.5720 | 2.5141 | 3.0673 | 1.7588 | | 1.7603 | H11 | 2.7590 | 2.1671 | 1.0892 | 4.0414 | 2.5720 | 3.1200 | 3.0673 | 2.5141 | 1.7588 | 1.7603 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.065 |
|
C1 |
C2 |
H7 |
108.745 |
| C1 |
C2 |
H8 |
108.745 |
|
C2 |
C1 |
F4 |
110.301 |
| C2 |
C1 |
H5 |
110.980 |
|
C2 |
C1 |
H6 |
110.980 |
| C2 |
C3 |
H9 |
111.032 |
|
C2 |
C3 |
H10 |
111.056 |
| C2 |
C3 |
H11 |
111.056 |
|
C3 |
C2 |
H7 |
110.416 |
| C3 |
C2 |
H8 |
110.416 |
|
F4 |
C1 |
H5 |
107.802 |
| F4 |
C1 |
H6 |
107.802 |
|
H5 |
C1 |
H6 |
108.862 |
| H7 |
C2 |
H8 |
107.352 |
|
H9 |
C3 |
H10 |
107.863 |
| H9 |
C3 |
H11 |
107.863 |
|
H10 |
C3 |
H11 |
107.824 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
