Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.355205 |
| Energy at 298.15K | -218.363023 |
| HF Energy | -218.355205 |
| Nuclear repulsion energy | 128.729771 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3138 |
3004 |
25.98 |
|
|
|
| 2 |
A' |
3067 |
2936 |
40.53 |
|
|
|
| 3 |
A' |
3056 |
2925 |
21.29 |
|
|
|
| 4 |
A' |
3046 |
2915 |
24.54 |
|
|
|
| 5 |
A' |
1522 |
1456 |
4.32 |
|
|
|
| 6 |
A' |
1508 |
1443 |
4.04 |
|
|
|
| 7 |
A' |
1493 |
1429 |
0.49 |
|
|
|
| 8 |
A' |
1429 |
1367 |
14.41 |
|
|
|
| 9 |
A' |
1413 |
1353 |
1.26 |
|
|
|
| 10 |
A' |
1332 |
1275 |
0.59 |
|
|
|
| 11 |
A' |
1146 |
1097 |
2.16 |
|
|
|
| 12 |
A' |
1090 |
1043 |
108.36 |
|
|
|
| 13 |
A' |
1055 |
1009 |
8.83 |
|
|
|
| 14 |
A' |
908 |
869 |
8.21 |
|
|
|
| 15 |
A' |
451 |
432 |
6.25 |
|
|
|
| 16 |
A' |
259 |
248 |
3.05 |
|
|
|
| 17 |
A" |
3129 |
2995 |
69.02 |
|
|
|
| 18 |
A" |
3107 |
2974 |
5.86 |
|
|
|
| 19 |
A" |
3084 |
2952 |
10.94 |
|
|
|
| 20 |
A" |
1501 |
1437 |
7.42 |
|
|
|
| 21 |
A" |
1318 |
1262 |
0.04 |
|
|
|
| 22 |
A" |
1272 |
1217 |
0.59 |
|
|
|
| 23 |
A" |
1193 |
1142 |
1.31 |
|
|
|
| 24 |
A" |
893 |
854 |
1.43 |
|
|
|
| 25 |
A" |
762 |
729 |
2.24 |
|
|
|
| 26 |
A" |
228 |
218 |
0.00 |
|
|
|
| 27 |
A" |
125 |
120 |
2.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21261.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20349.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.126 |
-0.775 |
0.000 |
| C2 |
0.000 |
0.726 |
0.000 |
| C3 |
-1.454 |
1.162 |
0.000 |
| F4 |
1.457 |
-1.147 |
0.000 |
| H5 |
-0.340 |
-1.209 |
0.885 |
| H6 |
-0.340 |
-1.209 |
-0.885 |
| H7 |
0.515 |
1.119 |
-0.876 |
| H8 |
0.515 |
1.119 |
0.876 |
| H9 |
-1.537 |
2.246 |
0.000 |
| H10 |
-1.980 |
0.791 |
-0.879 |
| H11 |
-1.980 |
0.791 |
0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5055 | 2.4994 | 1.3824 | 1.0911 | 1.0911 | 2.1225 | 2.1225 | 3.4481 | 2.7672 | 2.7672 |
C2 | 1.5055 | | 1.5183 | 2.3731 | 2.1551 | 2.1551 | 1.0896 | 1.0896 | 2.1621 | 2.1670 | 2.1670 | C3 | 2.4994 | 1.5183 | | 3.7161 | 2.7656 | 2.7656 | 2.1559 | 2.1559 | 1.0871 | 1.0892 | 1.0892 | F4 | 1.3824 | 2.3731 | 3.7161 | | 2.0045 | 2.0045 | 2.6061 | 2.6061 | 4.5257 | 4.0425 | 4.0425 | H5 | 1.0911 | 2.1551 | 2.7656 | 2.0045 | | 1.7710 | 3.0425 | 2.4810 | 3.7625 | 3.1311 | 2.5865 | H6 | 1.0911 | 2.1551 | 2.7656 | 2.0045 | 1.7710 | | 2.4810 | 3.0425 | 3.7625 | 2.5865 | 3.1311 | H7 | 2.1225 | 1.0896 | 2.1559 | 2.6061 | 3.0425 | 2.4810 | | 1.7511 | 2.4999 | 2.5164 | 3.0678 | H8 | 2.1225 | 1.0896 | 2.1559 | 2.6061 | 2.4810 | 3.0425 | 1.7511 | | 2.4999 | 3.0678 | 2.5164 | H9 | 3.4481 | 2.1621 | 1.0871 | 4.5257 | 3.7625 | 3.7625 | 2.4999 | 2.4999 | | 1.7564 | 1.7564 | H10 | 2.7672 | 2.1670 | 1.0892 | 4.0425 | 3.1311 | 2.5865 | 2.5164 | 3.0678 | 1.7564 | | 1.7584 | H11 | 2.7672 | 2.1670 | 1.0892 | 4.0425 | 2.5865 | 3.1311 | 3.0678 | 2.5164 | 1.7564 | 1.7584 | |
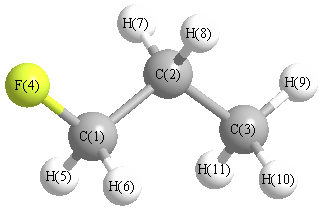 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.500 |
|
C1 |
C2 |
H7 |
108.693 |
| C1 |
C2 |
H8 |
108.693 |
|
C2 |
C1 |
F4 |
110.451 |
| C2 |
C1 |
H5 |
111.188 |
|
C2 |
C1 |
H6 |
111.188 |
| C2 |
C3 |
H9 |
111.083 |
|
C2 |
C3 |
H10 |
111.351 |
| C2 |
C3 |
H11 |
111.351 |
|
C3 |
C2 |
H7 |
110.439 |
| C3 |
C2 |
H8 |
110.439 |
|
F4 |
C1 |
H5 |
107.690 |
| F4 |
C1 |
H6 |
107.690 |
|
H5 |
C1 |
H6 |
108.494 |
| H7 |
C2 |
H8 |
106.943 |
|
H9 |
C3 |
H10 |
107.613 |
| H9 |
C3 |
H11 |
107.613 |
|
H10 |
C3 |
H11 |
107.643 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.092 |
|
|
|
| 2 |
C |
-0.225 |
|
|
|
| 3 |
C |
-0.368 |
|
|
|
| 4 |
F |
-0.235 |
|
|
|
| 5 |
H |
0.071 |
|
|
|
| 6 |
H |
0.071 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.113 |
|
|
|
| 11 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.701 |
0.791 |
0.000 |
1.876 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.254 |
2.125 |
0.000 |
| y |
2.125 |
-25.817 |
0.000 |
| z |
0.000 |
0.000 |
-24.566 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.063 |
2.125 |
0.000 |
| y |
2.125 |
0.593 |
0.000 |
| z |
0.000 |
0.000 |
2.470 |
|
| Polar |
|---|
| 3z2-r2 | 4.939 |
|---|
| x2-y2 | -2.437 |
|---|
| xy | 2.125 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.924 |
-0.346 |
0.000 |
| y |
-0.346 |
5.844 |
0.000 |
| z |
0.000 |
0.000 |
5.227 |
<r2> (average value of r
2) Å
2
| <r2> |
102.313 |
| (<r2>)1/2 |
10.115 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.355605 |
| Energy at 298.15K | |
| HF Energy | -218.355605 |
| Nuclear repulsion energy | 131.092268 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3010 |
22.20 |
|
|
|
| 2 |
A |
3131 |
2997 |
43.57 |
|
|
|
| 3 |
A |
3107 |
2974 |
43.05 |
|
|
|
| 4 |
A |
3093 |
2961 |
16.99 |
|
|
|
| 5 |
A |
3059 |
2928 |
24.60 |
|
|
|
| 6 |
A |
3057 |
2926 |
18.51 |
|
|
|
| 7 |
A |
3050 |
2920 |
32.09 |
|
|
|
| 8 |
A |
1514 |
1449 |
2.09 |
|
|
|
| 9 |
A |
1508 |
1444 |
8.01 |
|
|
|
| 10 |
A |
1496 |
1432 |
6.54 |
|
|
|
| 11 |
A |
1474 |
1411 |
2.79 |
|
|
|
| 12 |
A |
1427 |
1366 |
9.79 |
|
|
|
| 13 |
A |
1411 |
1350 |
4.69 |
|
|
|
| 14 |
A |
1380 |
1321 |
0.14 |
|
|
|
| 15 |
A |
1304 |
1249 |
1.62 |
|
|
|
| 16 |
A |
1279 |
1224 |
1.45 |
|
|
|
| 17 |
A |
1178 |
1128 |
1.47 |
|
|
|
| 18 |
A |
1132 |
1084 |
16.86 |
|
|
|
| 19 |
A |
1104 |
1057 |
50.47 |
|
|
|
| 20 |
A |
992 |
949 |
42.58 |
|
|
|
| 21 |
A |
921 |
881 |
2.90 |
|
|
|
| 22 |
A |
889 |
851 |
3.83 |
|
|
|
| 23 |
A |
769 |
736 |
0.67 |
|
|
|
| 24 |
A |
477 |
456 |
3.83 |
|
|
|
| 25 |
A |
308 |
294 |
1.09 |
|
|
|
| 26 |
A |
222 |
213 |
1.53 |
|
|
|
| 27 |
A |
142 |
136 |
2.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21284.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20371.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.791 |
0.508 |
0.298 |
| C2 |
-0.591 |
0.652 |
-0.283 |
| C3 |
-1.518 |
-0.480 |
0.118 |
| F4 |
1.387 |
-0.652 |
-0.171 |
| H5 |
1.433 |
1.342 |
0.016 |
| H6 |
0.758 |
0.439 |
1.386 |
| H7 |
-0.506 |
0.708 |
-1.368 |
| H8 |
-0.992 |
1.611 |
0.050 |
| H9 |
-2.500 |
-0.366 |
-0.336 |
| H10 |
-1.111 |
-1.438 |
-0.197 |
| H11 |
-1.651 |
-0.514 |
1.198 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5054 | 2.5181 | 1.3859 | 1.0897 | 1.0911 | 2.1204 | 2.1108 | 3.4632 | 2.7661 | 2.7963 |
C2 | 1.5054 | | 1.5175 | 2.3715 | 2.1587 | 2.1560 | 1.0901 | 1.0914 | 2.1643 | 2.1559 | 2.1628 | C3 | 2.5181 | 1.5175 | | 2.9244 | 3.4699 | 2.7628 | 2.1548 | 2.1579 | 1.0875 | 1.0872 | 1.0894 | F4 | 1.3859 | 2.3715 | 2.9244 | | 2.0034 | 2.0027 | 2.6200 | 3.2906 | 3.9008 | 2.6192 | 3.3354 | H5 | 1.0897 | 2.1587 | 3.4699 | 2.0034 | | 1.7747 | 2.4651 | 2.4396 | 4.3020 | 3.7748 | 3.7888 | H6 | 1.0911 | 2.1560 | 2.7628 | 2.0027 | 1.7747 | | 3.0421 | 2.4940 | 3.7715 | 3.0860 | 2.5973 | H7 | 2.1204 | 1.0901 | 2.1548 | 2.6200 | 2.4651 | 3.0421 | | 1.7499 | 2.4887 | 2.5185 | 3.0643 | H8 | 2.1108 | 1.0914 | 2.1579 | 3.2906 | 2.4396 | 2.4940 | 1.7499 | | 2.5164 | 3.0618 | 2.5040 | H9 | 3.4632 | 2.1643 | 1.0875 | 3.9008 | 4.3020 | 3.7715 | 2.4887 | 2.5164 | | 1.7600 | 1.7596 | H10 | 2.7661 | 2.1559 | 1.0872 | 2.6192 | 3.7748 | 3.0860 | 2.5185 | 3.0618 | 1.7600 | | 1.7589 | H11 | 2.7963 | 2.1628 | 1.0894 | 3.3354 | 3.7888 | 2.5973 | 3.0643 | 2.5040 | 1.7596 | 1.7589 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.817 |
|
C1 |
C2 |
H7 |
108.505 |
| C1 |
C2 |
H8 |
107.686 |
|
C2 |
C1 |
F4 |
110.147 |
| C2 |
C1 |
H5 |
111.580 |
|
C2 |
C1 |
H6 |
111.266 |
| C2 |
C3 |
H9 |
111.290 |
|
C2 |
C3 |
H10 |
110.639 |
| C2 |
C3 |
H11 |
111.053 |
|
C3 |
C2 |
H7 |
110.376 |
| C3 |
C2 |
H8 |
110.548 |
|
F4 |
C1 |
H5 |
107.445 |
| F4 |
C1 |
H6 |
107.300 |
|
H5 |
C1 |
H6 |
108.934 |
| H7 |
C2 |
H8 |
106.677 |
|
H9 |
C3 |
H10 |
108.051 |
| H9 |
C3 |
H11 |
107.852 |
|
H10 |
C3 |
H11 |
107.815 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.082 |
|
|
|
| 2 |
C |
-0.222 |
|
|
|
| 3 |
C |
-0.355 |
|
|
|
| 4 |
F |
-0.243 |
|
|
|
| 5 |
H |
0.085 |
|
|
|
| 6 |
H |
0.069 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.108 |
|
|
|
| 9 |
H |
0.119 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.967 |
1.423 |
0.449 |
1.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.881 |
1.654 |
0.651 |
| y |
1.654 |
-25.395 |
0.006 |
| z |
0.651 |
0.006 |
-24.747 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.811 |
1.654 |
0.651 |
| y |
1.654 |
0.419 |
0.006 |
| z |
0.651 |
0.006 |
1.391 |
|
| Polar |
|---|
| 3z2-r2 | 2.783 |
|---|
| x2-y2 | -1.486 |
|---|
| xy | 1.654 |
|---|
| xz | 0.651 |
|---|
| yz | 0.006 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.030 |
0.278 |
0.067 |
| y |
0.278 |
5.598 |
-0.044 |
| z |
0.067 |
-0.044 |
5.360 |
<r2> (average value of r
2) Å
2
| <r2> |
90.708 |
| (<r2>)1/2 |
9.524 |
