return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5F (Fluorobenzene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-330.965255
Energy at 298.15K-330.970923
HF Energy-329.670381
Nuclear repulsion energy270.749997
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3266 3101 0.47      
2 A1 3251 3087 9.13      
3 A1 3219 3056 0.14      
4 A1 1663 1579 51.01      
5 A1 1540 1462 82.45      
6 A1 1281 1217 85.24      
7 A1 1190 1130 8.88      
8 A1 1049 996 3.84      
9 A1 1031 978 0.00      
10 A1 831 789 21.99      
11 A1 523 497 3.70      
12 A2 986 936 0.00      
13 A2 853 810 0.00      
14 A2 429 407 0.00      
15 B1 995 945 0.02      
16 B1 929 882 8.10      
17 B1 788 748 72.86      
18 B1 709 673 15.61      
19 B1 520 494 9.13      
20 B1 242 230 0.08      
21 B2 3263 3098 1.91      
22 B2 3233 3070 5.81      
23 B2 1662 1578 6.32      
24 B2 1495 1419 1.09      
25 B2 1477 1402 0.77      
26 B2 1329 1262 0.44      
27 B2 1195 1135 0.19      
28 B2 1101 1045 7.43      
29 B2 620 589 0.12      
30 B2 407 386 2.00      

Unscaled Zero Point Vibrational Energy (zpe) 20537.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19498.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.19167 0.08643 0.05957

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
F1 0.000 0.000 2.268
C2 0.000 0.000 0.928
C3 0.000 1.208 0.259
C4 0.000 -1.208 0.259
C5 0.000 1.201 -1.129
C6 0.000 -1.201 -1.129
C7 0.000 0.000 -1.825
H8 0.000 2.124 0.821
H9 0.000 -2.124 0.821
H10 0.000 2.135 -1.663
H11 0.000 -2.135 -1.663
H12 0.000 0.000 -2.900

Atom - Atom Distances (Å)
  F1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
F11.34002.34422.34423.60233.60234.09242.56982.56984.47334.47335.1680
C21.34001.38101.38102.38132.38132.75252.12682.12683.35723.35723.8280
C32.34421.38102.41651.38752.78012.40861.07493.37962.13383.85643.3823
C42.34421.38102.41652.78011.38752.40863.37961.07493.85642.13383.3823
C53.60232.38131.38752.78012.40171.38802.15753.85461.07633.37842.1403
C63.60232.38132.78011.38752.40171.38803.85462.15753.37841.07632.1403
C74.09242.75252.40862.40861.38801.38803.39323.39322.14112.14111.0755
H82.56982.12681.07493.37962.15753.85463.39324.24832.48464.93094.2852
H92.56982.12683.37961.07493.85462.15753.39324.24834.93092.48464.2852
H104.47333.35722.13383.85641.07633.37842.14112.48464.93094.27012.4675
H114.47333.35723.85642.13383.37841.07632.14114.93092.48464.27012.4675
H125.16803.82803.38233.38232.14032.14031.07554.28524.28522.46752.4675

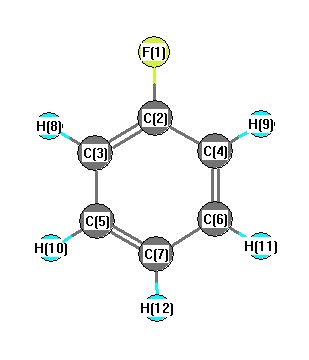
picture of Fluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
F1 C2 C3 118.968 F1 C2 C4 118.968
C2 C3 C5 118.662 C2 C3 H8 119.475
C2 C4 C6 118.662 C2 C4 H9 119.475
C3 C2 C4 122.064 C3 C5 C7 120.407
C3 C5 H10 119.474 C4 C6 C7 120.407
C4 C6 H11 119.474 C5 C3 H8 121.863
C5 C7 C6 119.798 C5 C7 H12 120.101
C6 C4 H9 121.863 C6 C7 H12 120.101
C7 C5 H10 120.119 C7 C6 H11 120.119
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability