All results from a given calculation for CH2FCH2CH2F (1,3-difluoropropane)
using model chemistry: CCSD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2 |
1A |
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -317.173141 |
| Energy at 298.15K | |
| HF Energy | -316.086666 |
| Nuclear repulsion energy | 188.153424 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.974 |
| C2 |
0.000 |
1.258 |
0.129 |
| C3 |
0.000 |
-1.258 |
0.129 |
| F4 |
1.178 |
1.316 |
-0.610 |
| F5 |
-1.178 |
-1.316 |
-0.610 |
| H6 |
0.884 |
-0.004 |
1.618 |
| H7 |
-0.884 |
0.004 |
1.618 |
| H8 |
-0.835 |
1.256 |
-0.574 |
| H9 |
-0.048 |
2.153 |
0.750 |
| H10 |
0.835 |
-1.256 |
-0.574 |
| H11 |
0.048 |
-2.153 |
0.750 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
F5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5156 | 1.5156 | 2.3725 | 2.3725 | 1.0935 | 1.0935 | 2.1615 | 2.1652 | 2.1615 | 2.1652 |
C2 | 1.5156 | | 2.5154 | 1.3915 | 2.9249 | 2.1427 | 2.1382 | 1.0910 | 1.0911 | 2.7404 | 3.4673 | C3 | 1.5156 | 2.5154 | | 2.9249 | 1.3915 | 2.1382 | 2.1427 | 2.7404 | 3.4673 | 1.0910 | 1.0911 | F4 | 2.3725 | 1.3915 | 2.9249 | | 3.5315 | 2.6062 | 3.3070 | 2.0134 | 2.0137 | 2.5949 | 3.8935 | F5 | 2.3725 | 2.9249 | 1.3915 | 3.5315 | | 3.3070 | 2.6062 | 2.5949 | 3.8935 | 2.0134 | 2.0137 | H6 | 1.0935 | 2.1427 | 2.1382 | 2.6062 | 3.3070 | | 1.7679 | 3.0571 | 2.5047 | 2.5251 | 2.4639 | H7 | 1.0935 | 2.1382 | 2.1427 | 3.3070 | 2.6062 | 1.7679 | | 2.5251 | 2.4639 | 3.0571 | 2.5047 | H8 | 2.1615 | 1.0910 | 2.7404 | 2.0134 | 2.5949 | 3.0571 | 2.5251 | | 1.7824 | 3.0163 | 3.7625 | H9 | 2.1652 | 1.0911 | 3.4673 | 2.0137 | 3.8935 | 2.5047 | 2.4639 | 1.7824 | | 3.7625 | 4.3072 | H10 | 2.1615 | 2.7404 | 1.0910 | 2.5949 | 2.0134 | 2.5251 | 3.0571 | 3.0163 | 3.7625 | | 1.7824 | H11 | 2.1652 | 3.4673 | 1.0911 | 3.8935 | 2.0137 | 2.4639 | 2.5047 | 3.7625 | 4.3072 | 1.7824 | |
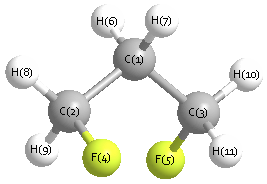 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.320 |
|
C1 |
C2 |
H8 |
110.996 |
| C1 |
C2 |
H9 |
111.287 |
|
C1 |
C3 |
F5 |
109.320 |
| C1 |
C3 |
H10 |
110.996 |
|
C1 |
C3 |
H11 |
111.287 |
| C2 |
C1 |
C3 |
112.164 |
|
C2 |
C1 |
H6 |
109.353 |
| C2 |
C1 |
H7 |
109.000 |
|
C3 |
C1 |
H6 |
109.000 |
| C3 |
C1 |
H7 |
109.353 |
|
F4 |
C2 |
H8 |
107.785 |
| F4 |
C2 |
H9 |
107.797 |
|
F5 |
C3 |
H10 |
107.785 |
| F5 |
C3 |
H11 |
107.797 |
|
H6 |
C1 |
H7 |
107.875 |
| H8 |
C2 |
H9 |
109.537 |
|
H10 |
C3 |
H11 |
109.537 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
