return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2CH2F (1,3-difluoropropane)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-317.178862
Energy at 298.15K 
HF Energy-316.087144
Nuclear repulsion energy189.200687
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3175 3015 22.61 44.08 0.69 0.81
2 A 3128 2970 2.39 228.92 0.01 0.03
3 A 3108 2951 5.08 80.47 0.18 0.30
4 A 1550 1472 0.64 9.30 0.73 0.85
5 A 1480 1405 9.36 5.27 0.71 0.83
6 A 1463 1389 6.64 1.30 0.05 0.10
7 A 1345 1277 0.85 13.13 0.68 0.81
8 A 1268 1204 0.74 5.07 0.60 0.75
9 A 1155 1096 23.07 1.76 0.74 0.85
10 A 1038 985 39.55 3.69 0.73 0.85
11 A 905 859 0.03 9.20 0.08 0.14
12 A 544 516 3.21 0.56 0.64 0.78
13 A 251 239 3.89 0.24 0.25 0.40
14 A 90 86 3.68 0.06 0.73 0.84
15 B 3176 3015 20.27 31.79 0.75 0.86
16 B 3152 2992 20.15 64.15 0.75 0.86
17 B 3126 2967 48.77 41.10 0.75 0.86
18 B 1550 1471 4.27 0.05 0.75 0.86
19 B 1441 1368 12.54 0.23 0.75 0.86
20 B 1414 1342 1.23 0.52 0.75 0.86
21 B 1290 1225 8.05 0.67 0.75 0.86
22 B 1148 1090 8.70 0.83 0.75 0.86
23 B 1122 1065 104.12 3.04 0.75 0.86
24 B 986 936 22.30 2.55 0.75 0.86
25 B 798 758 2.09 0.69 0.75 0.86
26 B 429 408 7.23 0.32 0.75 0.86
27 B 198 188 10.18 0.11 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 20164.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19144.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.31256 0.09911 0.09326

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.968
C2 0.000 1.248 0.129
C3 0.000 -1.248 0.129
F4 1.176 1.307 -0.606
F5 -1.176 -1.307 -0.606
H6 0.879 -0.001 1.607
H7 -0.879 0.001 1.607
H8 -0.828 1.244 -0.571
H9 -0.050 2.138 0.746
H10 0.828 -1.244 -0.571
H11 0.050 -2.138 0.746

Atom - Atom Distances (Å)
  C1 C2 C3 F4 F5 H6 H7 H8 H9 H10 H11
C11.50361.50362.35982.35981.08661.08662.14482.15002.14482.1500
C21.50362.49551.38802.90682.12552.12411.08401.08412.71713.4417
C31.50362.49552.90681.38802.12412.12552.71713.44171.08401.0841
F42.35981.38802.90683.51632.58833.29022.00522.00512.57463.8681
F52.35982.90681.38803.51633.29022.58832.57463.86812.00522.0051
H61.08662.12552.12412.58833.29021.75713.03432.48602.50822.4484
H71.08662.12412.12553.29022.58831.75712.50822.44843.03432.4860
H82.14481.08402.71712.00522.57463.03432.50821.77152.98833.7336
H92.15001.08413.44172.00513.86812.48602.44841.77153.73364.2769
H102.14482.71711.08402.57462.00522.50823.03432.98833.73361.7715
H112.15003.44171.08413.86812.00512.44842.48603.73364.27691.7715

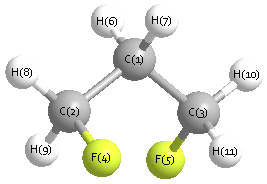
picture of 1,3-difluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 109.322 C1 C2 H8 110.925
C1 C2 H9 111.337 C1 C3 F5 109.322
C1 C3 H10 110.925 C1 C3 H11 111.337
C2 C1 C3 112.162 C2 C1 H6 109.227
C2 C1 H7 109.114 C3 C1 H6 109.114
C3 C1 H7 109.227 F4 C2 H8 107.787
F4 C2 H9 107.769 F5 C3 H10 107.787
F5 C3 H11 107.769 H6 C1 H7 107.903
H8 C2 H9 109.582 H10 C3 H11 109.582
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability