Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -245.178342 |
| Energy at 298.15K | |
| HF Energy | -245.178342 |
| Nuclear repulsion energy | 124.148857 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3853 |
3696 |
88.90 |
|
|
|
| 2 |
A' |
3807 |
3651 |
68.74 |
|
|
|
| 3 |
A' |
3668 |
3519 |
56.39 |
|
|
|
| 4 |
A' |
1864 |
1787 |
529.10 |
|
|
|
| 5 |
A' |
1622 |
1556 |
137.08 |
|
|
|
| 6 |
A' |
1452 |
1392 |
133.07 |
|
|
|
| 7 |
A' |
1239 |
1189 |
193.35 |
|
|
|
| 8 |
A' |
1085 |
1041 |
39.12 |
|
|
|
| 9 |
A' |
975 |
936 |
38.48 |
|
|
|
| 10 |
A' |
592 |
568 |
35.47 |
|
|
|
| 11 |
A' |
490 |
470 |
7.00 |
|
|
|
| 12 |
A" |
799 |
766 |
25.72 |
|
|
|
| 13 |
A" |
594 |
569 |
54.37 |
|
|
|
| 14 |
A" |
487 |
467 |
63.41 |
|
|
|
| 15 |
A" |
141i |
136i |
208.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11191.9 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 10735.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.128 |
0.000 |
| O2 |
-0.061 |
1.332 |
0.000 |
| N3 |
1.134 |
-0.604 |
0.000 |
| O4 |
-1.089 |
-0.674 |
0.000 |
| H5 |
2.009 |
-0.122 |
0.000 |
| H6 |
1.105 |
-1.604 |
0.000 |
| H7 |
-1.846 |
-0.079 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2054 | 1.3496 | 1.3527 | 2.0246 | 2.0545 | 1.8572 |
O2 | 1.2054 | | 2.2752 | 2.2540 | 2.5298 | 3.1591 | 2.2751 | N3 | 1.3496 | 2.2752 | | 2.2239 | 0.9997 | 1.0002 | 3.0250 | O4 | 1.3527 | 2.2540 | 2.2239 | | 3.1472 | 2.3831 | 0.9619 | H5 | 2.0246 | 2.5298 | 0.9997 | 3.1472 | | 1.7363 | 3.8549 | H6 | 2.0545 | 3.1591 | 1.0002 | 2.3831 | 1.7363 | | 3.3210 | H7 | 1.8572 | 2.2751 | 3.0250 | 0.9619 | 3.8549 | 3.3210 | |
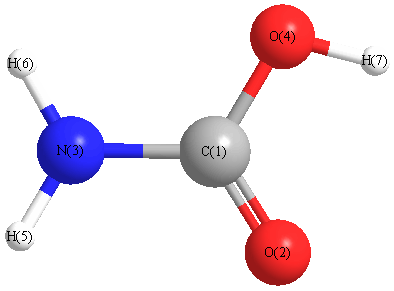 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.280 |
|
C1 |
N3 |
H6 |
121.217 |
| C1 |
O4 |
H7 |
105.469 |
|
O2 |
C1 |
N3 |
125.774 |
| O2 |
C1 |
O4 |
123.455 |
|
N3 |
C1 |
O4 |
110.771 |
| H5 |
N3 |
H6 |
120.503 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.304 |
|
|
|
| 2 |
O |
-0.360 |
|
|
|
| 3 |
N |
-0.217 |
|
|
|
| 4 |
O |
-0.277 |
|
|
|
| 5 |
H |
0.166 |
|
|
|
| 6 |
H |
0.161 |
|
|
|
| 7 |
H |
0.223 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.818 |
-2.401 |
0.000 |
2.537 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.248 |
-2.376 |
0.000 |
| y |
-2.376 |
-25.882 |
0.000 |
| z |
0.000 |
0.000 |
-23.716 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.551 |
-2.376 |
0.000 |
| y |
-2.376 |
-6.400 |
0.000 |
| z |
0.000 |
0.000 |
-3.151 |
|
| Polar |
|---|
| 3z2-r2 | -6.303 |
|---|
| x2-y2 | 10.634 |
|---|
| xy | -2.376 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.552 |
-0.306 |
0.000 |
| y |
-0.306 |
4.458 |
0.000 |
| z |
0.000 |
0.000 |
2.516 |
<r2> (average value of r
2) Å
2
| <r2> |
63.755 |
| (<r2>)1/2 |
7.985 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -245.178358 |
| Energy at 298.15K | -245.183024 |
| HF Energy | -245.178358 |
| Nuclear repulsion energy | 124.139484 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3854 |
3696 |
88.61 |
|
|
|
| 2 |
A |
3795 |
3641 |
64.56 |
|
|
|
| 3 |
A |
3661 |
3511 |
52.34 |
|
|
|
| 4 |
A |
1864 |
1788 |
521.27 |
|
|
|
| 5 |
A |
1625 |
1558 |
130.52 |
|
|
|
| 6 |
A |
1452 |
1393 |
133.93 |
|
|
|
| 7 |
A |
1242 |
1191 |
185.19 |
|
|
|
| 8 |
A |
1090 |
1045 |
48.35 |
|
|
|
| 9 |
A |
977 |
937 |
37.76 |
|
|
|
| 10 |
A |
798 |
766 |
29.77 |
|
|
|
| 11 |
A |
595 |
571 |
50.18 |
|
|
|
| 12 |
A |
587 |
563 |
51.42 |
|
|
|
| 13 |
A |
498 |
478 |
29.57 |
|
|
|
| 14 |
A |
479 |
459 |
28.77 |
|
|
|
| 15 |
A |
204 |
196 |
220.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11360.2 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 10896.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.039 |
0.122 |
-0.001 |
| O2 |
-0.460 |
1.251 |
0.004 |
| N3 |
1.265 |
-0.234 |
-0.033 |
| O4 |
-0.835 |
-0.971 |
0.002 |
| H5 |
1.944 |
0.490 |
0.086 |
| H6 |
1.531 |
-1.190 |
0.100 |
| H7 |
-1.735 |
-0.632 |
0.005 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2052 | 1.3517 | 1.3522 | 2.0188 | 2.0488 | 1.8560 |
O2 | 1.2052 | | 2.2768 | 2.2537 | 2.5234 | 3.1521 | 2.2739 | N3 | 1.3517 | 2.2768 | | 2.2250 | 1.0004 | 1.0010 | 3.0258 | O4 | 1.3522 | 2.2537 | 2.2250 | | 3.1407 | 2.3782 | 0.9620 | H5 | 2.0188 | 2.5234 | 1.0004 | 3.1407 | | 1.7300 | 3.8471 | H6 | 2.0488 | 3.1521 | 1.0010 | 2.3782 | 1.7300 | | 3.3149 | H7 | 1.8560 | 2.2739 | 3.0258 | 0.9620 | 3.8471 | 3.3149 | |
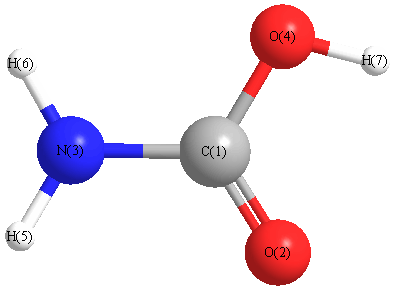 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.469 |
|
C1 |
N3 |
H6 |
120.384 |
| C1 |
O4 |
H7 |
105.402 |
|
O2 |
C1 |
N3 |
125.763 |
| O2 |
C1 |
O4 |
123.482 |
|
N3 |
C1 |
O4 |
110.747 |
| H5 |
N3 |
H6 |
119.632 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.305 |
|
|
|
| 2 |
O |
-0.359 |
|
|
|
| 3 |
N |
-0.219 |
|
|
|
| 4 |
O |
-0.276 |
|
|
|
| 5 |
H |
0.165 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
| 7 |
H |
0.222 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.446 |
-2.020 |
0.369 |
2.511 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.993 |
1.143 |
0.737 |
| y |
1.143 |
-26.300 |
-0.197 |
| z |
0.737 |
-0.197 |
-23.680 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.997 |
1.143 |
0.737 |
| y |
1.143 |
-6.963 |
-0.197 |
| z |
0.737 |
-0.197 |
-3.033 |
|
| Polar |
|---|
| 3z2-r2 | -6.067 |
|---|
| x2-y2 | 11.307 |
|---|
| xy | 1.143 |
|---|
| xz | 0.737 |
|---|
| yz | -0.197 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.721 |
-0.220 |
0.002 |
| y |
-0.220 |
4.301 |
0.003 |
| z |
0.002 |
0.003 |
2.526 |
<r2> (average value of r
2) Å
2
| <r2> |
63.757 |
| (<r2>)1/2 |
7.985 |
