Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -358.510701 |
| Energy at 298.15K | -358.515787 |
| Nuclear repulsion energy | 233.510637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3800 |
3645 |
85.92 |
|
|
|
| 2 |
A' |
3768 |
3614 |
83.78 |
|
|
|
| 3 |
A' |
3627 |
3479 |
59.03 |
|
|
|
| 4 |
A' |
1847 |
1772 |
42.92 |
|
|
|
| 5 |
A' |
1832 |
1758 |
579.05 |
|
|
|
| 6 |
A' |
1604 |
1538 |
120.81 |
|
|
|
| 7 |
A' |
1442 |
1383 |
22.80 |
|
|
|
| 8 |
A' |
1333 |
1279 |
68.21 |
|
|
|
| 9 |
A' |
1202 |
1153 |
270.58 |
|
|
|
| 10 |
A' |
1103 |
1058 |
2.03 |
|
|
|
| 11 |
A' |
788 |
756 |
9.45 |
|
|
|
| 12 |
A' |
616 |
591 |
72.58 |
|
|
|
| 13 |
A' |
531 |
510 |
0.22 |
|
|
|
| 14 |
A' |
415 |
398 |
3.84 |
|
|
|
| 15 |
A' |
269 |
258 |
15.89 |
|
|
|
| 16 |
A" |
850 |
815 |
5.04 |
|
|
|
| 17 |
A" |
681 |
653 |
111.26 |
|
|
|
| 18 |
A" |
637 |
611 |
28.57 |
|
|
|
| 19 |
A" |
442 |
424 |
12.87 |
|
|
|
| 20 |
A" |
337 |
324 |
180.08 |
|
|
|
| 21 |
A" |
68 |
65 |
4.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13596.1 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 13041.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.751 |
0.000 |
| C2 |
-0.056 |
-0.787 |
0.000 |
| O3 |
-1.092 |
-1.403 |
0.000 |
| O4 |
1.031 |
1.371 |
0.000 |
| O5 |
-1.208 |
1.295 |
0.000 |
| N6 |
1.184 |
-1.307 |
0.000 |
| H7 |
1.301 |
-2.302 |
0.000 |
| H8 |
1.978 |
-0.692 |
0.000 |
| H9 |
-1.085 |
2.252 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5395 | 2.4154 | 1.2028 | 1.3242 | 2.3743 | 3.3190 | 2.4481 | 1.8524 |
C2 | 1.5395 | | 1.2056 | 2.4163 | 2.3792 | 1.3443 | 2.0335 | 2.0354 | 3.2093 | O3 | 2.4154 | 1.2056 | | 3.4933 | 2.7002 | 2.2782 | 2.5560 | 3.1509 | 3.6557 | O4 | 1.2028 | 2.4163 | 3.4933 | | 2.2398 | 2.6821 | 3.6832 | 2.2698 | 2.2923 | O5 | 1.3242 | 2.3792 | 2.7002 | 2.2398 | | 3.5337 | 4.3850 | 3.7538 | 0.9657 | N6 | 2.3743 | 1.3443 | 2.2782 | 2.6821 | 3.5337 | | 1.0024 | 1.0037 | 4.2211 | H7 | 3.3190 | 2.0335 | 2.5560 | 3.6832 | 4.3850 | 1.0024 | | 1.7468 | 5.1418 | H8 | 2.4481 | 2.0354 | 3.1509 | 2.2698 | 3.7538 | 1.0037 | 1.7468 | | 4.2485 | H9 | 1.8524 | 3.2093 | 3.6557 | 2.2923 | 0.9657 | 4.2211 | 5.1418 | 4.2485 | |
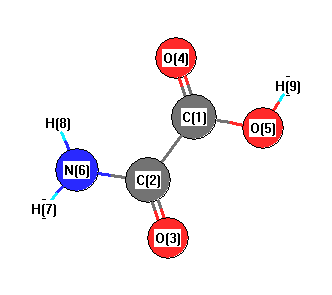 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.800 |
|
C1 |
C2 |
N6 |
110.663 |
| C1 |
O5 |
H9 |
106.943 |
|
C2 |
C1 |
O4 |
123.086 |
| C2 |
C1 |
O5 |
112.149 |
|
C2 |
N6 |
H7 |
119.409 |
| C2 |
N6 |
H8 |
119.492 |
|
O3 |
C2 |
N6 |
126.537 |
| O4 |
C1 |
O5 |
124.765 |
|
H7 |
N6 |
H8 |
121.099 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.273 |
|
|
|
| 2 |
C |
0.160 |
|
|
|
| 3 |
O |
-0.292 |
|
|
|
| 4 |
O |
-0.311 |
|
|
|
| 5 |
O |
-0.193 |
|
|
|
| 6 |
N |
-0.184 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.133 |
0.638 |
0.000 |
2.227 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.735 |
-10.184 |
0.000 |
| y |
-10.184 |
-28.388 |
0.000 |
| z |
0.000 |
0.000 |
-33.250 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.917 |
-10.184 |
0.000 |
| y |
-10.184 |
5.605 |
0.000 |
| z |
0.000 |
0.000 |
-1.688 |
|
| Polar |
|---|
| 3z2-r2 | -3.376 |
|---|
| x2-y2 | -6.348 |
|---|
| xy | -10.184 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.104 |
-0.150 |
0.000 |
| y |
-0.150 |
6.555 |
0.000 |
| z |
0.000 |
0.000 |
3.445 |
<r2> (average value of r
2) Å
2
| <r2> |
140.494 |
| (<r2>)1/2 |
11.853 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -358.519957 |
| Energy at 298.15K | -358.525308 |
| Nuclear repulsion energy | 235.140612 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3756 |
3603 |
91.14 |
|
|
|
| 2 |
A' |
3618 |
3470 |
80.96 |
|
|
|
| 3 |
A' |
3603 |
3456 |
144.09 |
|
|
|
| 4 |
A' |
1889 |
1812 |
265.17 |
|
|
|
| 5 |
A' |
1816 |
1742 |
324.94 |
|
|
|
| 6 |
A' |
1610 |
1545 |
73.11 |
|
|
|
| 7 |
A' |
1462 |
1402 |
203.74 |
|
|
|
| 8 |
A' |
1361 |
1305 |
360.39 |
|
|
|
| 9 |
A' |
1232 |
1182 |
3.13 |
|
|
|
| 10 |
A' |
1109 |
1064 |
3.87 |
|
|
|
| 11 |
A' |
817 |
784 |
9.19 |
|
|
|
| 12 |
A' |
638 |
612 |
13.18 |
|
|
|
| 13 |
A' |
549 |
527 |
1.98 |
|
|
|
| 14 |
A' |
403 |
387 |
7.94 |
|
|
|
| 15 |
A' |
268 |
257 |
45.38 |
|
|
|
| 16 |
A" |
849 |
814 |
0.02 |
|
|
|
| 17 |
A" |
778 |
746 |
94.60 |
|
|
|
| 18 |
A" |
671 |
644 |
2.19 |
|
|
|
| 19 |
A" |
480 |
461 |
119.48 |
|
|
|
| 20 |
A" |
391 |
375 |
98.52 |
|
|
|
| 21 |
A" |
120 |
115 |
7.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13710.2 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 13150.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
-0.793 |
0.000 |
| C2 |
0.000 |
0.744 |
0.000 |
| O3 |
-1.067 |
1.329 |
0.000 |
| O4 |
1.016 |
-1.446 |
0.000 |
| O5 |
-1.216 |
-1.277 |
0.000 |
| N6 |
1.218 |
1.284 |
0.000 |
| H7 |
1.333 |
2.280 |
0.000 |
| H8 |
2.013 |
0.670 |
0.000 |
| H9 |
-1.799 |
-0.494 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5376 | 2.3810 | 1.1974 | 1.3196 | 2.4020 | 3.3456 | 2.4794 | 1.8357 |
C2 | 1.5376 | | 1.2171 | 2.4145 | 2.3587 | 1.3321 | 2.0339 | 2.0148 | 2.1840 | O3 | 2.3810 | 1.2171 | | 3.4700 | 2.6101 | 2.2857 | 2.5822 | 3.1505 | 1.9641 | O4 | 1.1974 | 2.4145 | 3.4700 | | 2.2380 | 2.7377 | 3.7400 | 2.3397 | 2.9716 | O5 | 1.3196 | 2.3587 | 2.6101 | 2.2380 | | 3.5329 | 4.3762 | 3.7708 | 0.9765 | N6 | 2.4020 | 1.3321 | 2.2857 | 2.7377 | 3.5329 | | 1.0029 | 1.0050 | 3.5018 | H7 | 3.3456 | 2.0339 | 2.5822 | 3.7400 | 4.3762 | 1.0029 | | 1.7481 | 4.1841 | H8 | 2.4794 | 2.0148 | 3.1505 | 2.3397 | 3.7708 | 1.0050 | 1.7481 | | 3.9863 | H9 | 1.8357 | 2.1840 | 1.9641 | 2.9716 | 0.9765 | 3.5018 | 4.1841 | 3.9863 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.157 |
|
C1 |
C2 |
N6 |
113.461 |
| C1 |
O5 |
H9 |
105.184 |
|
C2 |
C1 |
O4 |
123.485 |
| C2 |
C1 |
O5 |
111.053 |
|
C2 |
N6 |
H7 |
120.505 |
| C2 |
N6 |
H8 |
118.437 |
|
O3 |
C2 |
N6 |
127.383 |
| O4 |
C1 |
O5 |
125.462 |
|
H7 |
N6 |
H8 |
121.058 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.269 |
|
|
|
| 2 |
C |
0.167 |
|
|
|
| 3 |
O |
-0.342 |
|
|
|
| 4 |
O |
-0.292 |
|
|
|
| 5 |
O |
-0.200 |
|
|
|
| 6 |
N |
-0.168 |
|
|
|
| 7 |
H |
0.162 |
|
|
|
| 8 |
H |
0.178 |
|
|
|
| 9 |
H |
0.226 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.538 |
2.578 |
0.000 |
3.002 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.327 |
7.308 |
0.000 |
| y |
7.308 |
-36.877 |
0.000 |
| z |
0.000 |
0.000 |
-33.183 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.703 |
7.308 |
0.000 |
| y |
7.308 |
-5.122 |
0.000 |
| z |
0.000 |
0.000 |
0.419 |
|
| Polar |
|---|
| 3z2-r2 | 0.838 |
|---|
| x2-y2 | 6.550 |
|---|
| xy | 7.308 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.331 |
0.022 |
0.000 |
| y |
0.022 |
6.166 |
0.000 |
| z |
0.000 |
0.000 |
3.421 |
<r2> (average value of r
2) Å
2
| <r2> |
137.984 |
| (<r2>)1/2 |
11.747 |
