Jump to
S1C2
Energy calculated at B3LYP/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -358.609191 |
| Energy at 298.15K | -358.614228 |
| Nuclear repulsion energy | 232.465118 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3738 |
3613 |
79.66 |
|
|
|
| 2 |
A' |
3723 |
3598 |
76.40 |
|
|
|
| 3 |
A' |
3587 |
3467 |
55.76 |
|
|
|
| 4 |
A' |
1807 |
1747 |
36.82 |
|
|
|
| 5 |
A' |
1795 |
1735 |
557.88 |
|
|
|
| 6 |
A' |
1598 |
1545 |
119.00 |
|
|
|
| 7 |
A' |
1411 |
1364 |
10.64 |
|
|
|
| 8 |
A' |
1316 |
1272 |
60.35 |
|
|
|
| 9 |
A' |
1181 |
1142 |
290.14 |
|
|
|
| 10 |
A' |
1095 |
1059 |
3.29 |
|
|
|
| 11 |
A' |
772 |
746 |
7.95 |
|
|
|
| 12 |
A' |
613 |
593 |
70.56 |
|
|
|
| 13 |
A' |
525 |
507 |
0.37 |
|
|
|
| 14 |
A' |
413 |
399 |
3.88 |
|
|
|
| 15 |
A' |
270 |
261 |
15.25 |
|
|
|
| 16 |
A" |
837 |
809 |
4.76 |
|
|
|
| 17 |
A" |
673 |
650 |
112.56 |
|
|
|
| 18 |
A" |
629 |
608 |
26.17 |
|
|
|
| 19 |
A" |
437 |
422 |
11.23 |
|
|
|
| 20 |
A" |
331 |
320 |
181.13 |
|
|
|
| 21 |
A" |
65 |
63 |
4.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13407.6 cm
-1
Scaled (by 0.9666) Zero Point Vibrational Energy (zpe) 12959.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.756 |
0.000 |
| C2 |
-0.053 |
-0.790 |
0.000 |
| O3 |
-1.091 |
-1.410 |
0.000 |
| O4 |
1.031 |
1.382 |
0.000 |
| O5 |
-1.217 |
1.298 |
0.000 |
| N6 |
1.192 |
-1.314 |
0.000 |
| H7 |
1.304 |
-2.313 |
0.000 |
| H8 |
1.991 |
-0.705 |
0.000 |
| H9 |
-1.102 |
2.261 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5467 | 2.4254 | 1.2063 | 1.3324 | 2.3885 | 3.3340 | 2.4695 | 1.8655 |
C2 | 1.5467 | | 1.2094 | 2.4276 | 2.3910 | 1.3504 | 2.0396 | 2.0458 | 3.2264 | O3 | 2.4254 | 1.2094 | | 3.5073 | 2.7118 | 2.2847 | 2.5597 | 3.1620 | 3.6714 | O4 | 1.2063 | 2.4276 | 3.5073 | | 2.2492 | 2.7014 | 3.7049 | 2.2975 | 2.3066 | O5 | 1.3324 | 2.3910 | 2.7118 | 2.2492 | | 3.5535 | 4.4041 | 3.7822 | 0.9694 | N6 | 2.3885 | 1.3504 | 2.2847 | 2.7014 | 3.5535 | | 1.0046 | 1.0054 | 4.2478 | H7 | 3.3340 | 2.0396 | 2.5597 | 3.7049 | 4.4041 | 1.0046 | | 1.7482 | 5.1680 | H8 | 2.4695 | 2.0458 | 3.1620 | 2.2975 | 3.7822 | 1.0054 | 1.7482 | | 4.2853 | H9 | 1.8655 | 3.2264 | 3.6714 | 2.3066 | 0.9694 | 4.2478 | 5.1680 | 4.2853 | |
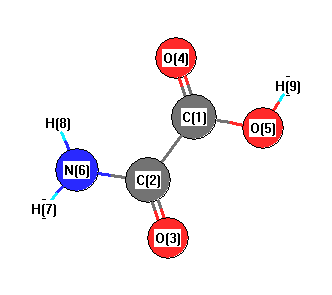 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.814 |
|
C1 |
C2 |
N6 |
110.889 |
| C1 |
O5 |
H9 |
107.225 |
|
C2 |
C1 |
O4 |
123.249 |
| C2 |
C1 |
O5 |
112.080 |
|
C2 |
N6 |
H7 |
119.292 |
| C2 |
N6 |
H8 |
119.838 |
|
O3 |
C2 |
N6 |
126.297 |
| O4 |
C1 |
O5 |
124.670 |
|
H7 |
N6 |
H8 |
120.869 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.274 |
|
|
0.554 |
| 2 |
C |
0.165 |
|
|
0.640 |
| 3 |
O |
-0.288 |
|
|
-0.503 |
| 4 |
O |
-0.306 |
|
|
-0.506 |
| 5 |
O |
-0.206 |
|
|
-0.541 |
| 6 |
N |
-0.204 |
|
|
-0.905 |
| 7 |
H |
0.168 |
|
|
0.426 |
| 8 |
H |
0.178 |
|
|
0.401 |
| 9 |
H |
0.219 |
|
|
0.434 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.144 |
0.611 |
0.000 |
2.229 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.152 |
0.594 |
0.000 |
2.232 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.814 |
7.955 |
0.004 |
| y |
7.955 |
-39.092 |
0.002 |
| z |
0.004 |
0.002 |
-33.477 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.471 |
7.955 |
0.004 |
| y |
7.955 |
-9.947 |
0.002 |
| z |
0.004 |
0.002 |
-1.524 |
|
| Polar |
|---|
| 3z2-r2 | -3.049 |
|---|
| x2-y2 | 14.278 |
|---|
| xy | 7.955 |
|---|
| xz | 0.004 |
|---|
| yz | 0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.825 |
0.285 |
0.000 |
| y |
0.285 |
7.101 |
0.000 |
| z |
0.000 |
0.000 |
3.457 |
<r2> (average value of r
2) Å
2
| <r2> |
141.860 |
| (<r2>)1/2 |
11.910 |
Jump to
S1C1
Energy calculated at B3LYP/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -358.615842 |
| Energy at 298.15K | -358.621139 |
| Nuclear repulsion energy | 233.993623 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3712 |
3588 |
84.25 |
|
|
|
| 2 |
A' |
3578 |
3459 |
109.63 |
|
|
|
| 3 |
A' |
3575 |
3455 |
95.94 |
|
|
|
| 4 |
A' |
1850 |
1788 |
249.45 |
|
|
|
| 5 |
A' |
1776 |
1717 |
314.37 |
|
|
|
| 6 |
A' |
1603 |
1549 |
74.98 |
|
|
|
| 7 |
A' |
1433 |
1385 |
182.11 |
|
|
|
| 8 |
A' |
1342 |
1297 |
363.84 |
|
|
|
| 9 |
A' |
1208 |
1167 |
13.12 |
|
|
|
| 10 |
A' |
1102 |
1065 |
2.10 |
|
|
|
| 11 |
A' |
800 |
773 |
9.42 |
|
|
|
| 12 |
A' |
633 |
612 |
13.32 |
|
|
|
| 13 |
A' |
542 |
524 |
1.93 |
|
|
|
| 14 |
A' |
403 |
389 |
6.60 |
|
|
|
| 15 |
A' |
270 |
261 |
42.98 |
|
|
|
| 16 |
A" |
837 |
809 |
0.04 |
|
|
|
| 17 |
A" |
755 |
730 |
95.21 |
|
|
|
| 18 |
A" |
662 |
640 |
2.80 |
|
|
|
| 19 |
A" |
473 |
457 |
112.97 |
|
|
|
| 20 |
A" |
384 |
372 |
103.52 |
|
|
|
| 21 |
A" |
115 |
111 |
7.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13526.0 cm
-1
Scaled (by 0.9666) Zero Point Vibrational Energy (zpe) 13074.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
-0.796 |
0.000 |
| C2 |
0.000 |
0.749 |
0.000 |
| O3 |
-1.068 |
1.340 |
0.000 |
| O4 |
1.018 |
-1.452 |
0.000 |
| O5 |
-1.221 |
-1.291 |
0.000 |
| N6 |
1.223 |
1.292 |
0.000 |
| H7 |
1.334 |
2.291 |
0.000 |
| H8 |
2.025 |
0.684 |
0.000 |
| H9 |
-1.820 |
-0.518 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5448 | 2.3935 | 1.2008 | 1.3281 | 2.4140 | 3.3584 | 2.4982 | 1.8530 |
C2 | 1.5448 | | 1.2209 | 2.4243 | 2.3769 | 1.3381 | 2.0392 | 2.0256 | 2.2172 | O3 | 2.3935 | 1.2209 | | 3.4849 | 2.6352 | 2.2913 | 2.5835 | 3.1615 | 2.0040 | O4 | 1.2008 | 2.4243 | 3.4849 | | 2.2444 | 2.7515 | 3.7561 | 2.3607 | 2.9878 | O5 | 1.3281 | 2.3769 | 2.6352 | 2.2444 | | 3.5556 | 4.3996 | 3.7987 | 0.9784 | N6 | 2.4140 | 1.3381 | 2.2913 | 2.7515 | 3.5556 | | 1.0051 | 1.0066 | 3.5404 | H7 | 3.3584 | 2.0392 | 2.5835 | 3.7561 | 4.3996 | 1.0051 | | 1.7496 | 4.2233 | H8 | 2.4982 | 2.0256 | 3.1615 | 2.3607 | 3.7987 | 1.0066 | 1.7496 | | 4.0279 | H9 | 1.8530 | 2.2172 | 2.0040 | 2.9878 | 0.9784 | 3.5404 | 4.2233 | 4.0279 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.406 |
|
C1 |
C2 |
N6 |
113.532 |
| C1 |
O5 |
H9 |
105.910 |
|
C2 |
C1 |
O4 |
123.524 |
| C2 |
C1 |
O5 |
111.434 |
|
C2 |
N6 |
H7 |
120.318 |
| C2 |
N6 |
H8 |
118.833 |
|
O3 |
C2 |
N6 |
127.062 |
| O4 |
C1 |
O5 |
125.042 |
|
H7 |
N6 |
H8 |
120.849 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.271 |
|
|
|
| 2 |
C |
0.169 |
|
|
|
| 3 |
O |
-0.338 |
|
|
|
| 4 |
O |
-0.287 |
|
|
|
| 5 |
O |
-0.212 |
|
|
|
| 6 |
N |
-0.189 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.230 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.516 |
2.553 |
0.000 |
2.969 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.603 |
7.364 |
0.000 |
| y |
7.364 |
-37.379 |
0.000 |
| z |
0.000 |
0.000 |
-33.414 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.794 |
7.364 |
0.000 |
| y |
7.364 |
-5.371 |
0.000 |
| z |
0.000 |
0.000 |
0.577 |
|
| Polar |
|---|
| 3z2-r2 | 1.153 |
|---|
| x2-y2 | 6.777 |
|---|
| xy | 7.364 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.453 |
0.048 |
0.000 |
| y |
0.048 |
6.286 |
0.000 |
| z |
0.000 |
0.000 |
3.438 |
<r2> (average value of r
2) Å
2
| <r2> |
139.495 |
| (<r2>)1/2 |
11.811 |
