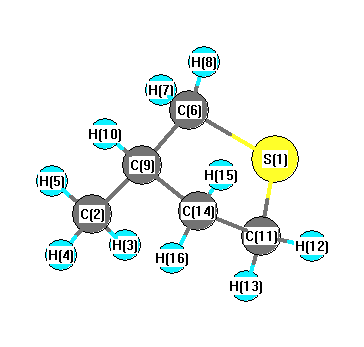
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
3005 |
14.71 |
|
|
|
| 2 |
A |
3133 |
2999 |
16.21 |
|
|
|
| 3 |
A |
3131 |
2997 |
28.86 |
|
|
|
| 4 |
A |
3128 |
2994 |
22.15 |
|
|
|
| 5 |
A |
3108 |
2975 |
27.55 |
|
|
|
| 6 |
A |
3084 |
2952 |
5.81 |
|
|
|
| 7 |
A |
3082 |
2950 |
59.02 |
|
|
|
| 8 |
A |
3066 |
2935 |
33.48 |
|
|
|
| 9 |
A |
3056 |
2924 |
12.09 |
|
|
|
| 10 |
A |
3055 |
2924 |
18.92 |
|
|
|
| 11 |
A |
1515 |
1450 |
4.61 |
|
|
|
| 12 |
A |
1505 |
1440 |
1.44 |
|
|
|
| 13 |
A |
1490 |
1426 |
5.34 |
|
|
|
| 14 |
A |
1484 |
1421 |
4.34 |
|
|
|
| 15 |
A |
1476 |
1412 |
4.68 |
|
|
|
| 16 |
A |
1405 |
1345 |
12.08 |
|
|
|
| 17 |
A |
1371 |
1312 |
3.74 |
|
|
|
| 18 |
A |
1342 |
1284 |
2.18 |
|
|
|
| 19 |
A |
1320 |
1264 |
2.01 |
|
|
|
| 20 |
A |
1297 |
1241 |
3.94 |
|
|
|
| 21 |
A |
1254 |
1201 |
8.61 |
|
|
|
| 22 |
A |
1232 |
1179 |
4.67 |
|
|
|
| 23 |
A |
1186 |
1135 |
1.60 |
|
|
|
| 24 |
A |
1136 |
1087 |
0.25 |
|
|
|
| 25 |
A |
1078 |
1032 |
0.40 |
|
|
|
| 26 |
A |
1067 |
1021 |
0.74 |
|
|
|
| 27 |
A |
1040 |
996 |
0.44 |
|
|
|
| 28 |
A |
1005 |
962 |
1.52 |
|
|
|
| 29 |
A |
967 |
925 |
0.31 |
|
|
|
| 30 |
A |
942 |
902 |
0.45 |
|
|
|
| 31 |
A |
905 |
867 |
2.36 |
|
|
|
| 32 |
A |
862 |
825 |
0.81 |
|
|
|
| 33 |
A |
818 |
783 |
1.44 |
|
|
|
| 34 |
A |
732 |
700 |
4.40 |
|
|
|
| 35 |
A |
706 |
676 |
1.84 |
|
|
|
| 36 |
A |
612 |
586 |
1.23 |
|
|
|
| 37 |
A |
496 |
475 |
0.07 |
|
|
|
| 38 |
A |
367 |
351 |
0.13 |
|
|
|
| 39 |
A |
342 |
328 |
0.10 |
|
|
|
| 40 |
A |
255 |
244 |
0.13 |
|
|
|
| 41 |
A |
190 |
181 |
0.18 |
|
|
|
| 42 |
A |
122 |
117 |
2.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31250.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29909.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.130 |
|
|
|
| 2 |
C |
-0.377 |
|
|
|
| 3 |
H |
0.133 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
| 5 |
H |
0.121 |
|
|
|
| 6 |
C |
-0.207 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
C |
-0.093 |
|
|
|
| 10 |
H |
0.119 |
|
|
|
| 11 |
C |
-0.231 |
|
|
|
| 12 |
H |
0.131 |
|
|
|
| 13 |
H |
0.137 |
|
|
|
| 14 |
C |
-0.231 |
|
|
|
| 15 |
H |
0.130 |
|
|
|
| 16 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.782 |
0.760 |
0.721 |
2.067 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.545 |
-1.812 |
-0.108 |
| y |
-1.812 |
-43.360 |
-1.162 |
| z |
-0.108 |
-1.162 |
-45.776 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.977 |
-1.812 |
-0.108 |
| y |
-1.812 |
3.801 |
-1.162 |
| z |
-0.108 |
-1.162 |
0.176 |
|
| Polar |
|---|
| 3z2-r2 | 0.352 |
|---|
| x2-y2 | -5.185 |
|---|
| xy | -1.812 |
|---|
| xz | -0.108 |
|---|
| yz | -1.162 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.714 |
0.069 |
0.344 |
| y |
0.069 |
11.301 |
-0.232 |
| z |
0.344 |
-0.232 |
9.697 |
<r2> (average value of r
2) Å
2
| <r2> |
190.093 |
| (<r2>)1/2 |
13.787 |