return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSCH3 (Thioacetone)

using model chemistry: MP4=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4=FULL/cc-pVTZ
 hartrees
Energy at 0K-515.516368
Energy at 298.15K-515.522038
HF Energy-514.670278
Nuclear repulsion energy152.939740
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3158 3041        
2 A1 3045 2932        
3 A1 1502 1446        
4 A1 1400 1348        
5 A1 1310 1261        
6 A1 1021 984        
7 A1 707 680        
8 A1 370 357        
9 A2 3086 2971        
10 A2 1482 1427        
11 A2 919 885        
12 A2 66 63        
13 B1 3093 2978        
14 B1 1510 1454        
15 B1 1072 1032        
16 B1 434 417        
17 B1 90 87        
18 B2 3157 3040        
19 B2 3037 2924        
20 B2 1479 1424        
21 B2 1391 1339        
22 B2 1227 1181        
23 B2 933 899        
24 B2 375 361        

Unscaled Zero Point Vibrational Energy (zpe) 17930.4 cm-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 17265.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVTZ
ABC
0.29457 0.16348 0.10935

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.256
S2 0.000 0.000 1.377
C3 0.000 1.263 -1.065
C4 0.000 -1.263 -1.065
H5 0.000 2.141 -0.429
H6 0.000 -2.141 -0.429
H7 0.876 1.282 -1.715
H8 -0.876 1.282 -1.715
H9 -0.876 -1.282 -1.715
H10 0.876 -1.282 -1.715

Atom - Atom Distances (Å)
  C1 S2 C3 C4 H5 H6 H7 H8 H9 H10
C11.63261.49951.49952.14822.14822.13072.13072.13072.1307
S21.63262.74862.74862.80112.80113.45953.45953.45953.4595
C31.49952.74862.52541.08413.46261.09111.09112.76892.7689
C41.49952.74862.52543.46261.08412.76892.76891.09111.0911
H52.14822.80111.08413.46264.28231.77691.77693.76023.7602
H62.14822.80113.46261.08414.28233.76023.76021.77691.7769
H72.13073.45951.09112.76891.77693.76021.75253.10612.5645
H82.13073.45951.09112.76891.77693.76021.75252.56453.1061
H92.13073.45952.76891.09113.76021.77693.10612.56451.7525
H102.13073.45952.76891.09113.76021.77692.56453.10611.7525

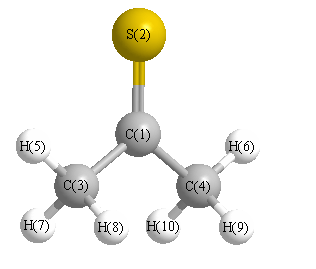
picture of Thioacetone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 111.484 C1 C3 H7 109.658
C1 C3 H8 109.658 C1 C4 H6 111.484
C1 C4 H9 109.658 C1 C4 H10 109.658
S2 C1 C3 122.641 S2 C1 C4 122.641
C3 C1 C4 114.718 H5 C3 H7 109.543
H5 C3 H8 109.543 H6 C4 H9 109.543
H6 C4 H10 109.543 H7 C3 H8 106.843
H9 C4 H10 106.843
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability