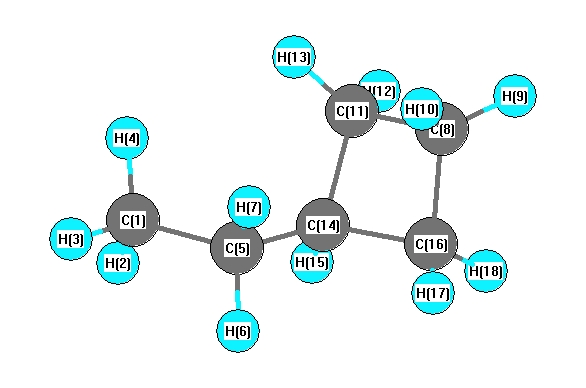
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3142 |
3007 |
73.83 |
|
|
|
| 2 |
A |
3126 |
2992 |
36.10 |
|
|
|
| 3 |
A |
3123 |
2989 |
17.21 |
|
|
|
| 4 |
A |
3120 |
2986 |
38.02 |
|
|
|
| 5 |
A |
3115 |
2981 |
14.18 |
|
|
|
| 6 |
A |
3088 |
2955 |
18.30 |
|
|
|
| 7 |
A |
3078 |
2946 |
63.25 |
|
|
|
| 8 |
A |
3071 |
2939 |
64.94 |
|
|
|
| 9 |
A |
3069 |
2937 |
29.84 |
|
|
|
| 10 |
A |
3057 |
2926 |
18.75 |
|
|
|
| 11 |
A |
3051 |
2920 |
23.87 |
|
|
|
| 12 |
A |
3033 |
2903 |
20.61 |
|
|
|
| 13 |
A |
1508 |
1444 |
4.19 |
|
|
|
| 14 |
A |
1504 |
1440 |
4.82 |
|
|
|
| 15 |
A |
1499 |
1435 |
5.98 |
|
|
|
| 16 |
A |
1485 |
1421 |
1.14 |
|
|
|
| 17 |
A |
1481 |
1417 |
1.82 |
|
|
|
| 18 |
A |
1470 |
1407 |
2.88 |
|
|
|
| 19 |
A |
1410 |
1349 |
4.74 |
|
|
|
| 20 |
A |
1397 |
1337 |
0.12 |
|
|
|
| 21 |
A |
1340 |
1283 |
4.23 |
|
|
|
| 22 |
A |
1309 |
1253 |
0.49 |
|
|
|
| 23 |
A |
1287 |
1232 |
2.46 |
|
|
|
| 24 |
A |
1281 |
1226 |
0.85 |
|
|
|
| 25 |
A |
1263 |
1208 |
0.06 |
|
|
|
| 26 |
A |
1257 |
1203 |
0.01 |
|
|
|
| 27 |
A |
1245 |
1191 |
0.12 |
|
|
|
| 28 |
A |
1190 |
1139 |
0.71 |
|
|
|
| 29 |
A |
1189 |
1138 |
0.05 |
|
|
|
| 30 |
A |
1137 |
1088 |
0.75 |
|
|
|
| 31 |
A |
1094 |
1047 |
0.09 |
|
|
|
| 32 |
A |
1038 |
994 |
0.06 |
|
|
|
| 33 |
A |
1010 |
966 |
0.35 |
|
|
|
| 34 |
A |
999 |
956 |
2.84 |
|
|
|
| 35 |
A |
970 |
928 |
0.44 |
|
|
|
| 36 |
A |
947 |
906 |
1.47 |
|
|
|
| 37 |
A |
932 |
892 |
1.02 |
|
|
|
| 38 |
A |
869 |
832 |
0.78 |
|
|
|
| 39 |
A |
822 |
787 |
0.41 |
|
|
|
| 40 |
A |
770 |
737 |
0.54 |
|
|
|
| 41 |
A |
757 |
725 |
1.44 |
|
|
|
| 42 |
A |
660 |
632 |
3.35 |
|
|
|
| 43 |
A |
414 |
396 |
0.08 |
|
|
|
| 44 |
A |
353 |
338 |
0.08 |
|
|
|
| 45 |
A |
240 |
230 |
0.03 |
|
|
|
| 46 |
A |
208 |
199 |
0.02 |
|
|
|
| 47 |
A |
150 |
144 |
0.01 |
|
|
|
| 48 |
A |
121 |
116 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36839.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 35259.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.364 |
|
|
|
| 2 |
H |
0.112 |
|
|
|
| 3 |
H |
0.119 |
|
|
|
| 4 |
H |
0.115 |
|
|
|
| 5 |
C |
-0.241 |
|
|
|
| 6 |
H |
0.116 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
C |
-0.236 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
H |
0.116 |
|
|
|
| 11 |
C |
-0.233 |
|
|
|
| 12 |
H |
0.112 |
|
|
|
| 13 |
H |
0.112 |
|
|
|
| 14 |
C |
-0.047 |
|
|
|
| 15 |
H |
0.104 |
|
|
|
| 16 |
C |
-0.232 |
|
|
|
| 17 |
H |
0.106 |
|
|
|
| 18 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.014 |
-0.047 |
0.044 |
0.066 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.435 |
-0.374 |
-0.580 |
| y |
-0.374 |
-40.460 |
-0.019 |
| z |
-0.580 |
-0.019 |
-39.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.490 |
-0.374 |
-0.580 |
| y |
-0.374 |
-0.528 |
-0.019 |
| z |
-0.580 |
-0.019 |
1.018 |
|
| Polar |
|---|
| 3z2-r2 | 2.036 |
|---|
| x2-y2 | 0.025 |
|---|
| xy | -0.374 |
|---|
| xz | -0.580 |
|---|
| yz | -0.019 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.549 |
0.065 |
0.077 |
| y |
0.065 |
9.941 |
0.036 |
| z |
0.077 |
0.036 |
9.303 |
<r2> (average value of r
2) Å
2
| <r2> |
190.330 |
| (<r2>)1/2 |
13.796 |