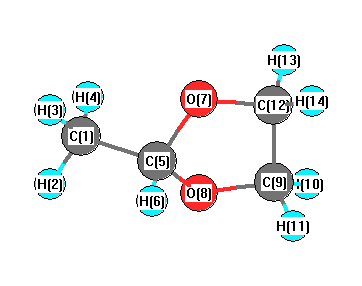
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3029 |
16.34 |
|
|
|
| 2 |
A |
3161 |
3026 |
12.60 |
|
|
|
| 3 |
A |
3144 |
3009 |
27.27 |
|
|
|
| 4 |
A |
3094 |
2962 |
40.82 |
|
|
|
| 5 |
A |
3077 |
2945 |
8.29 |
|
|
|
| 6 |
A |
3054 |
2923 |
64.58 |
|
|
|
| 7 |
A |
3019 |
2889 |
53.04 |
|
|
|
| 8 |
A |
2947 |
2821 |
92.20 |
|
|
|
| 9 |
A |
1533 |
1467 |
0.49 |
|
|
|
| 10 |
A |
1513 |
1448 |
4.62 |
|
|
|
| 11 |
A |
1486 |
1422 |
2.61 |
|
|
|
| 12 |
A |
1484 |
1421 |
7.49 |
|
|
|
| 13 |
A |
1445 |
1383 |
75.56 |
|
|
|
| 14 |
A |
1394 |
1334 |
5.43 |
|
|
|
| 15 |
A |
1389 |
1330 |
4.84 |
|
|
|
| 16 |
A |
1379 |
1320 |
0.35 |
|
|
|
| 17 |
A |
1343 |
1286 |
0.32 |
|
|
|
| 18 |
A |
1263 |
1208 |
23.21 |
|
|
|
| 19 |
A |
1235 |
1182 |
2.06 |
|
|
|
| 20 |
A |
1192 |
1141 |
125.03 |
|
|
|
| 21 |
A |
1168 |
1118 |
32.62 |
|
|
|
| 22 |
A |
1142 |
1093 |
56.62 |
|
|
|
| 23 |
A |
1119 |
1071 |
46.00 |
|
|
|
| 24 |
A |
1095 |
1048 |
22.84 |
|
|
|
| 25 |
A |
1072 |
1026 |
44.63 |
|
|
|
| 26 |
A |
967 |
926 |
8.01 |
|
|
|
| 27 |
A |
893 |
855 |
40.55 |
|
|
|
| 28 |
A |
886 |
848 |
7.70 |
|
|
|
| 29 |
A |
861 |
825 |
7.10 |
|
|
|
| 30 |
A |
705 |
674 |
1.44 |
|
|
|
| 31 |
A |
680 |
650 |
3.40 |
|
|
|
| 32 |
A |
506 |
484 |
4.29 |
|
|
|
| 33 |
A |
326 |
312 |
5.49 |
|
|
|
| 34 |
A |
219 |
210 |
0.55 |
|
|
|
| 35 |
A |
201 |
192 |
3.67 |
|
|
|
| 36 |
A |
61 |
58 |
4.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26607.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25466.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.329 |
|
|
|
| 2 |
H |
0.115 |
|
|
|
| 3 |
H |
0.118 |
|
|
|
| 4 |
H |
0.116 |
|
|
|
| 5 |
C |
0.209 |
|
|
|
| 6 |
H |
0.059 |
|
|
|
| 7 |
O |
-0.288 |
|
|
|
| 8 |
O |
-0.271 |
|
|
|
| 9 |
C |
-0.084 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
| 11 |
H |
0.095 |
|
|
|
| 12 |
C |
-0.022 |
|
|
|
| 13 |
H |
0.091 |
|
|
|
| 14 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.010 |
-0.235 |
0.472 |
1.140 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.324 |
-0.162 |
-0.702 |
| y |
-0.162 |
-40.422 |
-0.601 |
| z |
-0.702 |
-0.601 |
-35.757 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.766 |
-0.162 |
-0.702 |
| y |
-0.162 |
-6.881 |
-0.601 |
| z |
-0.702 |
-0.601 |
0.116 |
|
| Polar |
|---|
| 3z2-r2 | 0.231 |
|---|
| x2-y2 | 9.098 |
|---|
| xy | -0.162 |
|---|
| xz | -0.702 |
|---|
| yz | -0.601 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.349 |
0.028 |
-0.092 |
| y |
0.028 |
7.211 |
-0.131 |
| z |
-0.092 |
-0.131 |
7.104 |
<r2> (average value of r
2) Å
2
| <r2> |
148.230 |
| (<r2>)1/2 |
12.175 |