Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -192.785116 |
| Energy at 298.15K | |
| HF Energy | -191.980016 |
| Nuclear repulsion energy | 128.814216 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3152 |
2992 |
20.11 |
|
|
|
| 2 |
A1 |
3102 |
2945 |
0.46 |
|
|
|
| 3 |
A1 |
1573 |
1493 |
0.00 |
|
|
|
| 4 |
A1 |
1516 |
1440 |
3.98 |
|
|
|
| 5 |
A1 |
1399 |
1328 |
1.36 |
|
|
|
| 6 |
A1 |
1073 |
1019 |
4.02 |
|
|
|
| 7 |
A1 |
952 |
904 |
31.64 |
|
|
|
| 8 |
A1 |
811 |
770 |
5.26 |
|
|
|
| 9 |
A2 |
3137 |
2979 |
0.00 |
|
|
|
| 10 |
A2 |
1259 |
1196 |
0.00 |
|
|
|
| 11 |
A2 |
1192 |
1132 |
0.00 |
|
|
|
| 12 |
A2 |
847 |
804 |
0.00 |
|
|
|
| 13 |
B1 |
3204 |
3042 |
22.18 |
|
|
|
| 14 |
B1 |
3135 |
2977 |
56.32 |
|
|
|
| 15 |
B1 |
1220 |
1158 |
0.06 |
|
|
|
| 16 |
B1 |
1165 |
1106 |
2.46 |
|
|
|
| 17 |
B1 |
777 |
738 |
0.18 |
|
|
|
| 18 |
B1 |
74i |
70i |
3.59 |
|
|
|
| 19 |
B2 |
3095 |
2938 |
130.16 |
|
|
|
| 20 |
B2 |
1545 |
1466 |
1.22 |
|
|
|
| 21 |
B2 |
1337 |
1269 |
0.04 |
|
|
|
| 22 |
B2 |
1272 |
1208 |
9.52 |
|
|
|
| 23 |
B2 |
1067 |
1013 |
87.63 |
|
|
|
| 24 |
B2 |
974 |
925 |
7.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19365.3 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18385.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.070 |
| C2 |
0.000 |
0.000 |
-1.070 |
| C3 |
0.000 |
1.030 |
0.061 |
| C4 |
0.000 |
-1.030 |
0.061 |
| H5 |
0.884 |
0.000 |
-1.692 |
| H6 |
-0.884 |
0.000 |
-1.692 |
| H7 |
0.886 |
1.653 |
0.128 |
| H8 |
-0.886 |
1.653 |
0.128 |
| H9 |
-0.886 |
-1.653 |
0.128 |
| H10 |
0.886 |
-1.653 |
0.128 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1402 | 1.4411 | 1.4411 | 2.9000 | 2.9000 | 2.0993 | 2.0993 | 2.0993 | 2.0993 |
C2 | 2.1402 | | 1.5300 | 1.5300 | 1.0810 | 1.0810 | 2.2258 | 2.2258 | 2.2258 | 2.2258 | C3 | 1.4411 | 1.5300 | | 2.0591 | 2.2174 | 2.2174 | 1.0858 | 1.0858 | 2.8263 | 2.8263 | C4 | 1.4411 | 1.5300 | 2.0591 | | 2.2174 | 2.2174 | 2.8263 | 2.8263 | 1.0858 | 1.0858 | H5 | 2.9000 | 1.0810 | 2.2174 | 2.2174 | | 1.7689 | 2.4585 | 3.0298 | 3.0298 | 2.4585 | H6 | 2.9000 | 1.0810 | 2.2174 | 2.2174 | 1.7689 | | 3.0298 | 2.4585 | 2.4585 | 3.0298 | H7 | 2.0993 | 2.2258 | 1.0858 | 2.8263 | 2.4585 | 3.0298 | | 1.7723 | 3.7519 | 3.3068 | H8 | 2.0993 | 2.2258 | 1.0858 | 2.8263 | 3.0298 | 2.4585 | 1.7723 | | 3.3068 | 3.7519 | H9 | 2.0993 | 2.2258 | 2.8263 | 1.0858 | 3.0298 | 2.4585 | 3.7519 | 3.3068 | | 1.7723 | H10 | 2.0993 | 2.2258 | 2.8263 | 1.0858 | 2.4585 | 3.0298 | 3.3068 | 3.7519 | 1.7723 | |
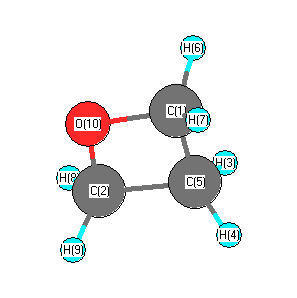 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.114 |
|
O1 |
C3 |
H7 |
111.587 |
| O1 |
C3 |
H8 |
111.587 |
|
O1 |
C4 |
C2 |
92.114 |
| O1 |
C4 |
H9 |
111.587 |
|
O1 |
C4 |
H10 |
111.587 |
| C2 |
C3 |
H7 |
115.572 |
|
C2 |
C3 |
H8 |
115.572 |
| C2 |
C4 |
H9 |
115.572 |
|
C2 |
C4 |
H10 |
115.572 |
| C3 |
O1 |
C4 |
91.189 |
|
C3 |
C2 |
C4 |
84.582 |
| C3 |
C2 |
H5 |
115.172 |
|
C3 |
C2 |
H6 |
115.172 |
| C4 |
C2 |
H5 |
115.172 |
|
C4 |
C2 |
H6 |
115.172 |
| H5 |
C2 |
H6 |
109.801 |
|
H7 |
C3 |
H8 |
109.409 |
| H9 |
C4 |
H10 |
109.409 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -192.785222 |
| Energy at 298.15K | -192.792558 |
| HF Energy | -191.979596 |
| Nuclear repulsion energy | 129.000231 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3203 |
3041 |
23.43 |
|
|
|
| 2 |
A' |
3152 |
2992 |
26.43 |
|
|
|
| 3 |
A' |
3143 |
2984 |
25.91 |
|
|
|
| 4 |
A' |
3090 |
2934 |
29.87 |
|
|
|
| 5 |
A' |
1571 |
1491 |
0.14 |
|
|
|
| 6 |
A' |
1513 |
1437 |
3.81 |
|
|
|
| 7 |
A' |
1395 |
1325 |
1.15 |
|
|
|
| 8 |
A' |
1227 |
1165 |
0.05 |
|
|
|
| 9 |
A' |
1171 |
1112 |
3.38 |
|
|
|
| 10 |
A' |
1076 |
1021 |
5.07 |
|
|
|
| 11 |
A' |
949 |
901 |
27.80 |
|
|
|
| 12 |
A' |
861 |
817 |
9.20 |
|
|
|
| 13 |
A' |
722 |
686 |
1.03 |
|
|
|
| 14 |
A' |
106 |
100 |
3.80 |
|
|
|
| 15 |
A" |
3147 |
2987 |
24.67 |
|
|
|
| 16 |
A" |
3085 |
2929 |
98.14 |
|
|
|
| 17 |
A" |
1542 |
1464 |
1.03 |
|
|
|
| 18 |
A" |
1334 |
1267 |
0.00 |
|
|
|
| 19 |
A" |
1282 |
1217 |
5.81 |
|
|
|
| 20 |
A" |
1242 |
1180 |
4.32 |
|
|
|
| 21 |
A" |
1185 |
1125 |
0.57 |
|
|
|
| 22 |
A" |
1056 |
1003 |
83.01 |
|
|
|
| 23 |
A" |
975 |
925 |
4.23 |
|
|
|
| 24 |
A" |
845 |
803 |
1.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19435.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18452.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.210 |
-1.047 |
0.000 |
| C2 |
-0.057 |
1.067 |
0.000 |
| C3 |
-0.057 |
-0.068 |
1.024 |
| C4 |
-0.057 |
-0.068 |
-1.024 |
| H5 |
0.860 |
1.641 |
0.000 |
| H6 |
-0.910 |
1.732 |
0.000 |
| H7 |
0.719 |
-0.053 |
1.781 |
| H8 |
-1.023 |
-0.240 |
1.494 |
| H9 |
0.719 |
-0.053 |
-1.781 |
| H10 |
-1.023 |
-0.240 |
-1.494 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1312 | 1.4415 | 1.4415 | 2.7651 | 2.9959 | 2.1020 | 2.0980 | 2.1020 | 2.0980 |
C2 | 2.1312 | | 1.5286 | 1.5286 | 1.0816 | 1.0812 | 2.2428 | 2.2074 | 2.2428 | 2.2074 | C3 | 1.4415 | 1.5286 | | 2.0473 | 2.1927 | 2.2391 | 1.0842 | 1.0877 | 2.9101 | 2.7016 | C4 | 1.4415 | 1.5286 | 2.0473 | | 2.1927 | 2.2391 | 2.9101 | 2.7016 | 1.0842 | 1.0877 | H5 | 2.7651 | 1.0816 | 2.1927 | 2.1927 | | 1.7726 | 2.4622 | 3.0517 | 2.4622 | 3.0517 | H6 | 2.9959 | 1.0812 | 2.2391 | 2.2391 | 1.7726 | | 3.0018 | 2.4760 | 3.0018 | 2.4760 | H7 | 2.1020 | 2.2428 | 1.0842 | 2.9101 | 2.4622 | 3.0018 | | 1.7746 | 3.5625 | 3.7136 | H8 | 2.0980 | 2.2074 | 1.0877 | 2.7016 | 3.0517 | 2.4760 | 1.7746 | | 3.7136 | 2.9871 | H9 | 2.1020 | 2.2428 | 2.9101 | 1.0842 | 2.4622 | 3.0018 | 3.5625 | 3.7136 | | 1.7746 | H10 | 2.0980 | 2.2074 | 2.7016 | 1.0877 | 3.0517 | 2.4760 | 3.7136 | 2.9871 | 1.7746 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.656 |
|
O1 |
C3 |
H7 |
111.886 |
| O1 |
C3 |
H8 |
111.328 |
|
O1 |
C4 |
C2 |
91.656 |
| O1 |
C4 |
H9 |
111.886 |
|
O1 |
C4 |
H10 |
111.328 |
| C2 |
C3 |
H7 |
117.264 |
|
C2 |
C3 |
H8 |
114.006 |
| C2 |
C4 |
H9 |
117.264 |
|
C2 |
C4 |
H10 |
114.006 |
| C3 |
O1 |
C4 |
90.489 |
|
C3 |
C2 |
C4 |
84.086 |
| C3 |
C2 |
H5 |
113.178 |
|
C3 |
C2 |
H6 |
117.148 |
| C4 |
C2 |
H5 |
113.178 |
|
C4 |
C2 |
H6 |
117.148 |
| H5 |
C2 |
H6 |
110.086 |
|
H7 |
C3 |
H8 |
109.590 |
| H9 |
C4 |
H10 |
109.590 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
