Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -193.083404 |
| Energy at 298.15K | |
| HF Energy | -193.083404 |
| Nuclear repulsion energy | 129.028827 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3110 |
2977 |
27.15 |
|
|
|
| 2 |
A1 |
3047 |
2917 |
0.89 |
|
|
|
| 3 |
A1 |
1547 |
1481 |
0.02 |
|
|
|
| 4 |
A1 |
1491 |
1427 |
3.45 |
|
|
|
| 5 |
A1 |
1387 |
1327 |
2.17 |
|
|
|
| 6 |
A1 |
1078 |
1031 |
8.62 |
|
|
|
| 7 |
A1 |
961 |
920 |
29.51 |
|
|
|
| 8 |
A1 |
811 |
776 |
5.10 |
|
|
|
| 9 |
A2 |
3083 |
2951 |
0.00 |
|
|
|
| 10 |
A2 |
1238 |
1185 |
0.00 |
|
|
|
| 11 |
A2 |
1172 |
1122 |
0.00 |
|
|
|
| 12 |
A2 |
836 |
800 |
0.00 |
|
|
|
| 13 |
B1 |
3165 |
3029 |
29.47 |
|
|
|
| 14 |
B1 |
3081 |
2949 |
76.31 |
|
|
|
| 15 |
B1 |
1203 |
1151 |
0.06 |
|
|
|
| 16 |
B1 |
1153 |
1104 |
2.49 |
|
|
|
| 17 |
B1 |
766 |
733 |
0.31 |
|
|
|
| 18 |
B1 |
25i |
24i |
3.21 |
|
|
|
| 19 |
B2 |
3038 |
2907 |
159.87 |
|
|
|
| 20 |
B2 |
1517 |
1452 |
1.39 |
|
|
|
| 21 |
B2 |
1319 |
1262 |
0.01 |
|
|
|
| 22 |
B2 |
1265 |
1211 |
9.36 |
|
|
|
| 23 |
B2 |
1079 |
1033 |
103.28 |
|
|
|
| 24 |
B2 |
966 |
925 |
4.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19144.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18323.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.062 |
| C2 |
0.000 |
0.000 |
-1.068 |
| C3 |
0.000 |
1.027 |
0.064 |
| C4 |
0.000 |
-1.027 |
0.064 |
| H5 |
0.885 |
0.000 |
-1.696 |
| H6 |
-0.885 |
0.000 |
-1.696 |
| H7 |
0.888 |
1.658 |
0.132 |
| H8 |
-0.888 |
1.658 |
0.132 |
| H9 |
-0.888 |
-1.658 |
0.132 |
| H10 |
0.888 |
-1.658 |
0.132 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1301 | 1.4322 | 1.4322 | 2.8963 | 2.8963 | 2.0978 | 2.0978 | 2.0978 | 2.0978 |
C2 | 2.1301 | | 1.5290 | 1.5290 | 1.0849 | 1.0849 | 2.2306 | 2.2306 | 2.2306 | 2.2306 | C3 | 1.4322 | 1.5290 | | 2.0548 | 2.2218 | 2.2218 | 1.0906 | 1.0906 | 2.8288 | 2.8288 | C4 | 1.4322 | 1.5290 | 2.0548 | | 2.2218 | 2.2218 | 2.8288 | 2.8288 | 1.0906 | 1.0906 | H5 | 2.8963 | 1.0849 | 2.2218 | 2.2218 | | 1.7696 | 2.4675 | 3.0380 | 3.0380 | 2.4675 | H6 | 2.8963 | 1.0849 | 2.2218 | 2.2218 | 1.7696 | | 3.0380 | 2.4675 | 2.4675 | 3.0380 | H7 | 2.0978 | 2.2306 | 1.0906 | 2.8288 | 2.4675 | 3.0380 | | 1.7751 | 3.7606 | 3.3153 | H8 | 2.0978 | 2.2306 | 1.0906 | 2.8288 | 3.0380 | 2.4675 | 1.7751 | | 3.3153 | 3.7606 | H9 | 2.0978 | 2.2306 | 2.8288 | 1.0906 | 3.0380 | 2.4675 | 3.7606 | 3.3153 | | 1.7751 | H10 | 2.0978 | 2.2306 | 2.8288 | 1.0906 | 2.4675 | 3.0380 | 3.3153 | 3.7606 | 1.7751 | |
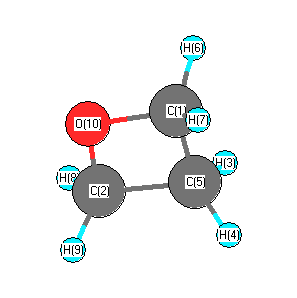 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.944 |
|
O1 |
C3 |
H7 |
111.795 |
| O1 |
C3 |
H8 |
111.795 |
|
O1 |
C4 |
C2 |
91.944 |
| O1 |
C4 |
H9 |
111.795 |
|
O1 |
C4 |
H10 |
111.795 |
| C2 |
C3 |
H7 |
115.738 |
|
C2 |
C3 |
H8 |
115.738 |
| C2 |
C4 |
H9 |
115.738 |
|
C2 |
C4 |
H10 |
115.738 |
| C3 |
O1 |
C4 |
91.674 |
|
C3 |
C2 |
C4 |
84.439 |
| C3 |
C2 |
H5 |
115.373 |
|
C3 |
C2 |
H6 |
115.373 |
| C4 |
C2 |
H5 |
115.373 |
|
C4 |
C2 |
H6 |
115.373 |
| H5 |
C2 |
H6 |
109.292 |
|
H7 |
C3 |
H8 |
108.934 |
| H9 |
C4 |
H10 |
108.934 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.294 |
|
|
|
| 2 |
C |
-0.286 |
|
|
|
| 3 |
C |
0.013 |
|
|
|
| 4 |
C |
0.013 |
|
|
|
| 5 |
H |
0.118 |
|
|
|
| 6 |
H |
0.118 |
|
|
|
| 7 |
H |
0.079 |
|
|
|
| 8 |
H |
0.079 |
|
|
|
| 9 |
H |
0.079 |
|
|
|
| 10 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.864 |
1.864 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.044 |
0.000 |
0.000 |
| y |
0.000 |
-22.981 |
0.000 |
| z |
0.000 |
0.000 |
-27.609 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.252 |
0.000 |
0.000 |
| y |
0.000 |
2.845 |
0.000 |
| z |
0.000 |
0.000 |
-4.096 |
|
| Polar |
|---|
| 3z2-r2 | -8.193 |
|---|
| x2-y2 | -1.062 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.082 |
0.000 |
0.000 |
| y |
0.000 |
6.214 |
0.000 |
| z |
0.000 |
0.000 |
5.254 |
<r2> (average value of r
2) Å
2
| <r2> |
65.652 |
| (<r2>)1/2 |
8.103 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -193.083405 |
| Energy at 298.15K | -193.090590 |
| HF Energy | -193.083405 |
| Nuclear repulsion energy | 129.047342 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
3029 |
29.65 |
|
|
|
| 2 |
A' |
3110 |
2977 |
27.50 |
|
|
|
| 3 |
A' |
3083 |
2951 |
70.33 |
|
|
|
| 4 |
A' |
3046 |
2915 |
7.04 |
|
|
|
| 5 |
A' |
1547 |
1480 |
0.03 |
|
|
|
| 6 |
A' |
1491 |
1427 |
3.44 |
|
|
|
| 7 |
A' |
1387 |
1327 |
2.15 |
|
|
|
| 8 |
A' |
1204 |
1152 |
0.06 |
|
|
|
| 9 |
A' |
1154 |
1104 |
2.62 |
|
|
|
| 10 |
A' |
1078 |
1032 |
8.73 |
|
|
|
| 11 |
A' |
961 |
920 |
29.31 |
|
|
|
| 12 |
A' |
819 |
784 |
5.06 |
|
|
|
| 13 |
A' |
757 |
725 |
0.67 |
|
|
|
| 14 |
A' |
29 |
27 |
3.24 |
|
|
|
| 15 |
A" |
3084 |
2952 |
4.98 |
|
|
|
| 16 |
A" |
3036 |
2906 |
154.02 |
|
|
|
| 17 |
A" |
1517 |
1452 |
1.37 |
|
|
|
| 18 |
A" |
1319 |
1262 |
0.02 |
|
|
|
| 19 |
A" |
1266 |
1212 |
8.87 |
|
|
|
| 20 |
A" |
1237 |
1183 |
0.57 |
|
|
|
| 21 |
A" |
1172 |
1121 |
0.06 |
|
|
|
| 22 |
A" |
1078 |
1032 |
102.65 |
|
|
|
| 23 |
A" |
966 |
925 |
4.08 |
|
|
|
| 24 |
A" |
836 |
800 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19169.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18346.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.065 |
-1.060 |
0.000 |
| C2 |
-0.018 |
1.068 |
0.000 |
| C3 |
-0.018 |
-0.065 |
1.027 |
| C4 |
-0.018 |
-0.065 |
-1.027 |
| H5 |
0.877 |
1.681 |
0.000 |
| H6 |
-0.892 |
1.709 |
0.000 |
| H7 |
0.838 |
-0.104 |
1.701 |
| H8 |
-0.934 |
-0.163 |
1.612 |
| H9 |
0.838 |
-0.104 |
-1.701 |
| H10 |
-0.934 |
-0.163 |
-1.612 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1293 | 1.4322 | 1.4322 | 2.8590 | 2.9300 | 2.0984 | 2.0973 | 2.0984 | 2.0973 |
C2 | 2.1293 | | 1.5288 | 1.5288 | 1.0850 | 1.0848 | 2.2357 | 2.2252 | 2.2357 | 2.2252 | C3 | 1.4322 | 1.5288 | | 2.0536 | 2.2146 | 2.2287 | 1.0901 | 1.0912 | 2.8589 | 2.7946 | C4 | 1.4322 | 1.5288 | 2.0536 | | 2.2146 | 2.2287 | 2.8589 | 2.7946 | 1.0901 | 1.0912 | H5 | 2.8590 | 1.0850 | 2.2146 | 2.2146 | | 1.7699 | 2.4662 | 3.0461 | 2.4662 | 3.0461 | H6 | 2.9300 | 1.0848 | 2.2287 | 2.2287 | 1.7699 | | 3.0293 | 2.4708 | 3.0293 | 2.4708 | H7 | 2.0984 | 2.2357 | 1.0901 | 2.8589 | 2.4662 | 3.0293 | | 1.7753 | 3.4013 | 3.7569 | H8 | 2.0973 | 2.2252 | 1.0912 | 2.7946 | 3.0461 | 2.4708 | 1.7753 | | 3.7569 | 3.2230 | H9 | 2.0984 | 2.2357 | 2.8589 | 1.0901 | 2.4662 | 3.0293 | 3.4013 | 3.7569 | | 1.7753 | H10 | 2.0973 | 2.2252 | 2.7946 | 1.0912 | 3.0461 | 2.4708 | 3.7569 | 3.2230 | 1.7753 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.902 |
|
O1 |
C3 |
H7 |
111.879 |
| O1 |
C3 |
H8 |
111.718 |
|
O1 |
C4 |
C2 |
91.902 |
| O1 |
C4 |
H9 |
111.879 |
|
O1 |
C4 |
H10 |
111.718 |
| C2 |
C3 |
H7 |
116.227 |
|
C2 |
C3 |
H8 |
115.258 |
| C2 |
C4 |
H9 |
116.227 |
|
C2 |
C4 |
H10 |
115.258 |
| C3 |
O1 |
C4 |
91.604 |
|
C3 |
C2 |
C4 |
84.389 |
| C3 |
C2 |
H5 |
114.771 |
|
C3 |
C2 |
H6 |
115.980 |
| C4 |
C2 |
H5 |
114.771 |
|
C4 |
C2 |
H6 |
115.980 |
| H5 |
C2 |
H6 |
109.313 |
|
H7 |
C3 |
H8 |
108.951 |
| H9 |
C4 |
H10 |
108.951 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.294 |
|
|
|
| 2 |
C |
-0.285 |
|
|
|
| 3 |
C |
0.013 |
|
|
|
| 4 |
C |
0.013 |
|
|
|
| 5 |
H |
0.118 |
|
|
|
| 6 |
H |
0.118 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.078 |
|
|
|
| 9 |
H |
0.081 |
|
|
|
| 10 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.125 |
1.858 |
0.000 |
1.862 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.064 |
0.210 |
0.000 |
| y |
0.210 |
-27.596 |
0.000 |
| z |
0.000 |
0.000 |
-22.975 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.222 |
0.210 |
0.000 |
| y |
0.210 |
-4.076 |
0.000 |
| z |
0.000 |
0.000 |
2.855 |
|
| Polar |
|---|
| 3z2-r2 | 5.709 |
|---|
| x2-y2 | 3.532 |
|---|
| xy | 0.210 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.086 |
-0.002 |
0.000 |
| y |
-0.002 |
5.256 |
0.000 |
| z |
0.000 |
0.000 |
6.213 |
<r2> (average value of r
2) Å
2
| <r2> |
65.626 |
| (<r2>)1/2 |
8.101 |
