return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O2 (1,3-Propanediol)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-269.159929
Energy at 298.15K-269.170000
Nuclear repulsion energy196.237328
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3883 3686 41.59      
2 A 3805 3613 150.52      
3 A 3143 2984 23.82      
4 A 3138 2979 38.93      
5 A 3107 2949 49.10      
6 A 3093 2936 4.25      
7 A 3064 2909 47.08      
8 A 3029 2876 59.61      
9 A 1553 1474 2.40      
10 A 1538 1461 1.50      
11 A 1484 1409 23.75      
12 A 1473 1399 40.80      
13 A 1466 1392 20.08      
14 A 1413 1341 1.41      
15 A 1394 1324 1.48      
16 A 1329 1262 17.13      
17 A 1300 1234 6.66      
18 A 1250 1187 6.01      
19 A 1215 1153 48.52      
20 A 1155 1097 25.99      
21 A 1127 1070 12.78      
22 A 1115 1058 130.91      
23 A 985 935 2.24      
24 A 932 885 0.32      
25 A 927 880 12.93      
26 A 842 799 9.25      
27 A 550 522 118.62      
28 A 508 483 2.32      
29 A 389 369 4.54      
30 A 338 321 14.85      
31 A 280 266 78.83      
32 A 217 206 3.97      
33 A 102 97 0.96      

Unscaled Zero Point Vibrational Energy (zpe) 25571.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 24277.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.25941 0.13408 0.09768

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.030 1.030 -0.351
C2 1.266 0.475 0.182
C3 -1.250 0.403 0.295
H4 -0.080 0.874 -1.425
H5 -0.035 2.102 -0.168
O6 1.297 -0.926 -0.093
O7 -1.474 -0.925 -0.127
H8 2.084 -1.297 0.311
H9 -0.614 -1.358 -0.088
H10 2.112 0.970 -0.291
H11 1.321 0.650 1.256
H12 -1.147 0.461 1.382
H13 -2.138 0.964 0.024

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 O6 O7 H8 H9 H10 H11 H12 H13
C11.50751.51561.08681.08802.37772.44063.21312.47242.14422.13342.13842.1421
C21.50752.52032.13372.11321.42783.09301.95602.64001.08801.08992.69563.4435
C31.51562.52032.13302.13942.89941.41193.74321.91183.45992.75651.09351.0847
H41.08682.13372.13301.75782.62912.62053.52312.65682.46993.03323.03132.5188
H51.08802.11322.13941.75783.30943.35244.03483.50972.43082.44482.51682.3993
O62.37771.42782.89942.62913.30942.77150.95931.95952.07342.07443.17343.9228
O72.44063.09301.41192.62053.35242.77153.60450.96374.05973.49422.07472.0081
H83.21311.95603.74323.52314.03480.95933.60452.72842.34612.29493.83114.7986
H92.47242.64001.91182.65683.50971.95950.96372.72843.59113.09612.39832.7800
H102.14421.08803.45992.46992.43082.07344.05972.34613.59111.76633.69884.2624
H112.13341.08992.75653.03322.44482.07443.49422.29493.09611.76632.47903.6862
H122.13842.69561.09353.03132.51683.17342.07473.83112.39833.69882.47901.7554
H132.14213.44351.08472.51882.39933.92282.00814.79862.78004.26243.68621.7554

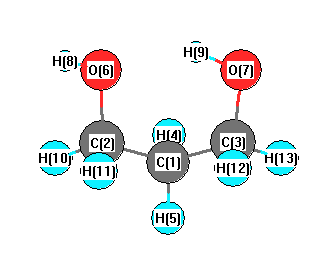
picture of 1,3-Propanediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O6 108.173 C1 C2 H10 110.360
C1 C2 H11 109.397 C1 C3 O7 112.908
C1 C3 H12 109.012 C1 C3 H13 109.826
C2 C1 C3 112.956 C2 C1 H4 109.600
C2 C1 H5 107.917 C2 O6 H8 108.452
C3 C1 H4 108.978 C3 C1 H5 109.410
C3 O7 H9 105.616 H4 C1 H5 107.853
O6 C2 H10 110.277 O6 C2 H11 110.239
O7 C3 H12 111.173 O7 C3 H13 106.353
H10 C2 H11 108.393 H12 C3 H13 107.398
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability