return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-307.187893
Energy at 298.15K-307.199314
HF Energy-305.946646
Nuclear repulsion energy266.980680
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3195 3033 23.90      
2 A' 3167 3007 18.69      
3 A' 3154 2995 18.85      
4 A' 3106 2949 15.25      
5 A' 3029 2876 88.11      
6 A' 2986 2835 69.92      
7 A' 1540 1462 5.95      
8 A' 1526 1448 0.46      
9 A' 1490 1414 5.76      
10 A' 1427 1354 13.24      
11 A' 1343 1275 5.46      
12 A' 1230 1168 85.77      
13 A' 1206 1145 94.75      
14 A' 1130 1073 13.93      
15 A' 1026 974 34.79      
16 A' 945 898 9.50      
17 A' 867 823 11.30      
18 A' 656 623 4.70      
19 A' 494 469 0.71      
20 A' 455 432 9.25      
21 A' 278 264 2.06      
22 A" 3165 3005 34.49      
23 A" 3024 2871 13.21      
24 A" 1523 1446 4.30      
25 A" 1458 1384 8.95      
26 A" 1405 1333 2.19      
27 A" 1386 1316 0.06      
28 A" 1356 1288 2.11      
29 A" 1280 1215 31.84      
30 A" 1249 1186 0.09      
31 A" 1115 1058 52.46      
32 A" 1058 1004 75.57      
33 A" 937 890 21.98      
34 A" 917 871 0.19      
35 A" 465 441 6.81      
36 A" 276 262 1.85      

Unscaled Zero Point Vibrational Energy (zpe) 27430.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 26042.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.16983 0.16265 0.09362

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.614 -1.190 0.000
O2 0.014 -0.758 1.166
O3 0.014 -0.758 -1.166
C4 0.014 0.652 1.227
C5 0.014 0.652 -1.227
C6 0.686 1.233 0.000
H7 -0.571 -2.270 0.000
H8 -1.660 -0.849 0.000
H9 0.538 0.923 2.135
H10 -1.013 1.025 1.296
H11 0.538 0.923 -2.135
H12 -1.013 1.025 -1.296
H13 1.735 0.955 0.000
H14 0.611 2.317 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.39311.39312.30112.30112.74981.08111.09963.21732.59743.21732.59743.18093.7151
O21.39312.33181.41202.77772.40331.99712.04192.01002.06213.74133.20912.69343.3427
O31.39312.33182.77771.41202.40331.99712.04193.74133.20912.01002.06212.69343.3427
C42.30111.41202.77772.45391.51453.22342.56151.08271.09473.41342.74972.13482.1525
C52.30112.77771.41202.45391.51453.22342.56153.41342.74971.08271.09472.13482.1525
C62.74982.40332.40331.51451.51453.72203.13632.16262.14692.16262.14691.08531.0867
H71.08111.99711.99713.22343.22343.72201.79003.99823.56863.99823.56863.96474.7373
H81.09962.04192.04192.56152.56153.13631.79003.53942.36893.53942.36893.84403.8962
H93.21732.01003.74131.08273.41342.16263.99823.53941.76594.27033.76702.44802.5510
H102.59742.06213.20911.09472.74972.14693.56862.36891.76593.76702.59293.03902.4468
H113.21733.74132.01003.41341.08272.16263.99823.53944.27033.76701.76592.44802.5510
H122.59743.20912.06212.74971.09472.14693.56862.36893.76702.59291.76593.03902.4468
H133.18092.69342.69342.13482.13481.08533.96473.84402.44803.03902.44803.03901.7662
H143.71513.34273.34272.15252.15251.08674.73733.89622.55102.44682.55102.44681.7662

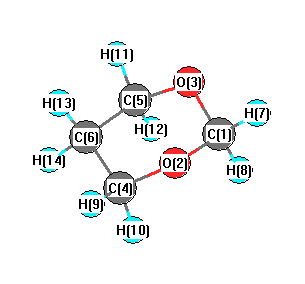
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.230 C1 O3 C5 110.230
O2 C1 O3 113.629 O2 C1 H7 106.964
O2 C1 H8 109.439 O2 C4 C6 110.362
O2 C4 H9 106.617 O2 C4 H10 110.056
O3 C1 H7 106.964 O3 C1 H8 109.439
O3 C5 C6 110.362 O3 C5 H11 106.617
O3 C5 H12 110.056 C4 C6 C5 108.220
C4 C6 H13 109.291 C4 C6 H14 110.603
C5 C6 H13 109.291 C5 C6 H14 110.603
C6 C4 H9 111.668 C6 C4 H10 109.691
C6 C5 H11 111.668 C6 C5 H12 109.691
H7 C1 H8 110.341 H9 C4 H10 108.385
H11 C5 H12 108.385 H13 C6 H14 108.809
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability