Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.064697 |
| Energy at 298.15K | -133.068890 |
| HF Energy | -132.516252 |
| Nuclear repulsion energy | 64.822483 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3455 |
3368 |
|
|
|
|
| 2 |
A' |
3273 |
3191 |
|
|
|
|
| 3 |
A' |
3165 |
3085 |
|
|
|
|
| 4 |
A' |
3084 |
3006 |
|
|
|
|
| 5 |
A' |
1519 |
1480 |
|
|
|
|
| 6 |
A' |
1467 |
1430 |
|
|
|
|
| 7 |
A' |
1364 |
1329 |
|
|
|
|
| 8 |
A' |
1248 |
1217 |
|
|
|
|
| 9 |
A' |
1108 |
1080 |
|
|
|
|
| 10 |
A' |
1001 |
976 |
|
|
|
|
| 11 |
A' |
486 |
473 |
|
|
|
|
| 12 |
A" |
1063 |
1036 |
|
|
|
|
| 13 |
A" |
812 |
792 |
|
|
|
|
| 14 |
A" |
687 |
670 |
|
|
|
|
| 15 |
A" |
536 |
522 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12133.4 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 11827.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| C2 |
1.154 |
-0.380 |
0.000 |
| N3 |
-1.193 |
-0.135 |
0.000 |
| H4 |
0.135 |
1.503 |
0.000 |
| H5 |
2.141 |
0.061 |
0.000 |
| H6 |
1.051 |
-1.457 |
0.000 |
| H7 |
-1.904 |
0.598 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4042 | 1.3159 | 1.0919 | 2.1712 | 2.1509 | 1.9119 |
C2 | 1.4042 | | 2.3600 | 2.1410 | 1.0810 | 1.0820 | 3.2104 | N3 | 1.3159 | 2.3600 | | 2.1095 | 3.3402 | 2.6042 | 1.0211 | H4 | 1.0919 | 2.1410 | 2.1095 | | 2.4706 | 3.0984 | 2.2310 | H5 | 2.1712 | 1.0810 | 3.3402 | 2.4706 | | 1.8689 | 4.0804 | H6 | 2.1509 | 1.0820 | 2.6042 | 3.0984 | 1.8689 | | 3.5989 | H7 | 1.9119 | 3.2104 | 1.0211 | 2.2310 | 4.0804 | 3.5989 | |
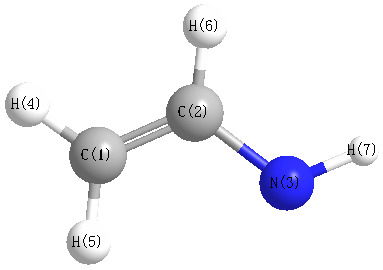 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.227 |
|
C1 |
C2 |
H6 |
119.232 |
| C1 |
N3 |
H7 |
109.139 |
|
C2 |
C1 |
N3 |
120.337 |
| C2 |
C1 |
H4 |
117.582 |
|
N3 |
C1 |
H4 |
122.081 |
| H5 |
C2 |
H6 |
119.541 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.064198 |
| Energy at 298.15K | -133.068340 |
| HF Energy | -132.515457 |
| Nuclear repulsion energy | 64.845955 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3396 |
3311 |
|
|
|
|
| 2 |
A' |
3255 |
3173 |
|
|
|
|
| 3 |
A' |
3144 |
3065 |
|
|
|
|
| 4 |
A' |
3132 |
3054 |
|
|
|
|
| 5 |
A' |
1503 |
1465 |
|
|
|
|
| 6 |
A' |
1452 |
1415 |
|
|
|
|
| 7 |
A' |
1382 |
1347 |
|
|
|
|
| 8 |
A' |
1229 |
1198 |
|
|
|
|
| 9 |
A' |
1129 |
1100 |
|
|
|
|
| 10 |
A' |
982 |
957 |
|
|
|
|
| 11 |
A' |
482 |
470 |
|
|
|
|
| 12 |
A" |
1082 |
1055 |
|
|
|
|
| 13 |
A" |
809 |
789 |
|
|
|
|
| 14 |
A" |
683 |
666 |
|
|
|
|
| 15 |
A" |
475 |
463 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12067.5 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 11763.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.455 |
0.000 |
| C2 |
1.128 |
-0.384 |
0.000 |
| N3 |
-1.257 |
0.063 |
0.000 |
| H4 |
0.166 |
1.530 |
0.000 |
| H5 |
2.130 |
0.024 |
0.000 |
| H6 |
1.009 |
-1.461 |
0.000 |
| H7 |
-1.274 |
-0.962 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4053 | 1.3164 | 1.0881 | 2.1729 | 2.1653 | 1.9052 |
C2 | 1.4053 | | 2.4264 | 2.1419 | 1.0816 | 1.0841 | 2.4707 | N3 | 1.3164 | 2.4264 | | 2.0433 | 3.3870 | 2.7313 | 1.0254 | H4 | 1.0881 | 2.1419 | 2.0433 | | 2.4751 | 3.1078 | 2.8779 | H5 | 2.1729 | 1.0816 | 3.3870 | 2.4751 | | 1.8604 | 3.5438 | H6 | 2.1653 | 1.0841 | 2.7313 | 3.1078 | 1.8604 | | 2.3373 | H7 | 1.9052 | 2.4707 | 1.0254 | 2.8779 | 3.5438 | 2.3373 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.253 |
|
C1 |
C2 |
H6 |
120.326 |
| C1 |
N3 |
H7 |
108.248 |
|
C2 |
C1 |
N3 |
126.098 |
| C2 |
C1 |
H4 |
117.857 |
|
N3 |
C1 |
H4 |
116.044 |
| H5 |
C2 |
H6 |
118.421 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
