Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.047156 |
| Energy at 298.15K | -133.051417 |
| HF Energy | -132.515701 |
| Nuclear repulsion energy | 65.685736 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3542 |
3362 |
1.42 |
|
|
|
| 2 |
A' |
3341 |
3172 |
0.79 |
|
|
|
| 3 |
A' |
3240 |
3076 |
3.86 |
|
|
|
| 4 |
A' |
3129 |
2970 |
37.18 |
|
|
|
| 5 |
A' |
1540 |
1462 |
15.15 |
|
|
|
| 6 |
A' |
1496 |
1420 |
12.03 |
|
|
|
| 7 |
A' |
1410 |
1339 |
18.06 |
|
|
|
| 8 |
A' |
1260 |
1196 |
29.84 |
|
|
|
| 9 |
A' |
1064 |
1010 |
10.02 |
|
|
|
| 10 |
A' |
1022 |
971 |
9.91 |
|
|
|
| 11 |
A' |
499 |
474 |
15.23 |
|
|
|
| 12 |
A" |
1173 |
1114 |
0.18 |
|
|
|
| 13 |
A" |
870 |
826 |
5.08 |
|
|
|
| 14 |
A" |
745 |
707 |
107.69 |
|
|
|
| 15 |
A" |
535 |
508 |
1.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12432.7 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 11803.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.412 |
0.000 |
| C2 |
1.134 |
-0.402 |
0.000 |
| N3 |
-1.175 |
-0.103 |
0.000 |
| H4 |
0.166 |
1.485 |
0.000 |
| H5 |
2.123 |
0.015 |
0.000 |
| H6 |
1.004 |
-1.468 |
0.000 |
| H7 |
-1.876 |
0.629 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3961 | 1.2827 | 1.0855 | 2.1598 | 2.1311 | 1.8888 |
C2 | 1.3961 | | 2.3280 | 2.1205 | 1.0728 | 1.0738 | 3.1822 | N3 | 1.2827 | 2.3280 | | 2.0785 | 3.2996 | 2.5703 | 1.0145 | H4 | 1.0855 | 2.1205 | 2.0785 | | 2.4473 | 3.0690 | 2.2145 | H5 | 2.1598 | 1.0728 | 3.2996 | 2.4473 | | 1.8574 | 4.0461 | H6 | 2.1311 | 1.0738 | 2.5703 | 3.0690 | 1.8574 | | 3.5626 | H7 | 1.8888 | 3.1822 | 1.0145 | 2.2145 | 4.0461 | 3.5626 | |
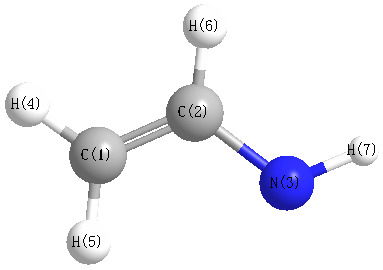 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.485 |
|
C1 |
C2 |
H6 |
118.687 |
| C1 |
N3 |
H7 |
110.065 |
|
C2 |
C1 |
N3 |
120.636 |
| C2 |
C1 |
H4 |
116.850 |
|
N3 |
C1 |
H4 |
122.514 |
| H5 |
C2 |
H6 |
119.827 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.046418 |
| Energy at 298.15K | -133.050675 |
| HF Energy | -132.514989 |
| Nuclear repulsion energy | 65.699611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3487 |
3311 |
2.06 |
|
|
|
| 2 |
A' |
3323 |
3155 |
2.70 |
|
|
|
| 3 |
A' |
3225 |
3062 |
2.84 |
|
|
|
| 4 |
A' |
3182 |
3021 |
16.56 |
|
|
|
| 5 |
A' |
1539 |
1461 |
8.69 |
|
|
|
| 6 |
A' |
1475 |
1400 |
0.63 |
|
|
|
| 7 |
A' |
1418 |
1346 |
19.15 |
|
|
|
| 8 |
A' |
1248 |
1185 |
45.91 |
|
|
|
| 9 |
A' |
1090 |
1035 |
33.03 |
|
|
|
| 10 |
A' |
1009 |
958 |
0.09 |
|
|
|
| 11 |
A' |
504 |
479 |
9.22 |
|
|
|
| 12 |
A" |
1189 |
1129 |
47.30 |
|
|
|
| 13 |
A" |
893 |
848 |
71.68 |
|
|
|
| 14 |
A" |
754 |
716 |
0.65 |
|
|
|
| 15 |
A" |
508 |
483 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12421.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 11793.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.446 |
0.000 |
| C2 |
1.109 |
-0.403 |
0.000 |
| N3 |
-1.234 |
0.091 |
0.000 |
| H4 |
0.195 |
1.510 |
0.000 |
| H5 |
2.109 |
-0.014 |
0.000 |
| H6 |
0.971 |
-1.469 |
0.000 |
| H7 |
-1.290 |
-0.926 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3963 | 1.2841 | 1.0816 | 2.1588 | 2.1474 | 1.8828 |
C2 | 1.3963 | | 2.3945 | 2.1193 | 1.0735 | 1.0755 | 2.4551 | N3 | 1.2841 | 2.3945 | | 2.0138 | 3.3452 | 2.7018 | 1.0185 | H4 | 1.0816 | 2.1193 | 2.0138 | | 2.4462 | 3.0783 | 2.8525 | H5 | 2.1588 | 1.0735 | 3.3452 | 2.4462 | | 1.8478 | 3.5195 | H6 | 2.1474 | 1.0755 | 2.7018 | 3.0783 | 1.8478 | | 2.3256 | H7 | 1.8828 | 2.4551 | 1.0185 | 2.8525 | 3.5195 | 2.3256 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.329 |
|
C1 |
C2 |
H6 |
120.075 |
| C1 |
N3 |
H7 |
109.164 |
|
C2 |
C1 |
N3 |
126.547 |
| C2 |
C1 |
H4 |
117.018 |
|
N3 |
C1 |
H4 |
116.435 |
| H5 |
C2 |
H6 |
118.596 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
