All results from a given calculation for HN=C=C(CN)2 (Dicyanoketenimine)
using model chemistry: CCSD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -316.655260 |
| Energy at 298.15K | |
| HF Energy | -315.425450 |
| Nuclear repulsion energy | 211.505348 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.008 |
-0.056 |
0.000 |
| C2 |
-0.008 |
1.283 |
0.000 |
| N3 |
0.139 |
2.485 |
0.000 |
| C4 |
-0.008 |
-0.764 |
1.247 |
| C5 |
-0.008 |
-0.764 |
-1.247 |
| N6 |
-0.008 |
-1.335 |
2.261 |
| N7 |
-0.008 |
-1.335 |
-2.261 |
| H8 |
-0.671 |
3.100 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
C4 |
C5 |
N6 |
N7 |
H8 |
| C1 | | 1.3397 | 2.5459 | 1.4335 | 1.4335 | 2.5981 | 2.5981 | 3.2252 |
C2 | 1.3397 | | 1.2110 | 2.3968 | 2.3968 | 3.4600 | 3.4600 | 1.9338 | N3 | 2.5459 | 1.2110 | | 3.4831 | 3.4831 | 4.4422 | 4.4422 | 1.0170 | C4 | 1.4335 | 2.3968 | 3.4831 | | 2.4938 | 1.1645 | 3.5546 | 4.1136 | C5 | 1.4335 | 2.3968 | 3.4831 | 2.4938 | | 3.5546 | 1.1645 | 4.1136 | N6 | 2.5981 | 3.4600 | 4.4422 | 1.1645 | 3.5546 | | 4.5229 | 5.0225 | N7 | 2.5981 | 3.4600 | 4.4422 | 3.5546 | 1.1645 | 4.5229 | | 5.0225 | H8 | 3.2252 | 1.9338 | 1.0170 | 4.1136 | 4.1136 | 5.0225 | 5.0225 | |
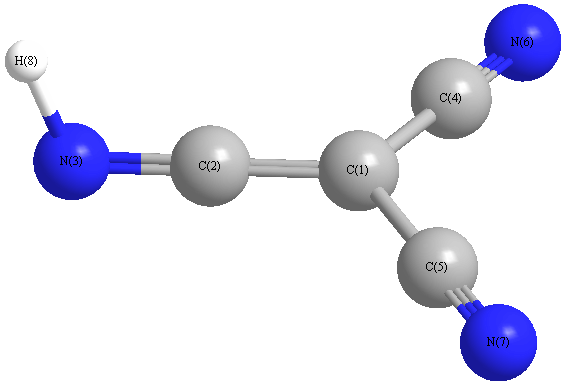 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
173.020 |
|
C1 |
C4 |
N6 |
179.841 |
| C1 |
C5 |
N7 |
179.841 |
|
C2 |
C1 |
C4 |
119.563 |
| C2 |
C1 |
C5 |
119.563 |
|
C2 |
N3 |
H8 |
120.204 |
| C4 |
C1 |
C5 |
120.874 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
