Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.678713 |
| Energy at 298.15K | |
| HF Energy | -151.139170 |
| Nuclear repulsion energy | 52.294468 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3468 |
3289 |
|
|
|
|
| 2 |
Σ |
2087 |
1980 |
|
|
|
|
| 3 |
Σ |
1279 |
1214 |
|
|
|
|
| 4 |
Π |
562 |
533 |
|
|
|
|
| 4 |
Π |
509 |
483 |
|
|
|
|
| 5 |
Π |
381 |
361 |
|
|
|
|
| 5 |
Π |
464i |
440i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3910.6 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 3709.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
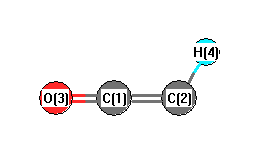
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.019 |
| C2 |
0.000 |
0.000 |
-1.247 |
| O3 |
0.000 |
0.000 |
1.209 |
| H4 |
0.000 |
0.000 |
-2.309 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2661 | 1.1899 | 2.3278 |
C2 | 1.2661 | | 2.4560 | 1.0618 | O3 | 1.1899 | 2.4560 | | 3.5178 | H4 | 2.3278 | 1.0618 | 3.5178 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.682444 |
| Energy at 298.15K | |
| HF Energy | -151.138260 |
| Nuclear repulsion energy | 52.100682 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3324 |
3153 |
|
|
|
|
| 2 |
A' |
2059 |
1953 |
|
|
|
|
| 3 |
A' |
1214 |
1152 |
|
|
|
|
| 4 |
A' |
577 |
547 |
|
|
|
|
| 5 |
A' |
520 |
493 |
|
|
|
|
| 6 |
A" |
493 |
468 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4093.2 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 3882.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.060 |
0.000 |
| C2 |
1.123 |
-0.611 |
0.000 |
| O3 |
-1.111 |
0.449 |
0.000 |
| H4 |
2.147 |
-0.287 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3086 | 1.1767 | 2.1751 |
C2 | 1.3086 | | 2.4726 | 1.0739 | O3 | 1.1767 | 2.4726 | | 3.3400 | H4 | 2.1751 | 1.0739 | 3.3400 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
131.589 |
|
C2 |
C1 |
O3 |
168.404 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
