Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.677300 |
| Energy at 298.15K | |
| HF Energy | -151.139396 |
| Nuclear repulsion energy | 52.346734 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3468 |
3381 |
|
|
|
|
| 2 |
Σ |
2090 |
2037 |
|
|
|
|
| 3 |
Σ |
1290 |
1257 |
|
|
|
|
| 4 |
Π |
560 |
546 |
|
|
|
|
| 4 |
Π |
514 |
501 |
|
|
|
|
| 5 |
Π |
387 |
377 |
|
|
|
|
| 5 |
Π |
428i |
418i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3939.8 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 3840.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.020 |
| C2 |
0.000 |
0.000 |
-1.246 |
| O3 |
0.000 |
0.000 |
1.208 |
| H4 |
0.000 |
0.000 |
-2.307 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2656 | 1.1880 | 2.3273 |
C2 | 1.2656 | | 2.4535 | 1.0617 | O3 | 1.1880 | 2.4535 | | 3.5153 | H4 | 2.3273 | 1.0617 | 3.5153 | |
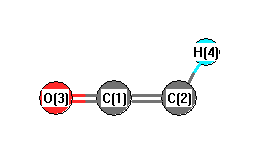 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.681002 |
| Energy at 298.15K | |
| HF Energy | -151.138563 |
| Nuclear repulsion energy | 52.153021 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3324 |
3241 |
|
|
|
|
| 2 |
A' |
2080 |
2027 |
|
|
|
|
| 3 |
A' |
1225 |
1194 |
|
|
|
|
| 4 |
A' |
579 |
565 |
|
|
|
|
| 5 |
A' |
524 |
511 |
|
|
|
|
| 6 |
A" |
493 |
481 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4112.7 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 4009.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.059 |
0.000 |
| C2 |
1.111 |
-0.631 |
0.000 |
| O3 |
-1.101 |
0.469 |
0.000 |
| H4 |
2.140 |
-0.323 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3081 | 1.1747 | 2.1740 |
C2 | 1.3081 | | 2.4705 | 1.0739 | O3 | 1.1747 | 2.4705 | | 3.3364 | H4 | 2.1740 | 1.0739 | 3.3364 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
131.503 |
|
C2 |
C1 |
O3 |
168.569 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
