Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.679926 |
| Energy at 298.15K | |
| HF Energy | -151.140825 |
| Nuclear repulsion energy | 52.875337 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3535 |
3357 |
216.95 |
27.06 |
0.27 |
0.43 |
| 2 |
Σ |
2458 |
2334 |
1479.94 |
125.68 |
0.22 |
0.37 |
| 3 |
Σ |
1380 |
1310 |
26.40 |
1.88 |
0.03 |
0.05 |
| 4 |
Π |
616 |
585 |
0.12 |
1.90 |
0.75 |
0.86 |
| 4 |
Π |
595 |
565 |
15.12 |
6.07 |
0.75 |
0.86 |
| 5 |
Π |
439 |
417 |
26.41 |
0.29 |
0.75 |
0.86 |
| 5 |
Π |
474i |
450i |
148.49 |
8.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4274.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 4058.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
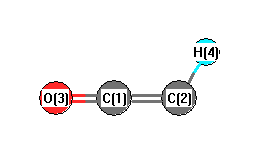
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.028 |
| C2 |
0.000 |
0.000 |
-1.238 |
| O3 |
0.000 |
0.000 |
1.194 |
| H4 |
0.000 |
0.000 |
-2.292 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2665 | 1.1656 | 2.3198 |
C2 | 1.2665 | | 2.4321 | 1.0533 | O3 | 1.1656 | 2.4321 | | 3.4854 | H4 | 2.3198 | 1.0533 | 3.4854 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.684733 |
| Energy at 298.15K | |
| HF Energy | -151.140068 |
| Nuclear repulsion energy | 52.515372 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3377 |
3206 |
52.14 |
65.85 |
0.30 |
0.46 |
| 2 |
A' |
2378 |
2258 |
1108.24 |
542.49 |
0.12 |
0.22 |
| 3 |
A' |
1260 |
1196 |
1.54 |
63.82 |
0.41 |
0.58 |
| 4 |
A' |
649 |
616 |
174.81 |
22.64 |
0.03 |
0.06 |
| 5 |
A' |
590 |
560 |
60.81 |
50.25 |
0.14 |
0.25 |
| 6 |
A" |
544 |
516 |
5.54 |
32.90 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4398.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 4176.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.050 |
0.000 |
| C2 |
0.991 |
-0.804 |
0.000 |
| O3 |
-0.998 |
0.642 |
0.000 |
| H4 |
2.041 |
-0.615 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3084 | 1.1606 | 2.1462 |
C2 | 1.3084 | | 2.4595 | 1.0668 | O3 | 1.1606 | 2.4595 | | 3.2885 | H4 | 2.1462 | 1.0668 | 3.2885 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
128.983 |
|
C2 |
C1 |
O3 |
169.905 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
