Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.722911 |
| Energy at 298.15K | |
| HF Energy | -151.140313 |
| Nuclear repulsion energy | 52.632446 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.025 |
| C2 |
0.000 |
0.000 |
-1.242 |
| O3 |
0.000 |
0.000 |
1.200 |
| H4 |
0.000 |
0.000 |
-2.298 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2675 | 1.1751 | 2.3227 |
C2 | 1.2675 | | 2.4425 | 1.0553 | O3 | 1.1751 | 2.4425 | | 3.4978 | H4 | 2.3227 | 1.0553 | 3.4978 | |
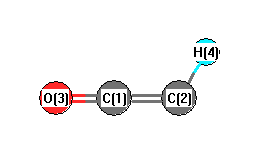 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.727885 |
| Energy at 298.15K | |
| HF Energy | -151.138981 |
| Nuclear repulsion energy | 52.259865 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3334 |
3210 |
|
|
|
|
| 2 |
A' |
2290 |
2206 |
|
|
|
|
| 3 |
A' |
1232 |
1186 |
|
|
|
|
| 4 |
A' |
637 |
614 |
|
|
|
|
| 5 |
A' |
575 |
554 |
|
|
|
|
| 6 |
A" |
522 |
502 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4295.0 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 4135.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.053 |
0.000 |
| C2 |
1.055 |
-0.727 |
0.000 |
| O3 |
-1.052 |
0.561 |
0.000 |
| H4 |
2.088 |
-0.449 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3125 | 1.1687 | 2.1476 |
C2 | 1.3125 | | 2.4705 | 1.0696 | O3 | 1.1687 | 2.4705 | | 3.2990 | H4 | 2.1476 | 1.0696 | 3.2990 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
128.435 |
|
C2 |
C1 |
O3 |
169.286 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
