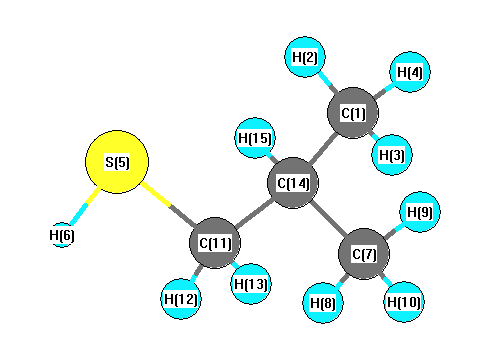
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3000 |
28.72 |
|
|
|
| 2 |
A |
3130 |
2995 |
32.52 |
|
|
|
| 3 |
A |
3128 |
2994 |
27.07 |
|
|
|
| 4 |
A |
3121 |
2987 |
30.87 |
|
|
|
| 5 |
A |
3117 |
2983 |
8.36 |
|
|
|
| 6 |
A |
3070 |
2938 |
30.75 |
|
|
|
| 7 |
A |
3051 |
2920 |
21.13 |
|
|
|
| 8 |
A |
3047 |
2916 |
23.31 |
|
|
|
| 9 |
A |
3043 |
2913 |
2.30 |
|
|
|
| 10 |
A |
2729 |
2612 |
3.93 |
|
|
|
| 11 |
A |
1506 |
1441 |
9.51 |
|
|
|
| 12 |
A |
1503 |
1439 |
9.18 |
|
|
|
| 13 |
A |
1493 |
1429 |
3.90 |
|
|
|
| 14 |
A |
1485 |
1421 |
0.55 |
|
|
|
| 15 |
A |
1475 |
1412 |
3.68 |
|
|
|
| 16 |
A |
1418 |
1357 |
8.74 |
|
|
|
| 17 |
A |
1401 |
1341 |
7.59 |
|
|
|
| 18 |
A |
1371 |
1313 |
1.65 |
|
|
|
| 19 |
A |
1356 |
1298 |
0.75 |
|
|
|
| 20 |
A |
1270 |
1215 |
16.62 |
|
|
|
| 21 |
A |
1237 |
1184 |
4.70 |
|
|
|
| 22 |
A |
1201 |
1149 |
4.83 |
|
|
|
| 23 |
A |
1153 |
1104 |
2.73 |
|
|
|
| 24 |
A |
1083 |
1037 |
2.72 |
|
|
|
| 25 |
A |
985 |
943 |
1.53 |
|
|
|
| 26 |
A |
972 |
931 |
0.73 |
|
|
|
| 27 |
A |
931 |
891 |
1.64 |
|
|
|
| 28 |
A |
907 |
868 |
1.68 |
|
|
|
| 29 |
A |
875 |
837 |
1.94 |
|
|
|
| 30 |
A |
811 |
776 |
2.75 |
|
|
|
| 31 |
A |
739 |
707 |
1.10 |
|
|
|
| 32 |
A |
411 |
394 |
0.11 |
|
|
|
| 33 |
A |
382 |
366 |
0.88 |
|
|
|
| 34 |
A |
329 |
315 |
0.11 |
|
|
|
| 35 |
A |
244 |
234 |
0.07 |
|
|
|
| 36 |
A |
227 |
217 |
4.37 |
|
|
|
| 37 |
A |
212 |
203 |
7.52 |
|
|
|
| 38 |
A |
196 |
188 |
3.97 |
|
|
|
| 39 |
A |
98 |
93 |
3.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28918.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27677.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.355 |
|
|
|
| 2 |
H |
0.133 |
|
|
|
| 3 |
H |
0.113 |
|
|
|
| 4 |
H |
0.117 |
|
|
|
| 5 |
S |
-0.199 |
|
|
|
| 6 |
H |
0.108 |
|
|
|
| 7 |
C |
-0.363 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
C |
-0.209 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.134 |
|
|
|
| 14 |
C |
-0.106 |
|
|
|
| 15 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.050 |
-1.044 |
0.508 |
1.566 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.104 |
2.333 |
-1.545 |
| y |
2.333 |
-40.179 |
-1.004 |
| z |
-1.545 |
-1.004 |
-41.442 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.293 |
2.333 |
-1.545 |
| y |
2.333 |
1.093 |
-1.004 |
| z |
-1.545 |
-1.004 |
-0.800 |
|
| Polar |
|---|
| 3z2-r2 | -1.600 |
|---|
| x2-y2 | -0.924 |
|---|
| xy | 2.333 |
|---|
| xz | -1.545 |
|---|
| yz | -1.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.116 |
-0.094 |
0.015 |
| y |
-0.094 |
9.612 |
-0.255 |
| z |
0.015 |
-0.255 |
8.335 |
<r2> (average value of r
2) Å
2
| <r2> |
198.399 |
| (<r2>)1/2 |
14.085 |