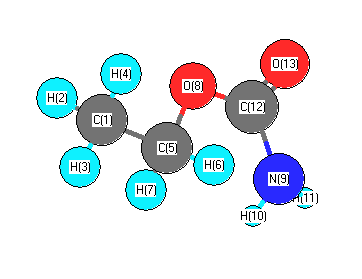
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3756 |
3595 |
50.18 |
|
|
|
| 2 |
A |
3635 |
3479 |
27.30 |
|
|
|
| 3 |
A |
3157 |
3021 |
18.74 |
|
|
|
| 4 |
A |
3150 |
3015 |
17.56 |
|
|
|
| 5 |
A |
3072 |
2940 |
13.46 |
|
|
|
| 6 |
A |
3071 |
2939 |
21.97 |
|
|
|
| 7 |
A |
3017 |
2888 |
38.59 |
|
|
|
| 8 |
A |
1866 |
1786 |
551.27 |
|
|
|
| 9 |
A |
1618 |
1548 |
145.38 |
|
|
|
| 10 |
A |
1525 |
1459 |
1.68 |
|
|
|
| 11 |
A |
1501 |
1437 |
3.29 |
|
|
|
| 12 |
A |
1483 |
1420 |
6.68 |
|
|
|
| 13 |
A |
1431 |
1370 |
11.88 |
|
|
|
| 14 |
A |
1405 |
1345 |
50.19 |
|
|
|
| 15 |
A |
1356 |
1298 |
339.30 |
|
|
|
| 16 |
A |
1313 |
1256 |
0.90 |
|
|
|
| 17 |
A |
1181 |
1130 |
4.66 |
|
|
|
| 18 |
A |
1167 |
1117 |
58.55 |
|
|
|
| 19 |
A |
1128 |
1079 |
72.46 |
|
|
|
| 20 |
A |
1091 |
1044 |
33.25 |
|
|
|
| 21 |
A |
1004 |
961 |
14.01 |
|
|
|
| 22 |
A |
873 |
836 |
11.40 |
|
|
|
| 23 |
A |
838 |
803 |
0.62 |
|
|
|
| 24 |
A |
785 |
752 |
25.39 |
|
|
|
| 25 |
A |
579 |
554 |
17.89 |
|
|
|
| 26 |
A |
556 |
532 |
6.93 |
|
|
|
| 27 |
A |
526 |
504 |
50.87 |
|
|
|
| 28 |
A |
381 |
365 |
2.71 |
|
|
|
| 29 |
A |
354 |
338 |
140.52 |
|
|
|
| 30 |
A |
257 |
246 |
0.27 |
|
|
|
| 31 |
A |
217 |
208 |
3.17 |
|
|
|
| 32 |
A |
121 |
116 |
0.48 |
|
|
|
| 33 |
A |
94 |
90 |
1.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23753.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22734.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.344 |
|
|
|
| 2 |
H |
0.128 |
|
|
|
| 3 |
H |
0.109 |
|
|
|
| 4 |
H |
0.130 |
|
|
|
| 5 |
C |
-0.054 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.087 |
|
|
|
| 8 |
O |
-0.179 |
|
|
|
| 9 |
N |
-0.279 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
| 12 |
C |
0.272 |
|
|
|
| 13 |
O |
-0.332 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.496 |
4.307 |
-0.535 |
5.007 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.781 |
-5.960 |
0.829 |
| y |
-5.960 |
-35.757 |
-1.240 |
| z |
0.829 |
-1.240 |
-35.998 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.096 |
-5.960 |
0.829 |
| y |
-5.960 |
0.133 |
-1.240 |
| z |
0.829 |
-1.240 |
-0.229 |
|
| Polar |
|---|
| 3z2-r2 | -0.459 |
|---|
| x2-y2 | -0.024 |
|---|
| xy | -5.960 |
|---|
| xz | 0.829 |
|---|
| yz | -1.240 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.323 |
0.246 |
-0.048 |
| y |
0.246 |
7.547 |
-0.041 |
| z |
-0.048 |
-0.041 |
5.600 |
<r2> (average value of r
2) Å
2
| <r2> |
189.463 |
| (<r2>)1/2 |
13.765 |