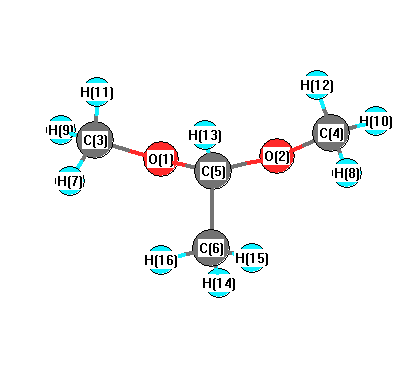
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3022 |
17.12 |
|
|
|
| 2 |
A |
3146 |
3011 |
23.14 |
|
|
|
| 3 |
A |
3068 |
2936 |
20.37 |
|
|
|
| 4 |
A |
3051 |
2920 |
86.83 |
|
|
|
| 5 |
A |
3001 |
2872 |
62.45 |
|
|
|
| 6 |
A |
2914 |
2789 |
62.83 |
|
|
|
| 7 |
A |
1510 |
1446 |
3.97 |
|
|
|
| 8 |
A |
1498 |
1433 |
3.57 |
|
|
|
| 9 |
A |
1483 |
1419 |
15.55 |
|
|
|
| 10 |
A |
1469 |
1406 |
2.57 |
|
|
|
| 11 |
A |
1405 |
1344 |
65.49 |
|
|
|
| 12 |
A |
1373 |
1314 |
1.14 |
|
|
|
| 13 |
A |
1269 |
1214 |
25.41 |
|
|
|
| 14 |
A |
1192 |
1141 |
37.71 |
|
|
|
| 15 |
A |
1179 |
1128 |
0.44 |
|
|
|
| 16 |
A |
1131 |
1083 |
7.94 |
|
|
|
| 17 |
A |
1089 |
1042 |
48.26 |
|
|
|
| 18 |
A |
851 |
815 |
6.27 |
|
|
|
| 19 |
A |
534 |
511 |
6.09 |
|
|
|
| 20 |
A |
447 |
428 |
10.19 |
|
|
|
| 21 |
A |
243 |
233 |
2.56 |
|
|
|
| 22 |
A |
167 |
159 |
1.53 |
|
|
|
| 23 |
A |
65 |
62 |
3.80 |
|
|
|
| 24 |
A |
3146 |
3011 |
16.43 |
|
|
|
| 25 |
A |
3141 |
3006 |
35.41 |
|
|
|
| 26 |
A |
3051 |
2920 |
14.57 |
|
|
|
| 27 |
A |
2999 |
2871 |
59.98 |
|
|
|
| 28 |
A |
1512 |
1447 |
7.47 |
|
|
|
| 29 |
A |
1489 |
1425 |
2.35 |
|
|
|
| 30 |
A |
1485 |
1421 |
7.75 |
|
|
|
| 31 |
A |
1472 |
1408 |
0.69 |
|
|
|
| 32 |
A |
1394 |
1334 |
51.45 |
|
|
|
| 33 |
A |
1218 |
1166 |
205.85 |
|
|
|
| 34 |
A |
1179 |
1129 |
108.41 |
|
|
|
| 35 |
A |
1178 |
1127 |
124.39 |
|
|
|
| 36 |
A |
1095 |
1048 |
14.80 |
|
|
|
| 37 |
A |
924 |
884 |
6.67 |
|
|
|
| 38 |
A |
461 |
441 |
2.08 |
|
|
|
| 39 |
A |
338 |
323 |
0.89 |
|
|
|
| 40 |
A |
217 |
208 |
0.04 |
|
|
|
| 41 |
A |
191 |
183 |
0.02 |
|
|
|
| 42 |
A |
54 |
52 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30891.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29566.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.237 |
|
|
|
| 2 |
O |
-0.237 |
|
|
|
| 3 |
C |
-0.170 |
|
|
|
| 4 |
C |
-0.170 |
|
|
|
| 5 |
C |
0.219 |
|
|
|
| 6 |
C |
-0.355 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.081 |
|
|
|
| 9 |
H |
0.128 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.078 |
|
|
|
| 12 |
H |
0.078 |
|
|
|
| 13 |
H |
0.032 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.108 |
|
|
|
| 16 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.856 |
2.129 |
0.000 |
2.295 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.492 |
-1.216 |
0.000 |
| y |
-1.216 |
-39.777 |
0.000 |
| z |
0.000 |
0.000 |
-32.437 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.385 |
-1.216 |
0.000 |
| y |
-1.216 |
-4.313 |
0.000 |
| z |
0.000 |
0.000 |
6.698 |
|
| Polar |
|---|
| 3z2-r2 | 13.396 |
|---|
| x2-y2 | 1.286 |
|---|
| xy | -1.216 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.210 |
-0.129 |
0.000 |
| y |
-0.129 |
7.560 |
0.000 |
| z |
0.000 |
0.000 |
10.310 |
<r2> (average value of r
2) Å
2
| <r2> |
196.797 |
| (<r2>)1/2 |
14.028 |