Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -269.319530 |
| Energy at 298.15K | -269.326214 |
| HF Energy | -269.319530 |
| Nuclear repulsion energy | 224.073606 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3305 |
3163 |
0.81 |
|
|
|
| 2 |
A' |
3281 |
3141 |
1.07 |
|
|
|
| 3 |
A' |
3269 |
3129 |
4.64 |
|
|
|
| 4 |
A' |
3160 |
3024 |
9.35 |
|
|
|
| 5 |
A' |
3057 |
2926 |
27.20 |
|
|
|
| 6 |
A' |
1668 |
1597 |
15.99 |
|
|
|
| 7 |
A' |
1570 |
1502 |
20.91 |
|
|
|
| 8 |
A' |
1493 |
1429 |
7.26 |
|
|
|
| 9 |
A' |
1432 |
1371 |
5.72 |
|
|
|
| 10 |
A' |
1412 |
1351 |
0.58 |
|
|
|
| 11 |
A' |
1273 |
1219 |
18.42 |
|
|
|
| 12 |
A' |
1260 |
1206 |
0.40 |
|
|
|
| 13 |
A' |
1196 |
1145 |
18.95 |
|
|
|
| 14 |
A' |
1132 |
1084 |
4.09 |
|
|
|
| 15 |
A' |
1042 |
997 |
23.85 |
|
|
|
| 16 |
A' |
994 |
952 |
1.84 |
|
|
|
| 17 |
A' |
951 |
911 |
15.54 |
|
|
|
| 18 |
A' |
901 |
862 |
3.62 |
|
|
|
| 19 |
A' |
665 |
637 |
0.87 |
|
|
|
| 20 |
A' |
334 |
320 |
2.03 |
|
|
|
| 21 |
A" |
3113 |
2979 |
13.09 |
|
|
|
| 22 |
A" |
1473 |
1409 |
7.08 |
|
|
|
| 23 |
A" |
1063 |
1017 |
1.72 |
|
|
|
| 24 |
A" |
888 |
850 |
0.13 |
|
|
|
| 25 |
A" |
822 |
787 |
17.64 |
|
|
|
| 26 |
A" |
753 |
721 |
56.86 |
|
|
|
| 27 |
A" |
649 |
621 |
1.33 |
|
|
|
| 28 |
A" |
622 |
595 |
6.40 |
|
|
|
| 29 |
A" |
245 |
234 |
5.43 |
|
|
|
| 30 |
A" |
125 |
119 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21573.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20648.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.028 |
-0.241 |
0.000 |
| C2 |
0.000 |
0.641 |
0.000 |
| C3 |
0.495 |
-1.484 |
0.000 |
| C4 |
-1.175 |
-0.030 |
0.000 |
| C5 |
-0.851 |
-1.418 |
0.000 |
| C6 |
0.353 |
2.075 |
0.000 |
| H7 |
1.195 |
-2.297 |
0.000 |
| H8 |
-2.154 |
0.412 |
0.000 |
| H9 |
-1.531 |
-2.249 |
0.000 |
| H10 |
-0.550 |
2.677 |
0.000 |
| H11 |
0.941 |
2.338 |
0.879 |
| H12 |
0.941 |
2.338 |
-0.879 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3552 | 1.3521 | 2.2131 | 2.2171 | 2.4128 | 2.0626 | 3.2490 | 3.2532 | 3.3177 | 2.7265 | 2.7265 |
C2 | 1.3552 | | 2.1822 | 1.3528 | 2.2280 | 1.4766 | 3.1724 | 2.1665 | 3.2714 | 2.1084 | 2.1298 | 2.1298 | C3 | 1.3521 | 2.1822 | | 2.2143 | 1.3478 | 3.5618 | 1.0733 | 3.2580 | 2.1660 | 4.2899 | 3.9470 | 3.9470 | C4 | 2.2131 | 1.3528 | 2.2143 | | 1.4252 | 2.6009 | 3.2801 | 1.0747 | 2.2483 | 2.7773 | 3.2944 | 3.2944 | C5 | 2.2171 | 2.2280 | 1.3478 | 1.4252 | | 3.6945 | 2.2275 | 2.2464 | 1.0745 | 4.1053 | 4.2530 | 4.2530 | C6 | 2.4128 | 1.4766 | 3.5618 | 2.6009 | 3.6945 | | 4.4526 | 3.0089 | 4.7173 | 1.0856 | 1.0895 | 1.0895 | H7 | 2.0626 | 3.1724 | 1.0733 | 3.2801 | 2.2275 | 4.4526 | | 4.3083 | 2.7270 | 5.2713 | 4.7247 | 4.7247 | H8 | 3.2490 | 2.1665 | 3.2580 | 1.0747 | 2.2464 | 3.0089 | 4.3083 | | 2.7336 | 2.7750 | 3.7498 | 3.7498 | H9 | 3.2532 | 3.2714 | 2.1660 | 2.2483 | 1.0745 | 4.7173 | 2.7270 | 2.7336 | | 5.0229 | 5.2847 | 5.2847 | H10 | 3.3177 | 2.1084 | 4.2899 | 2.7773 | 4.1053 | 1.0856 | 5.2713 | 2.7750 | 5.0229 | | 1.7637 | 1.7637 | H11 | 2.7265 | 2.1298 | 3.9470 | 3.2944 | 4.2530 | 1.0895 | 4.7247 | 3.7498 | 5.2847 | 1.7637 | | 1.7582 | H12 | 2.7265 | 2.1298 | 3.9470 | 3.2944 | 4.2530 | 1.0895 | 4.7247 | 3.7498 | 5.2847 | 1.7637 | 1.7582 | |
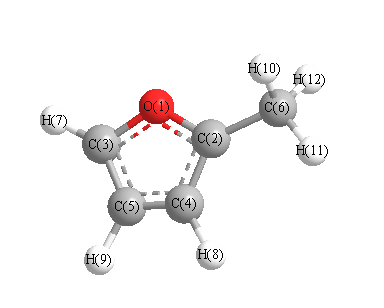 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.617 |
|
O1 |
C2 |
C6 |
116.804 |
| O1 |
C3 |
C5 |
110.406 |
|
O1 |
C3 |
H7 |
116.040 |
| C2 |
O1 |
C3 |
107.422 |
|
C2 |
C4 |
C5 |
106.614 |
| C2 |
C4 |
H8 |
125.993 |
|
C2 |
C6 |
H10 |
109.807 |
| C2 |
C6 |
H11 |
111.308 |
|
C2 |
C6 |
H12 |
111.308 |
| C3 |
C5 |
C6 |
73.792 |
|
C3 |
C5 |
H9 |
126.446 |
| C4 |
C2 |
C6 |
133.579 |
|
C4 |
C5 |
H9 |
127.614 |
| C5 |
C3 |
H7 |
133.554 |
|
C5 |
C4 |
H8 |
127.393 |
| H10 |
C6 |
H11 |
108.361 |
|
H10 |
C6 |
H12 |
108.361 |
| H11 |
C6 |
H12 |
107.589 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.145 |
|
|
|
| 2 |
C |
0.171 |
|
|
|
| 3 |
C |
-0.064 |
|
|
|
| 4 |
C |
-0.232 |
|
|
|
| 5 |
C |
-0.184 |
|
|
|
| 6 |
C |
-0.349 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.141 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.404 |
0.392 |
0.000 |
0.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.050 |
-0.460 |
0.000 |
| y |
-0.460 |
-30.188 |
0.000 |
| z |
0.000 |
0.000 |
-37.666 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.123 |
-0.460 |
0.000 |
| y |
-0.460 |
5.670 |
0.000 |
| z |
0.000 |
0.000 |
-5.548 |
|
| Polar |
|---|
| 3z2-r2 | -11.096 |
|---|
| x2-y2 | -3.862 |
|---|
| xy | -0.460 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.855 |
0.356 |
0.000 |
| y |
0.356 |
10.769 |
0.000 |
| z |
0.000 |
0.000 |
5.433 |
<r2> (average value of r
2) Å
2
| <r2> |
139.202 |
| (<r2>)1/2 |
11.798 |
