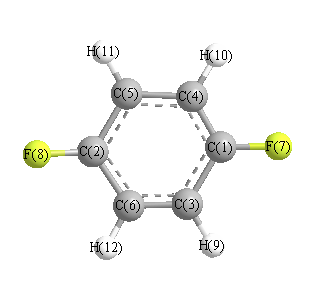
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3274 |
3108 |
0.00 |
267.25 |
0.17 |
0.28 |
| 2 |
Ag |
1678 |
1593 |
0.00 |
5.22 |
0.61 |
0.76 |
| 3 |
Ag |
1307 |
1241 |
0.00 |
20.82 |
0.11 |
0.20 |
| 4 |
Ag |
1173 |
1113 |
0.00 |
6.70 |
0.64 |
0.78 |
| 5 |
Ag |
876 |
832 |
0.00 |
33.27 |
0.08 |
0.14 |
| 6 |
Ag |
456 |
433 |
0.00 |
4.60 |
0.43 |
0.60 |
| 7 |
Au |
979 |
929 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
442 |
420 |
0.00 |
0.00 |
0.00 |
0.00 |
| 9 |
B1g |
831 |
789 |
0.00 |
0.15 |
0.75 |
0.86 |
| 10 |
B1u |
3252 |
3088 |
2.18 |
0.00 |
0.31 |
0.47 |
| 11 |
B1u |
1562 |
1483 |
261.69 |
0.00 |
0.26 |
0.00 |
| 12 |
B1u |
1253 |
1190 |
158.52 |
0.00 |
0.40 |
0.00 |
| 13 |
B1u |
1038 |
985 |
1.42 |
0.00 |
0.00 |
0.00 |
| 14 |
B1u |
763 |
724 |
54.09 |
0.00 |
0.00 |
0.00 |
| 15 |
B2g |
957 |
909 |
0.00 |
0.29 |
0.75 |
0.86 |
| 16 |
B2g |
722 |
685 |
0.00 |
0.57 |
0.75 |
0.86 |
| 17 |
B2g |
389 |
369 |
0.00 |
3.31 |
0.75 |
0.86 |
| 18 |
B2u |
3273 |
3107 |
0.08 |
0.00 |
0.40 |
0.57 |
| 19 |
B2u |
1488 |
1412 |
0.97 |
0.00 |
0.51 |
0.68 |
| 20 |
B2u |
1440 |
1367 |
0.02 |
0.00 |
0.51 |
0.68 |
| 21 |
B2u |
1116 |
1060 |
12.89 |
0.00 |
0.50 |
0.67 |
| 22 |
B2u |
351 |
333 |
4.04 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3256 |
3091 |
0.00 |
109.30 |
0.75 |
0.86 |
| 24 |
B3g |
1671 |
1586 |
0.00 |
5.71 |
0.75 |
0.86 |
| 25 |
B3g |
1299 |
1233 |
0.00 |
1.75 |
0.75 |
0.86 |
| 26 |
B3g |
646 |
613 |
0.00 |
6.52 |
0.75 |
0.86 |
| 27 |
B3g |
445 |
422 |
0.00 |
0.04 |
0.75 |
0.86 |
| 28 |
B3u |
875 |
831 |
80.11 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
528 |
502 |
17.31 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
163 |
155 |
1.51 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 18751.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17802.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.