Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -578.050014 |
| Energy at 298.15K | -578.057755 |
| HF Energy | -577.247035 |
| Nuclear repulsion energy | 159.825788 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3173 |
3013 |
19.47 |
|
|
|
| 2 |
A' |
3131 |
2973 |
24.77 |
|
|
|
| 3 |
A' |
3111 |
2954 |
6.17 |
|
|
|
| 4 |
A' |
3092 |
2936 |
17.29 |
|
|
|
| 5 |
A' |
1534 |
1456 |
7.30 |
|
|
|
| 6 |
A' |
1518 |
1441 |
0.55 |
|
|
|
| 7 |
A' |
1509 |
1433 |
1.66 |
|
|
|
| 8 |
A' |
1425 |
1353 |
2.81 |
|
|
|
| 9 |
A' |
1387 |
1317 |
7.84 |
|
|
|
| 10 |
A' |
1299 |
1233 |
18.73 |
|
|
|
| 11 |
A' |
1145 |
1087 |
0.36 |
|
|
|
| 12 |
A' |
1079 |
1024 |
1.39 |
|
|
|
| 13 |
A' |
934 |
887 |
13.65 |
|
|
|
| 14 |
A' |
787 |
747 |
26.80 |
|
|
|
| 15 |
A' |
372 |
354 |
1.90 |
|
|
|
| 16 |
A' |
236 |
224 |
1.69 |
|
|
|
| 17 |
A" |
3182 |
3021 |
20.92 |
|
|
|
| 18 |
A" |
3163 |
3003 |
15.53 |
|
|
|
| 19 |
A" |
3139 |
2981 |
0.07 |
|
|
|
| 20 |
A" |
1528 |
1451 |
8.18 |
|
|
|
| 21 |
A" |
1341 |
1273 |
0.00 |
|
|
|
| 22 |
A" |
1266 |
1202 |
0.37 |
|
|
|
| 23 |
A" |
1113 |
1057 |
0.98 |
|
|
|
| 24 |
A" |
882 |
838 |
0.10 |
|
|
|
| 25 |
A" |
765 |
726 |
2.83 |
|
|
|
| 26 |
A" |
249 |
237 |
0.01 |
|
|
|
| 27 |
A" |
125 |
119 |
1.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21243.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20168.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.575 |
0.000 |
| C2 |
0.888 |
-0.641 |
0.000 |
| C3 |
2.354 |
-0.252 |
0.000 |
| Cl4 |
-1.725 |
0.147 |
0.000 |
| H5 |
0.183 |
1.181 |
0.879 |
| H6 |
0.183 |
1.181 |
-0.879 |
| H7 |
0.655 |
-1.247 |
-0.872 |
| H8 |
0.655 |
-1.247 |
0.872 |
| H9 |
2.993 |
-1.129 |
0.000 |
| H10 |
2.604 |
0.339 |
-0.877 |
| H11 |
2.604 |
0.339 |
0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5052 | 2.4952 | 1.7775 | 1.0838 | 1.0838 | 2.1226 | 2.1226 | 3.4435 | 2.7581 | 2.7581 |
C2 | 1.5052 | | 1.5176 | 2.7290 | 2.1422 | 2.1422 | 1.0867 | 1.0867 | 2.1610 | 2.1625 | 2.1625 | C3 | 2.4952 | 1.5176 | | 4.0990 | 2.7457 | 2.7457 | 2.1535 | 2.1535 | 1.0847 | 1.0864 | 1.0864 | Cl4 | 1.7775 | 2.7290 | 4.0990 | | 2.3422 | 2.3422 | 2.8925 | 2.8925 | 4.8874 | 4.4216 | 4.4216 | H5 | 1.0838 | 2.1422 | 2.7457 | 2.3422 | | 1.7585 | 3.0301 | 2.4729 | 3.7417 | 3.1072 | 2.5632 | H6 | 1.0838 | 2.1422 | 2.7457 | 2.3422 | 1.7585 | | 2.4729 | 3.0301 | 3.7417 | 2.5632 | 3.1072 | H7 | 2.1226 | 1.0867 | 2.1535 | 2.8925 | 3.0301 | 2.4729 | | 1.7439 | 2.4980 | 2.5125 | 3.0613 | H8 | 2.1226 | 1.0867 | 2.1535 | 2.8925 | 2.4729 | 3.0301 | 1.7439 | | 2.4980 | 3.0613 | 2.5125 | H9 | 3.4435 | 2.1610 | 1.0847 | 4.8874 | 3.7417 | 3.7417 | 2.4980 | 2.4980 | | 1.7530 | 1.7530 | H10 | 2.7581 | 2.1625 | 1.0864 | 4.4216 | 3.1072 | 2.5632 | 2.5125 | 3.0613 | 1.7530 | | 1.7541 | H11 | 2.7581 | 2.1625 | 1.0864 | 4.4216 | 2.5632 | 3.1072 | 3.0613 | 2.5125 | 1.7530 | 1.7541 | |
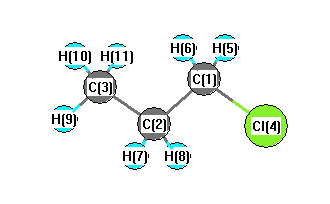 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.266 |
|
C1 |
C2 |
H7 |
108.889 |
| C1 |
C2 |
H8 |
108.889 |
|
C2 |
C1 |
Cl4 |
112.204 |
| C2 |
C1 |
H5 |
110.619 |
|
C2 |
C1 |
H6 |
110.619 |
| C2 |
C3 |
H9 |
111.195 |
|
C2 |
C3 |
H10 |
111.209 |
| C2 |
C3 |
H11 |
111.209 |
|
C3 |
C2 |
H7 |
110.474 |
| C3 |
C2 |
H8 |
110.474 |
|
Cl4 |
C1 |
H5 |
107.395 |
| Cl4 |
C1 |
H6 |
107.395 |
|
H5 |
C1 |
H6 |
108.450 |
| H7 |
C2 |
H8 |
106.721 |
|
H9 |
C3 |
H10 |
107.693 |
| H9 |
C3 |
H11 |
107.693 |
|
H10 |
C3 |
H11 |
107.666 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -578.050294 |
| Energy at 298.15K | -578.058156 |
| Nuclear repulsion energy | 164.095825 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3023 |
9.48 |
|
|
|
| 2 |
A |
3182 |
3021 |
14.35 |
|
|
|
| 3 |
A |
3168 |
3007 |
24.96 |
|
|
|
| 4 |
A |
3138 |
2979 |
20.54 |
|
|
|
| 5 |
A |
3125 |
2967 |
3.90 |
|
|
|
| 6 |
A |
3096 |
2939 |
17.82 |
|
|
|
| 7 |
A |
3090 |
2934 |
17.09 |
|
|
|
| 8 |
A |
1534 |
1457 |
6.12 |
|
|
|
| 9 |
A |
1527 |
1450 |
7.19 |
|
|
|
| 10 |
A |
1507 |
1430 |
3.69 |
|
|
|
| 11 |
A |
1500 |
1424 |
4.27 |
|
|
|
| 12 |
A |
1428 |
1356 |
6.52 |
|
|
|
| 13 |
A |
1394 |
1324 |
0.02 |
|
|
|
| 14 |
A |
1353 |
1284 |
22.61 |
|
|
|
| 15 |
A |
1303 |
1237 |
3.93 |
|
|
|
| 16 |
A |
1259 |
1195 |
0.51 |
|
|
|
| 17 |
A |
1134 |
1076 |
0.84 |
|
|
|
| 18 |
A |
1105 |
1049 |
1.07 |
|
|
|
| 19 |
A |
1075 |
1021 |
2.26 |
|
|
|
| 20 |
A |
926 |
879 |
8.35 |
|
|
|
| 21 |
A |
888 |
843 |
3.13 |
|
|
|
| 22 |
A |
811 |
770 |
15.83 |
|
|
|
| 23 |
A |
700 |
665 |
11.69 |
|
|
|
| 24 |
A |
429 |
407 |
0.94 |
|
|
|
| 25 |
A |
307 |
291 |
0.47 |
|
|
|
| 26 |
A |
226 |
214 |
1.01 |
|
|
|
| 27 |
A |
142 |
135 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21265.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20189.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.180 |
0.880 |
0.310 |
| C2 |
-1.134 |
0.562 |
-0.359 |
| C3 |
-1.767 |
-0.719 |
0.145 |
| Cl4 |
1.425 |
-0.344 |
-0.069 |
| H5 |
0.577 |
1.830 |
-0.022 |
| H6 |
0.079 |
0.893 |
1.389 |
| H7 |
-1.799 |
1.403 |
-0.171 |
| H8 |
-0.979 |
0.512 |
-1.433 |
| H9 |
-2.721 |
-0.897 |
-0.339 |
| H10 |
-1.122 |
-1.568 |
-0.050 |
| H11 |
-1.938 |
-0.666 |
1.217 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5080 | 2.5245 | 1.7869 | 1.0817 | 1.0835 | 2.1027 | 2.1250 | 3.4628 | 2.7967 | 2.7747 |
C2 | 1.5080 | | 1.5151 | 2.7300 | 2.1561 | 2.1529 | 1.0887 | 1.0863 | 2.1556 | 2.1530 | 2.1536 | C3 | 2.5245 | 1.5151 | | 3.2207 | 3.4668 | 2.7481 | 2.1454 | 2.1508 | 1.0845 | 1.0841 | 1.0865 | Cl4 | 1.7869 | 2.7300 | 3.2207 | | 2.3339 | 2.3380 | 3.6687 | 2.8933 | 4.1909 | 2.8263 | 3.6148 | H5 | 1.0817 | 2.1561 | 3.4668 | 2.3339 | | 1.7650 | 2.4194 | 2.4798 | 4.2909 | 3.7999 | 3.7539 | H6 | 1.0835 | 2.1529 | 2.7481 | 2.3380 | 1.7650 | | 2.4941 | 3.0374 | 3.7454 | 3.0941 | 2.5554 | H7 | 2.1027 | 1.0887 | 2.1454 | 3.6687 | 2.4194 | 2.4941 | | 1.7495 | 2.4830 | 3.0500 | 2.4949 | H8 | 2.1250 | 1.0863 | 2.1508 | 2.8933 | 2.4798 | 3.0374 | 1.7495 | | 2.4930 | 2.5023 | 3.0543 | H9 | 3.4628 | 2.1556 | 1.0845 | 4.1909 | 4.2909 | 3.7454 | 2.4830 | 2.4930 | | 1.7577 | 1.7568 | H10 | 2.7967 | 2.1530 | 1.0841 | 2.8263 | 3.7999 | 3.0941 | 3.0500 | 2.5023 | 1.7577 | | 1.7565 | H11 | 2.7747 | 2.1536 | 1.0865 | 3.6148 | 3.7539 | 2.5554 | 2.4949 | 3.0543 | 1.7568 | 1.7565 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.241 |
|
C1 |
C2 |
H7 |
107.040 |
| C1 |
C2 |
H8 |
108.900 |
|
C2 |
C1 |
Cl4 |
111.622 |
| C2 |
C1 |
H5 |
111.667 |
|
C2 |
C1 |
H6 |
111.296 |
| C2 |
C3 |
H9 |
110.950 |
|
C2 |
C3 |
H10 |
110.762 |
| C2 |
C3 |
H11 |
110.664 |
|
C3 |
C2 |
H7 |
109.878 |
| C3 |
C2 |
H8 |
110.452 |
|
Cl4 |
C1 |
H5 |
106.288 |
| Cl4 |
C1 |
H6 |
106.498 |
|
H5 |
C1 |
H6 |
109.212 |
| H7 |
C2 |
H8 |
107.093 |
|
H9 |
C3 |
H10 |
108.290 |
| H9 |
C3 |
H11 |
108.033 |
|
H10 |
C3 |
H11 |
108.031 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
