Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -578.780323 |
| Energy at 298.15K | -578.787933 |
| Nuclear repulsion energy | 159.257307 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3136 |
3002 |
26.06 |
|
|
|
| 2 |
A' |
3100 |
2967 |
25.92 |
|
|
|
| 3 |
A' |
3074 |
2942 |
9.11 |
|
|
|
| 4 |
A' |
3056 |
2925 |
21.79 |
|
|
|
| 5 |
A' |
1508 |
1444 |
7.51 |
|
|
|
| 6 |
A' |
1493 |
1429 |
0.58 |
|
|
|
| 7 |
A' |
1487 |
1423 |
1.74 |
|
|
|
| 8 |
A' |
1409 |
1348 |
2.63 |
|
|
|
| 9 |
A' |
1369 |
1310 |
6.09 |
|
|
|
| 10 |
A' |
1277 |
1222 |
20.72 |
|
|
|
| 11 |
A' |
1127 |
1079 |
0.43 |
|
|
|
| 12 |
A' |
1056 |
1011 |
1.45 |
|
|
|
| 13 |
A' |
916 |
876 |
14.32 |
|
|
|
| 14 |
A' |
754 |
721 |
35.46 |
|
|
|
| 15 |
A' |
360 |
344 |
2.66 |
|
|
|
| 16 |
A' |
225 |
215 |
1.89 |
|
|
|
| 17 |
A" |
3160 |
3025 |
20.41 |
|
|
|
| 18 |
A" |
3130 |
2995 |
28.49 |
|
|
|
| 19 |
A" |
3106 |
2972 |
0.03 |
|
|
|
| 20 |
A" |
1496 |
1432 |
8.12 |
|
|
|
| 21 |
A" |
1319 |
1263 |
0.00 |
|
|
|
| 22 |
A" |
1244 |
1191 |
0.27 |
|
|
|
| 23 |
A" |
1093 |
1046 |
0.99 |
|
|
|
| 24 |
A" |
867 |
830 |
0.12 |
|
|
|
| 25 |
A" |
748 |
716 |
3.54 |
|
|
|
| 26 |
A" |
218 |
209 |
0.01 |
|
|
|
| 27 |
A" |
116 |
111 |
1.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20921.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20023.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.583 |
0.000 |
| C2 |
0.897 |
-0.630 |
0.000 |
| C3 |
2.363 |
-0.228 |
0.000 |
| Cl4 |
-1.732 |
0.124 |
0.000 |
| H5 |
0.154 |
1.195 |
0.883 |
| H6 |
0.154 |
1.195 |
-0.883 |
| H7 |
0.672 |
-1.239 |
-0.874 |
| H8 |
0.672 |
-1.239 |
0.874 |
| H9 |
3.006 |
-1.105 |
0.000 |
| H10 |
2.611 |
0.364 |
-0.880 |
| H11 |
2.611 |
0.364 |
0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5089 | 2.4984 | 1.7916 | 1.0856 | 1.0856 | 2.1293 | 2.1293 | 3.4473 | 2.7641 | 2.7641 |
C2 | 1.5089 | | 1.5203 | 2.7354 | 2.1596 | 2.1596 | 1.0888 | 1.0888 | 2.1613 | 2.1681 | 2.1681 | C3 | 2.4984 | 1.5203 | | 4.1103 | 2.7719 | 2.7719 | 2.1558 | 2.1558 | 1.0876 | 1.0890 | 1.0890 | Cl4 | 1.7916 | 2.7354 | 4.1103 | | 2.3418 | 2.3418 | 2.8985 | 2.8985 | 4.8947 | 4.4378 | 4.4378 | H5 | 1.0856 | 2.1596 | 2.7719 | 2.3418 | | 1.7662 | 3.0463 | 2.4883 | 3.7685 | 3.1360 | 2.5936 | H6 | 1.0856 | 2.1596 | 2.7719 | 2.3418 | 1.7662 | | 2.4883 | 3.0463 | 3.7685 | 2.5936 | 3.1360 | H7 | 2.1293 | 1.0888 | 2.1558 | 2.8985 | 3.0463 | 2.4883 | | 1.7487 | 2.4958 | 2.5161 | 3.0672 | H8 | 2.1293 | 1.0888 | 2.1558 | 2.8985 | 2.4883 | 3.0463 | 1.7487 | | 2.4958 | 3.0672 | 2.5161 | H9 | 3.4473 | 2.1613 | 1.0876 | 4.8947 | 3.7685 | 3.7685 | 2.4958 | 2.4958 | | 1.7576 | 1.7576 | H10 | 2.7641 | 2.1681 | 1.0890 | 4.4378 | 3.1360 | 2.5936 | 2.5161 | 3.0672 | 1.7576 | | 1.7596 | H11 | 2.7641 | 2.1681 | 1.0890 | 4.4378 | 2.5936 | 3.1360 | 3.0672 | 2.5161 | 1.7576 | 1.7596 | |
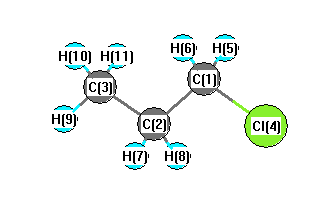 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.131 |
|
C1 |
C2 |
H7 |
109.039 |
| C1 |
C2 |
H8 |
109.039 |
|
C2 |
C1 |
Cl4 |
111.666 |
| C2 |
C1 |
H5 |
111.649 |
|
C2 |
C1 |
H6 |
111.649 |
| C2 |
C3 |
H9 |
110.852 |
|
C2 |
C3 |
H10 |
111.314 |
| C2 |
C3 |
H11 |
111.314 |
|
C3 |
C2 |
H7 |
110.340 |
| C3 |
C2 |
H8 |
110.340 |
|
Cl4 |
C1 |
H5 |
106.359 |
| Cl4 |
C1 |
H6 |
106.359 |
|
H5 |
C1 |
H6 |
108.871 |
| H7 |
C2 |
H8 |
106.842 |
|
H9 |
C3 |
H10 |
107.704 |
| H9 |
C3 |
H11 |
107.704 |
|
H10 |
C3 |
H11 |
107.782 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.208 |
|
|
|
| 3 |
C |
-0.363 |
|
|
|
| 4 |
Cl |
-0.177 |
|
|
|
| 5 |
H |
0.140 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.126 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.187 |
0.265 |
0.000 |
2.203 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.445 |
0.118 |
0.000 |
| y |
0.118 |
-32.440 |
0.000 |
| z |
0.000 |
0.000 |
-32.516 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.967 |
0.118 |
0.000 |
| y |
0.118 |
1.540 |
0.000 |
| z |
0.000 |
0.000 |
1.427 |
|
| Polar |
|---|
| 3z2-r2 | 2.853 |
|---|
| x2-y2 | -3.004 |
|---|
| xy | 0.118 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.374 |
-0.008 |
0.000 |
| y |
-0.008 |
6.471 |
0.000 |
| z |
0.000 |
0.000 |
6.024 |
<r2> (average value of r
2) Å
2
| <r2> |
150.904 |
| (<r2>)1/2 |
12.284 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -578.780376 |
| Energy at 298.15K | -578.788110 |
| Nuclear repulsion energy | 163.432924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3028 |
11.63 |
|
|
|
| 2 |
A |
3146 |
3011 |
19.75 |
|
|
|
| 3 |
A |
3132 |
2997 |
35.06 |
|
|
|
| 4 |
A |
3107 |
2974 |
22.08 |
|
|
|
| 5 |
A |
3092 |
2959 |
4.23 |
|
|
|
| 6 |
A |
3060 |
2928 |
20.69 |
|
|
|
| 7 |
A |
3049 |
2918 |
21.91 |
|
|
|
| 8 |
A |
1506 |
1441 |
5.29 |
|
|
|
| 9 |
A |
1496 |
1432 |
8.81 |
|
|
|
| 10 |
A |
1481 |
1417 |
3.64 |
|
|
|
| 11 |
A |
1475 |
1412 |
3.86 |
|
|
|
| 12 |
A |
1410 |
1349 |
7.08 |
|
|
|
| 13 |
A |
1379 |
1320 |
0.11 |
|
|
|
| 14 |
A |
1331 |
1274 |
23.90 |
|
|
|
| 15 |
A |
1281 |
1226 |
3.99 |
|
|
|
| 16 |
A |
1238 |
1185 |
0.46 |
|
|
|
| 17 |
A |
1114 |
1066 |
0.83 |
|
|
|
| 18 |
A |
1085 |
1039 |
1.10 |
|
|
|
| 19 |
A |
1052 |
1007 |
2.83 |
|
|
|
| 20 |
A |
909 |
870 |
9.13 |
|
|
|
| 21 |
A |
871 |
834 |
3.45 |
|
|
|
| 22 |
A |
790 |
756 |
19.25 |
|
|
|
| 23 |
A |
672 |
643 |
16.71 |
|
|
|
| 24 |
A |
417 |
399 |
1.31 |
|
|
|
| 25 |
A |
289 |
277 |
0.64 |
|
|
|
| 26 |
A |
200 |
191 |
1.15 |
|
|
|
| 27 |
A |
135 |
129 |
0.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20939.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20040.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.180 |
0.883 |
0.306 |
| C2 |
-1.140 |
0.563 |
-0.353 |
| C3 |
-1.780 |
-0.719 |
0.144 |
| Cl4 |
1.435 |
-0.346 |
-0.068 |
| H5 |
0.581 |
1.830 |
-0.035 |
| H6 |
0.094 |
0.902 |
1.389 |
| H7 |
-1.803 |
1.409 |
-0.161 |
| H8 |
-0.994 |
0.519 |
-1.432 |
| H9 |
-2.732 |
-0.895 |
-0.351 |
| H10 |
-1.135 |
-1.573 |
-0.050 |
| H11 |
-1.963 |
-0.674 |
1.217 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5099 | 2.5364 | 1.7953 | 1.0845 | 1.0859 | 2.1041 | 2.1290 | 3.4750 | 2.8084 | 2.8013 |
C2 | 1.5099 | | 1.5166 | 2.7449 | 2.1611 | 2.1615 | 1.0923 | 1.0894 | 2.1593 | 2.1572 | 2.1619 | C3 | 2.5364 | 1.5166 | | 3.2430 | 3.4795 | 2.7729 | 2.1501 | 2.1528 | 1.0876 | 1.0871 | 1.0894 | Cl4 | 1.7953 | 2.7449 | 3.2430 | | 2.3378 | 2.3404 | 3.6837 | 2.9165 | 4.2126 | 2.8480 | 3.6477 | H5 | 1.0845 | 2.1611 | 3.4795 | 2.3378 | | 1.7684 | 2.4243 | 2.4802 | 4.3023 | 3.8118 | 3.7838 | H6 | 1.0859 | 2.1615 | 2.7729 | 2.3404 | 1.7684 | | 2.5010 | 3.0474 | 3.7742 | 3.1155 | 2.5976 | H7 | 2.1041 | 1.0923 | 2.1501 | 3.6837 | 2.4243 | 2.5010 | | 1.7502 | 2.4923 | 3.0579 | 2.5027 | H8 | 2.1290 | 1.0894 | 2.1528 | 2.9165 | 2.4802 | 3.0474 | 1.7502 | | 2.4885 | 2.5115 | 3.0627 | H9 | 3.4750 | 2.1593 | 1.0876 | 4.2126 | 4.3023 | 3.7742 | 2.4923 | 2.4885 | | 1.7607 | 1.7597 | H10 | 2.8084 | 2.1572 | 1.0871 | 2.8480 | 3.8118 | 3.1155 | 3.0579 | 2.5115 | 1.7607 | | 1.7599 | H11 | 2.8013 | 2.1619 | 1.0894 | 3.6477 | 3.7838 | 2.5976 | 2.5027 | 3.0627 | 1.7597 | 1.7599 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.870 |
|
C1 |
C2 |
H7 |
106.819 |
| C1 |
C2 |
H8 |
108.908 |
|
C2 |
C1 |
Cl4 |
112.010 |
| C2 |
C1 |
H5 |
111.766 |
|
C2 |
C1 |
H6 |
111.714 |
| C2 |
C3 |
H9 |
110.949 |
|
C2 |
C3 |
H10 |
110.811 |
| C2 |
C3 |
H11 |
111.055 |
|
C3 |
C2 |
H7 |
109.933 |
| C3 |
C2 |
H8 |
110.322 |
|
Cl4 |
C1 |
H5 |
105.885 |
| Cl4 |
C1 |
H6 |
106.004 |
|
H5 |
C1 |
H6 |
109.131 |
| H7 |
C2 |
H8 |
106.690 |
|
H9 |
C3 |
H10 |
108.112 |
| H9 |
C3 |
H11 |
107.865 |
|
H10 |
C3 |
H11 |
107.918 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.159 |
|
|
|
| 2 |
C |
-0.218 |
|
|
|
| 3 |
C |
-0.352 |
|
|
|
| 4 |
Cl |
-0.176 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.141 |
|
|
|
| 7 |
H |
0.119 |
|
|
|
| 8 |
H |
0.132 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.132 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.640 |
1.275 |
0.288 |
2.097 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.647 |
0.236 |
0.133 |
| y |
0.236 |
-31.798 |
0.434 |
| z |
0.133 |
0.434 |
-32.618 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.440 |
0.236 |
0.133 |
| y |
0.236 |
1.835 |
0.434 |
| z |
0.133 |
0.434 |
0.605 |
|
| Polar |
|---|
| 3z2-r2 | 1.210 |
|---|
| x2-y2 | -2.849 |
|---|
| xy | 0.236 |
|---|
| xz | 0.133 |
|---|
| yz | 0.434 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.285 |
-0.436 |
-0.086 |
| y |
-0.436 |
7.145 |
0.118 |
| z |
-0.086 |
0.118 |
6.201 |
<r2> (average value of r
2) Å
2
| <r2> |
128.792 |
| (<r2>)1/2 |
11.349 |
