return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3OC2H5 (Ethane, methoxy-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-194.013092
Energy at 298.15K-194.022143
HF Energy-193.179709
Nuclear repulsion energy134.586120
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3184 3023 24.04      
2 A' 3183 3022 16.62      
3 A' 3105 2948 13.81      
4 A' 3040 2887 74.18      
5 A' 3029 2876 33.68      
6 A' 1559 1480 1.77      
7 A' 1539 1461 4.76      
8 A' 1528 1451 5.02      
9 A' 1496 1420 0.39      
10 A' 1445 1372 32.65      
11 A' 1408 1337 0.80      
12 A' 1254 1191 48.57      
13 A' 1192 1132 124.73      
14 A' 1134 1077 3.08      
15 A' 1066 1012 18.95      
16 A' 888 843 10.50      
17 A' 473 449 0.66      
18 A' 292 277 2.61      
19 A" 3187 3026 18.68      
20 A" 3091 2935 45.29      
21 A" 3055 2900 58.76      
22 A" 1520 1443 7.95      
23 A" 1512 1436 4.87      
24 A" 1321 1254 2.02      
25 A" 1217 1156 9.57      
26 A" 1185 1125 0.01      
27 A" 833 791 0.22      
28 A" 283 268 1.78      
29 A" 221 209 1.38      
30 A" 119 113 2.31      

Unscaled Zero Point Vibrational Energy (zpe) 24179.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 22955.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.94876 0.14135 0.13208

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.313 1.218 0.000
O2 0.002 0.715 0.000
C3 0.000 -0.693 0.000
C4 -1.428 -1.164 0.000
H5 1.250 2.299 0.000
H6 1.863 0.894 0.885
H7 1.863 0.894 -0.885
H8 0.528 -1.066 -0.881
H9 0.528 -1.066 0.881
H10 -1.470 -2.249 0.000
H11 -1.943 -0.795 0.880
H12 -1.943 -0.795 -0.880

Atom - Atom Distances (Å)
  C1 O2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.40442.31873.63161.08261.09141.09142.57102.57104.44543.92803.9280
O21.40441.40812.36122.01712.06912.06912.05582.05583.30882.61472.6147
C32.31871.40811.50383.24292.60282.60281.09281.09282.14012.13532.1353
C43.63162.36121.50384.37823.98163.98162.14792.14791.08511.08411.0841
H51.08262.01713.24294.37821.76991.76993.55283.55285.29914.53294.5329
H61.09142.06912.60283.98161.76991.76952.95672.37174.66584.16454.5229
H71.09142.06912.60283.98161.76991.76952.37172.95674.66584.52294.1645
H82.57102.05581.09282.14793.55282.95672.37171.76152.48343.04652.4863
H92.57102.05581.09282.14793.55282.37172.95671.76152.48342.48633.0465
H104.44543.30882.14011.08515.29914.66584.66582.48342.48341.76341.7634
H113.92802.61472.13531.08414.53294.16454.52293.04652.48631.76341.7595
H123.92802.61472.13531.08414.53294.52294.16452.48633.04651.76341.7595

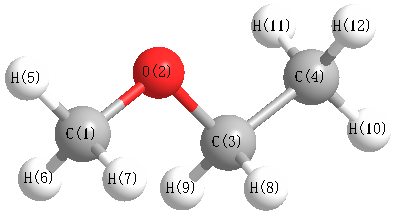
picture of Ethane, methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 111.061 O2 C1 H5 107.690
O2 C1 H6 111.384 O2 C1 H7 111.384
O2 C3 C4 108.324 O2 C3 H8 109.932
O2 C3 H9 109.932 C3 C4 H10 110.468
C3 C4 H11 110.151 C3 C4 H12 110.151
C4 C3 H8 110.625 C4 C3 H9 110.625
H5 C1 H6 109.002 H5 C1 H7 109.002
H6 C1 H7 108.329 H8 C3 H9 107.404
H10 C4 H11 108.765 H10 C4 H12 108.765
H11 C4 H12 108.493
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability