return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,3-dichlorobenzene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-1150.127131
Energy at 298.15K-1150.131469
HF Energy-1148.649284
Nuclear repulsion energy455.390430
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3276 3110 1.56      
2 A1 3270 3105 0.56      
3 A1 3233 3069 2.85      
4 A1 1631 1548 22.89      
5 A1 1448 1375 14.14      
6 A1 1168 1109 21.49      
7 A1 1108 1052 10.42      
8 A1 1025 973 3.08      
9 A1 677 643 1.38      
10 A1 410 389 2.13      
11 A1 198 188 0.34      
12 A2 934 887 0.00      
13 A2 564 536 0.00      
14 A2 208 197 0.00      
15 B1 1005 954 0.21      
16 B1 924 878 14.71      
17 B1 816 774 35.93      
18 B1 711 675 17.98      
19 B1 453 430 4.24      
20 B1 173 164 0.12      
21 B2 3264 3099 0.03      
22 B2 1634 1552 70.25      
23 B2 1505 1429 56.33      
24 B2 1478 1403 6.92      
25 B2 1296 1231 4.73      
26 B2 1201 1141 0.23      
27 B2 1114 1058 19.36      
28 B2 814 772 68.06      
29 B2 436 414 2.19      
30 B2 369 351 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 18172.2 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17252.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.09544 0.02900 0.02224

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.064
C2 0.000 1.206 1.380
C3 0.000 -1.206 1.380
C4 0.000 1.190 -0.006
C5 0.000 -1.190 -0.006
C6 0.000 0.000 -0.714
Cl7 0.000 2.682 -0.875
Cl8 0.000 -2.682 -0.875
H9 0.000 0.000 3.140
H10 0.000 2.144 1.904
H11 0.000 -2.144 1.904
H12 0.000 0.000 -1.788

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.38651.38652.38712.38712.77833.97903.97901.07622.14982.14983.8521
C21.38652.41191.38552.76742.41662.69524.49482.13381.07453.39063.3898
C31.38652.41192.76741.38552.41664.49482.69522.13383.39061.07453.3898
C42.38711.38552.76742.37951.38491.72743.96853.36322.13473.84182.1432
C52.38712.76741.38552.37951.38493.96851.72743.36323.84182.13472.1432
C62.77832.41662.41661.38491.38492.68722.68723.85453.38413.38411.0738
Cl73.97902.69524.49481.72743.96852.68725.36474.82872.83075.56922.8335
Cl83.97904.49482.69523.96851.72742.68725.36474.82875.56922.83072.8335
H91.07622.13382.13383.36323.36323.85454.82874.82872.47472.47474.9283
H102.14981.07453.39062.13473.84183.38412.83075.56922.47474.28784.2695
H112.14983.39061.07453.84182.13473.38415.56922.83072.47474.28784.2695
H123.85213.38983.38982.14322.14321.07382.83352.83354.92834.26954.2695

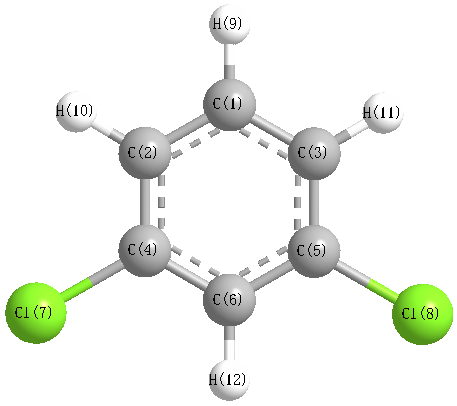
picture of 1,3-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 118.896 C1 C2 H10 121.237
C1 C3 C5 118.896 C1 C3 H11 121.237
C2 C1 C3 120.869 C2 C1 H9 119.565
C2 C4 C6 121.454 C2 C4 Cl7 119.551
C3 C1 H9 119.565 C3 C5 C6 121.454
C3 C5 Cl8 119.551 C4 C2 H10 119.867
C4 C6 C5 118.431 C4 C6 H12 120.785
C5 C3 H11 119.867 C5 C6 H12 120.785
C6 C4 Cl7 118.995 C6 C5 Cl8 118.995
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability