return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propanedial)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-266.743399
Energy at 298.15K-266.747563
HF Energy-265.730647
Nuclear repulsion energy156.935608
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3190 3028 3.68      
2 A 3110 2953 0.26      
3 A 3008 2856 23.00      
4 A 3004 2852 114.28      
5 A 1795 1704 42.14      
6 A 1754 1666 231.87      
7 A 1475 1401 12.38      
8 A 1438 1365 10.12      
9 A 1434 1362 3.23      
10 A 1280 1215 37.95      
11 A 1219 1157 10.50      
12 A 1115 1058 0.40      
13 A 1095 1039 65.62      
14 A 954 905 8.12      
15 A 910 864 39.08      
16 A 851 808 8.83      
17 A 553 525 14.31      
18 A 473 449 9.65      
19 A 284 269 6.21      
20 A 91 86 12.80      
21 A 22 21 13.13      

Unscaled Zero Point Vibrational Energy (zpe) 14526.6 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 13791.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.58029 0.09377 0.08821

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.016 0.720 0.363
C2 -1.057 -0.358 0.185
C3 1.234 0.299 -0.359
O4 -2.154 -0.166 -0.281
O5 1.971 -0.564 0.062
H6 0.219 0.778 1.423
H7 -0.409 1.662 0.001
H8 -0.734 -1.355 0.509
H9 1.428 0.787 -1.322

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.50911.50402.40232.38501.08721.08242.20092.2202
C21.50912.44501.20693.03802.10952.12901.09753.1238
C31.50402.44503.42101.21002.10582.16472.71371.0970
O42.40231.20693.42104.15863.07012.54322.01343.8502
O52.38503.03801.21004.15862.59243.25872.85422.0086
H61.08722.10952.10583.07012.59241.78802.50852.9994
H71.08242.12902.16472.54323.25871.78803.07672.4268
H82.20091.09752.71372.01342.85422.50853.07673.5520
H92.22023.12381.09703.85022.00862.99942.42683.5520

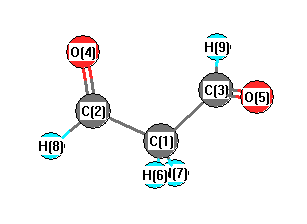
picture of propanedial state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 124.004 C1 C2 H8 114.284
C1 C3 O5 122.621 C1 C3 H9 116.338
C2 C1 C3 108.482 C2 C1 H6 107.574
C2 C1 H7 109.374 C3 C1 H6 107.629
C3 C1 H7 112.622 O4 C2 H8 121.710
O5 C3 H9 120.990 H6 C1 H7 110.998
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/cc-pVTZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.160      
2 C 0.186      
3 C 0.173      
4 O -0.330      
5 O -0.342      
6 H 0.128      
7 H 0.129      
8 H 0.107      
9 H 0.110      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000